Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -1070.826835 |
| Energy at 298.15K | -1070.829571 |
| HF Energy | -1070.826835 |
| Nuclear repulsion energy | 265.589415 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3337 |
3024 |
1.88 |
|
|
|
| 2 |
A' |
3163 |
2867 |
32.28 |
|
|
|
| 3 |
A' |
2016 |
1827 |
244.83 |
|
|
|
| 4 |
A' |
1522 |
1380 |
7.19 |
|
|
|
| 5 |
A' |
1345 |
1219 |
13.24 |
|
|
|
| 6 |
A' |
1157 |
1048 |
25.69 |
|
|
|
| 7 |
A' |
855 |
775 |
33.26 |
|
|
|
| 8 |
A' |
474 |
429 |
6.51 |
|
|
|
| 9 |
A' |
357 |
323 |
27.74 |
|
|
|
| 10 |
A' |
275 |
249 |
1.99 |
|
|
|
| 11 |
A" |
1361 |
1234 |
32.18 |
|
|
|
| 12 |
A" |
1137 |
1031 |
16.97 |
|
|
|
| 13 |
A" |
812 |
736 |
123.96 |
|
|
|
| 14 |
A" |
310 |
281 |
4.15 |
|
|
|
| 15 |
A" |
91 |
83 |
11.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9104.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 8252.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.385 |
-0.013 |
0.000 |
| C2 |
-0.170 |
1.399 |
0.000 |
| H3 |
1.457 |
0.001 |
0.000 |
| Cl4 |
-0.170 |
-0.838 |
1.460 |
| Cl5 |
-0.170 |
-0.838 |
-1.460 |
| O6 |
0.534 |
2.339 |
0.000 |
| H7 |
-1.257 |
1.470 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5176 | 1.0721 | 1.7669 | 1.7669 | 2.3571 | 2.2132 |
C2 | 1.5176 | | 2.1453 | 2.6719 | 2.6719 | 1.1745 | 1.0899 | H3 | 1.0721 | 2.1453 | | 2.3421 | 2.3421 | 2.5137 | 3.0866 | Cl4 | 1.7669 | 2.6719 | 2.3421 | | 2.9208 | 3.5673 | 2.9401 | Cl5 | 1.7669 | 2.6719 | 2.3421 | 2.9208 | | 3.5673 | 2.9401 | O6 | 2.3571 | 1.1745 | 2.5137 | 3.5673 | 3.5673 | | 1.9914 | H7 | 2.2132 | 1.0899 | 3.0866 | 2.9401 | 2.9401 | 1.9914 | |
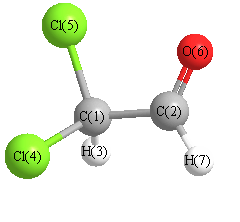 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
121.709 |
|
C1 |
C2 |
H7 |
115.183 |
| C2 |
C1 |
H3 |
110.692 |
|
C2 |
C1 |
Cl4 |
108.643 |
| C2 |
C1 |
Cl5 |
108.643 |
|
H3 |
C1 |
Cl4 |
108.689 |
| H3 |
C1 |
Cl5 |
108.689 |
|
Cl4 |
C1 |
Cl5 |
111.492 |
| O6 |
C2 |
H7 |
123.108 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.168 |
|
|
|
| 2 |
C |
0.239 |
|
|
|
| 3 |
H |
0.103 |
|
|
|
| 4 |
Cl |
-0.106 |
|
|
|
| 5 |
Cl |
-0.106 |
|
|
|
| 6 |
O |
-0.350 |
|
|
|
| 7 |
H |
0.052 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.488 |
-0.921 |
0.000 |
1.042 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.791 |
-3.456 |
0.000 |
| y |
-3.456 |
-49.544 |
0.000 |
| z |
0.000 |
0.000 |
-43.081 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.521 |
-3.456 |
0.000 |
| y |
-3.456 |
-7.608 |
0.000 |
| z |
0.000 |
0.000 |
2.087 |
|
| Polar |
|---|
| 3z2-r2 | 4.174 |
|---|
| x2-y2 | 8.753 |
|---|
| xy | -3.456 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.795 |
0.695 |
0.000 |
| y |
0.695 |
7.429 |
0.000 |
| z |
0.000 |
0.000 |
7.752 |
<r2> (average value of r
2) Å
2
| <r2> |
190.148 |
| (<r2>)1/2 |
13.789 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -1070.823297 |
| Energy at 298.15K | |
| HF Energy | -1070.823297 |
| Nuclear repulsion energy | 268.159014 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3279 |
2972 |
6.59 |
71.51 |
0.19 |
0.32 |
| 2 |
A |
3142 |
2848 |
45.15 |
110.96 |
0.29 |
0.45 |
| 3 |
A |
2027 |
1837 |
206.31 |
17.95 |
0.41 |
0.58 |
| 4 |
A |
1522 |
1380 |
22.95 |
2.54 |
0.37 |
0.54 |
| 5 |
A |
1404 |
1273 |
7.96 |
3.02 |
0.75 |
0.86 |
| 6 |
A |
1345 |
1219 |
18.73 |
5.16 |
0.71 |
0.83 |
| 7 |
A |
1145 |
1038 |
9.83 |
2.27 |
0.47 |
0.64 |
| 8 |
A |
1028 |
932 |
10.92 |
3.60 |
0.65 |
0.79 |
| 9 |
A |
897 |
813 |
86.90 |
9.26 |
0.74 |
0.85 |
| 10 |
A |
695 |
630 |
48.59 |
12.59 |
0.12 |
0.22 |
| 11 |
A |
665 |
603 |
34.93 |
6.59 |
0.42 |
0.59 |
| 12 |
A |
376 |
341 |
1.36 |
2.72 |
0.17 |
0.29 |
| 13 |
A |
297 |
269 |
4.17 |
3.31 |
0.62 |
0.77 |
| 14 |
A |
237 |
215 |
3.72 |
1.11 |
0.70 |
0.83 |
| 15 |
A |
91 |
83 |
14.80 |
1.47 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 9074.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 8225.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.091 |
-0.034 |
0.507 |
| C2 |
-0.751 |
-1.263 |
0.178 |
| H3 |
0.207 |
0.034 |
1.575 |
| Cl4 |
1.714 |
-0.297 |
-0.161 |
| Cl5 |
-0.623 |
1.462 |
-0.057 |
| O6 |
-1.818 |
-1.231 |
-0.304 |
| H7 |
-0.258 |
-2.196 |
0.456 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5250 | 1.0767 | 1.7755 | 1.7503 | 2.3939 | 2.1907 |
C2 | 1.5250 | | 2.1332 | 2.6695 | 2.7372 | 1.1707 | 1.0922 | H3 | 1.0767 | 2.1332 | | 2.3233 | 2.3212 | 3.0379 | 2.5384 | Cl4 | 1.7755 | 2.6695 | 2.3233 | | 2.9271 | 3.6561 | 2.8064 | Cl5 | 1.7503 | 2.7372 | 2.3212 | 2.9271 | | 2.9560 | 3.7117 | O6 | 2.3939 | 1.1707 | 3.0379 | 3.6561 | 2.9560 | | 1.9858 | H7 | 2.1907 | 1.0922 | 2.5384 | 2.8064 | 3.7117 | 1.9858 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.733 |
|
C1 |
C2 |
H7 |
112.610 |
| C2 |
C1 |
H3 |
108.936 |
|
C2 |
C1 |
Cl4 |
107.716 |
| C2 |
C1 |
Cl5 |
113.202 |
|
H3 |
C1 |
Cl4 |
106.500 |
| H3 |
C1 |
Cl5 |
108.009 |
|
Cl4 |
C1 |
Cl5 |
112.231 |
| O6 |
C2 |
H7 |
122.652 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.166 |
|
|
|
| 2 |
C |
0.220 |
|
|
|
| 3 |
H |
0.081 |
|
|
|
| 4 |
Cl |
-0.114 |
|
|
|
| 5 |
Cl |
-0.072 |
|
|
|
| 6 |
O |
-0.329 |
|
|
|
| 7 |
H |
0.049 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.488 |
-0.889 |
2.089 |
2.714 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.746 |
-2.257 |
-1.439 |
| y |
-2.257 |
-41.602 |
-1.689 |
| z |
-1.439 |
-1.689 |
-40.287 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.802 |
-2.257 |
-1.439 |
| y |
-2.257 |
2.915 |
-1.689 |
| z |
-1.439 |
-1.689 |
4.887 |
|
| Polar |
|---|
| 3z2-r2 | 9.774 |
|---|
| x2-y2 | -7.144 |
|---|
| xy | -2.257 |
|---|
| xz | -1.439 |
|---|
| yz | -1.689 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.234 |
-0.661 |
0.137 |
| y |
-0.661 |
7.241 |
-0.264 |
| z |
0.137 |
-0.264 |
5.326 |
<r2> (average value of r
2) Å
2
| <r2> |
183.724 |
| (<r2>)1/2 |
13.555 |
