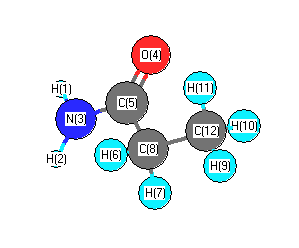
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3967 |
3595 |
52.79 |
|
|
|
| 2 |
A |
3830 |
3472 |
53.90 |
|
|
|
| 3 |
A |
3255 |
2950 |
34.99 |
|
|
|
| 4 |
A |
3236 |
2933 |
34.24 |
|
|
|
| 5 |
A |
3189 |
2891 |
14.98 |
|
|
|
| 6 |
A |
3177 |
2880 |
26.75 |
|
|
|
| 7 |
A |
3161 |
2865 |
32.99 |
|
|
|
| 8 |
A |
1935 |
1754 |
411.22 |
|
|
|
| 9 |
A |
1769 |
1603 |
121.66 |
|
|
|
| 10 |
A |
1618 |
1467 |
10.63 |
|
|
|
| 11 |
A |
1608 |
1457 |
5.75 |
|
|
|
| 12 |
A |
1590 |
1441 |
5.56 |
|
|
|
| 13 |
A |
1549 |
1404 |
17.14 |
|
|
|
| 14 |
A |
1531 |
1387 |
54.21 |
|
|
|
| 15 |
A |
1399 |
1268 |
0.01 |
|
|
|
| 16 |
A |
1382 |
1253 |
163.37 |
|
|
|
| 17 |
A |
1218 |
1104 |
2.45 |
|
|
|
| 18 |
A |
1205 |
1092 |
0.46 |
|
|
|
| 19 |
A |
1143 |
1036 |
11.43 |
|
|
|
| 20 |
A |
1071 |
971 |
2.03 |
|
|
|
| 21 |
A |
885 |
802 |
9.06 |
|
|
|
| 22 |
A |
850 |
771 |
1.57 |
|
|
|
| 23 |
A |
669 |
606 |
8.87 |
|
|
|
| 24 |
A |
653 |
592 |
13.92 |
|
|
|
| 25 |
A |
538 |
488 |
2.84 |
|
|
|
| 26 |
A |
476 |
432 |
4.44 |
|
|
|
| 27 |
A |
274 |
248 |
10.15 |
|
|
|
| 28 |
A |
234 |
212 |
0.02 |
|
|
|
| 29 |
A |
117 |
106 |
228.90 |
|
|
|
| 30 |
A |
20 |
18 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23774.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 21548.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.157 |
|
|
|
| 2 |
C |
-0.082 |
|
|
|
| 3 |
C |
0.455 |
|
|
|
| 4 |
O |
-0.488 |
|
|
|
| 5 |
N |
-0.428 |
|
|
|
| 6 |
H |
0.059 |
|
|
|
| 7 |
H |
0.067 |
|
|
|
| 8 |
H |
0.063 |
|
|
|
| 9 |
H |
0.055 |
|
|
|
| 10 |
H |
0.057 |
|
|
|
| 11 |
H |
0.189 |
|
|
|
| 12 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.725 |
-3.829 |
0.143 |
3.900 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.818 |
-2.507 |
0.286 |
| y |
-2.507 |
-33.533 |
-0.194 |
| z |
0.286 |
-0.194 |
-31.406 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.652 |
-2.507 |
0.286 |
| y |
-2.507 |
-4.921 |
-0.194 |
| z |
0.286 |
-0.194 |
-1.731 |
|
| Polar |
|---|
| 3z2-r2 | -3.462 |
|---|
| x2-y2 | 7.715 |
|---|
| xy | -2.507 |
|---|
| xz | 0.286 |
|---|
| yz | -0.194 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.294 |
-0.289 |
-0.012 |
| y |
-0.289 |
6.864 |
0.000 |
| z |
-0.012 |
0.000 |
5.190 |
<r2> (average value of r
2) Å
2
| <r2> |
127.168 |
| (<r2>)1/2 |
11.277 |