Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -302.801630 |
| Energy at 298.15K | -302.807394 |
| HF Energy | -302.801630 |
| Nuclear repulsion energy | 182.189534 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4119 |
3734 |
102.40 |
|
|
|
| 2 |
A |
4106 |
3721 |
131.22 |
|
|
|
| 3 |
A |
3209 |
2908 |
21.73 |
|
|
|
| 4 |
A |
3180 |
2882 |
33.72 |
|
|
|
| 5 |
A |
1990 |
1804 |
415.25 |
|
|
|
| 6 |
A |
1623 |
1471 |
8.68 |
|
|
|
| 7 |
A |
1611 |
1460 |
0.15 |
|
|
|
| 8 |
A |
1461 |
1325 |
140.76 |
|
|
|
| 9 |
A |
1396 |
1266 |
54.80 |
|
|
|
| 10 |
A |
1373 |
1245 |
0.89 |
|
|
|
| 11 |
A |
1300 |
1179 |
151.83 |
|
|
|
| 12 |
A |
1234 |
1119 |
253.78 |
|
|
|
| 13 |
A |
1142 |
1035 |
3.19 |
|
|
|
| 14 |
A |
929 |
842 |
39.20 |
|
|
|
| 15 |
A |
711 |
645 |
27.94 |
|
|
|
| 16 |
A |
695 |
630 |
140.10 |
|
|
|
| 17 |
A |
547 |
496 |
21.59 |
|
|
|
| 18 |
A |
511 |
463 |
23.55 |
|
|
|
| 19 |
A |
300 |
272 |
12.14 |
|
|
|
| 20 |
A |
290 |
263 |
88.09 |
|
|
|
| 21 |
A |
79 |
72 |
19.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15902.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 14414.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.744 |
-0.729 |
0.003 |
| C2 |
0.516 |
0.098 |
-0.002 |
| O3 |
1.609 |
-0.640 |
-0.003 |
| O4 |
0.526 |
1.281 |
0.002 |
| O5 |
-1.878 |
0.062 |
-0.005 |
| H6 |
-0.718 |
-1.385 |
-0.859 |
| H7 |
-0.722 |
-1.354 |
0.889 |
| H8 |
-1.621 |
0.970 |
0.002 |
| H9 |
2.366 |
-0.074 |
0.005 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5068 | 2.3549 | 2.3769 | 1.3826 | 1.0840 | 1.0848 | 1.9114 | 3.1785 |
C2 | 1.5068 | | 1.3188 | 1.1827 | 2.3941 | 2.1109 | 2.1061 | 2.3083 | 1.8585 | O3 | 2.3549 | 1.3188 | | 2.2044 | 3.5571 | 2.5897 | 2.5964 | 3.6093 | 0.9452 | O4 | 2.3769 | 1.1827 | 2.2044 | | 2.6952 | 3.0651 | 3.0475 | 2.1701 | 2.2849 | O5 | 1.3826 | 2.3941 | 3.5571 | 2.6952 | | 2.0420 | 2.0351 | 0.9427 | 4.2465 | H6 | 1.0840 | 2.1109 | 2.5897 | 3.0651 | 2.0420 | | 1.7486 | 2.6648 | 3.4613 | H7 | 1.0848 | 2.1061 | 2.5964 | 3.0475 | 2.0351 | 1.7486 | | 2.6449 | 3.4583 | H8 | 1.9114 | 2.3083 | 3.6093 | 2.1701 | 0.9427 | 2.6648 | 2.6449 | | 4.1221 | H9 | 3.1785 | 1.8585 | 0.9452 | 2.2849 | 4.2465 | 3.4613 | 3.4583 | 4.1221 | |
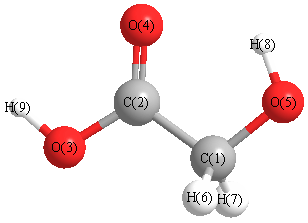 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
112.737 |
|
C1 |
C2 |
O4 |
123.766 |
| C1 |
O5 |
H8 |
109.107 |
|
C2 |
C1 |
O5 |
111.835 |
| C2 |
C1 |
H6 |
108.021 |
|
C2 |
C1 |
H7 |
107.606 |
| C2 |
O3 |
H9 |
109.239 |
|
O3 |
C2 |
O4 |
123.495 |
| O5 |
C1 |
H6 |
111.181 |
|
O5 |
C1 |
H7 |
110.558 |
| H6 |
C1 |
H7 |
107.457 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.199 |
|
|
|
| 2 |
C |
0.484 |
|
|
|
| 3 |
O |
-0.373 |
|
|
|
| 4 |
O |
-0.465 |
|
|
|
| 5 |
O |
-0.435 |
|
|
|
| 6 |
H |
0.055 |
|
|
|
| 7 |
H |
0.054 |
|
|
|
| 8 |
H |
0.238 |
|
|
|
| 9 |
H |
0.243 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.999 |
-1.140 |
0.029 |
2.302 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.600 |
-0.417 |
-0.050 |
| y |
-0.417 |
-30.994 |
0.012 |
| z |
-0.050 |
0.012 |
-28.094 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.944 |
-0.417 |
-0.050 |
| y |
-0.417 |
-2.648 |
0.012 |
| z |
-0.050 |
0.012 |
1.703 |
|
| Polar |
|---|
| 3z2-r2 | 3.407 |
|---|
| x2-y2 | 2.395 |
|---|
| xy | -0.417 |
|---|
| xz | -0.050 |
|---|
| yz | 0.012 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.126 |
-0.066 |
0.000 |
| y |
-0.066 |
5.250 |
0.000 |
| z |
0.000 |
0.000 |
3.593 |
<r2> (average value of r
2) Å
2
| <r2> |
109.413 |
| (<r2>)1/2 |
10.460 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -302.801630 |
| Energy at 298.15K | -302.807394 |
| HF Energy | -302.801630 |
| Nuclear repulsion energy | 182.192374 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4119 |
3734 |
102.44 |
|
|
|
| 2 |
A |
4106 |
3721 |
131.17 |
|
|
|
| 3 |
A |
3208 |
2908 |
21.72 |
|
|
|
| 4 |
A |
3180 |
2882 |
33.73 |
|
|
|
| 5 |
A |
1990 |
1804 |
415.23 |
|
|
|
| 6 |
A |
1623 |
1471 |
8.67 |
|
|
|
| 7 |
A |
1611 |
1460 |
0.15 |
|
|
|
| 8 |
A |
1461 |
1325 |
140.35 |
|
|
|
| 9 |
A |
1396 |
1265 |
55.57 |
|
|
|
| 10 |
A |
1374 |
1245 |
0.24 |
|
|
|
| 11 |
A |
1300 |
1179 |
151.87 |
|
|
|
| 12 |
A |
1234 |
1119 |
254.16 |
|
|
|
| 13 |
A |
1142 |
1035 |
3.15 |
|
|
|
| 14 |
A |
929 |
842 |
39.18 |
|
|
|
| 15 |
A |
711 |
645 |
27.86 |
|
|
|
| 16 |
A |
695 |
630 |
140.23 |
|
|
|
| 17 |
A |
547 |
496 |
21.51 |
|
|
|
| 18 |
A |
511 |
463 |
23.57 |
|
|
|
| 19 |
A |
299 |
271 |
8.52 |
|
|
|
| 20 |
A |
290 |
263 |
91.60 |
|
|
|
| 21 |
A |
79 |
71 |
19.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15902.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 14414.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.771 |
-0.677 |
0.190 |
| C2 |
0.520 |
0.085 |
-0.008 |
| O3 |
1.590 |
-0.672 |
-0.112 |
| O4 |
0.593 |
1.266 |
0.092 |
| O5 |
-1.883 |
0.060 |
-0.191 |
| H6 |
-0.737 |
-1.595 |
-0.376 |
| H7 |
-0.826 |
-0.952 |
1.243 |
| H8 |
-1.686 |
0.979 |
-0.099 |
| H9 |
2.352 |
-0.115 |
-0.170 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5132 | 2.3805 | 2.3769 | 1.3876 | 1.0790 | 1.0900 | 1.9142 | 3.1942 |
C2 | 1.5132 | | 1.3147 | 1.1873 | 2.4105 | 2.1312 | 2.1114 | 2.3820 | 1.8498 | O3 | 2.3805 | 1.3147 | | 2.1889 | 3.5503 | 2.5178 | 2.7847 | 3.6684 | 0.9460 | O4 | 2.3769 | 1.1873 | 2.1889 | | 2.7689 | 3.1903 | 2.8748 | 2.3049 | 2.2513 | O5 | 1.3876 | 2.4105 | 3.5503 | 2.7689 | | 2.0219 | 2.0491 | 0.9445 | 4.2389 | H6 | 1.0790 | 2.1312 | 2.5178 | 3.1903 | 2.0219 | | 1.7446 | 2.7577 | 3.4323 | H7 | 1.0900 | 2.1114 | 2.7847 | 2.8748 | 2.0491 | 1.7446 | | 2.5043 | 3.5782 | H8 | 1.9142 | 2.3820 | 3.6684 | 2.3049 | 0.9445 | 2.7577 | 2.5043 | | 4.1840 | H9 | 3.1942 | 1.8498 | 0.9460 | 2.2513 | 4.2389 | 3.4323 | 3.5782 | 4.1840 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
114.483 |
|
C1 |
C2 |
O4 |
122.871 |
| C1 |
O5 |
H8 |
108.853 |
|
C2 |
C1 |
O5 |
112.329 |
| C2 |
C1 |
H6 |
109.462 |
|
C2 |
C1 |
H7 |
107.290 |
| C2 |
O3 |
H9 |
108.717 |
|
O3 |
C2 |
O4 |
121.974 |
| O5 |
C1 |
H6 |
109.480 |
|
O5 |
C1 |
H7 |
111.029 |
| H6 |
C1 |
H7 |
107.091 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.199 |
|
|
|
| 2 |
C |
0.484 |
|
|
|
| 3 |
O |
-0.373 |
|
|
|
| 4 |
O |
-0.465 |
|
|
|
| 5 |
O |
-0.435 |
|
|
|
| 6 |
H |
0.054 |
|
|
|
| 7 |
H |
0.054 |
|
|
|
| 8 |
H |
0.238 |
|
|
|
| 9 |
H |
0.243 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.999 |
-1.139 |
-0.002 |
2.301 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.602 |
-0.416 |
0.004 |
| y |
-0.416 |
-30.994 |
-0.000 |
| z |
0.004 |
-0.000 |
-28.093 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.942 |
-0.416 |
0.004 |
| y |
-0.416 |
-2.646 |
-0.000 |
| z |
0.004 |
-0.000 |
1.705 |
|
| Polar |
|---|
| 3z2-r2 | 3.409 |
|---|
| x2-y2 | 2.392 |
|---|
| xy | -0.416 |
|---|
| xz | 0.004 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.125 |
-0.066 |
0.000 |
| y |
-0.066 |
5.250 |
0.000 |
| z |
0.000 |
0.000 |
3.593 |
<r2> (average value of r
2) Å
2
| <r2> |
109.410 |
| (<r2>)1/2 |
10.460 |
