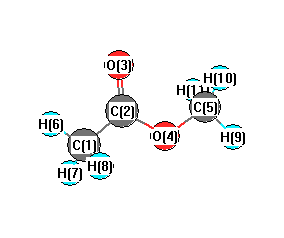
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3295 |
2987 |
14.48 |
|
|
|
| 2 |
A' |
3292 |
2984 |
23.47 |
|
|
|
| 3 |
A' |
3200 |
2901 |
38.56 |
|
|
|
| 4 |
A' |
3190 |
2891 |
4.29 |
|
|
|
| 5 |
A' |
1975 |
1790 |
358.76 |
|
|
|
| 6 |
A' |
1625 |
1473 |
6.94 |
|
|
|
| 7 |
A' |
1613 |
1462 |
27.05 |
|
|
|
| 8 |
A' |
1586 |
1437 |
13.65 |
|
|
|
| 9 |
A' |
1535 |
1392 |
80.18 |
|
|
|
| 10 |
A' |
1412 |
1280 |
428.41 |
|
|
|
| 11 |
A' |
1329 |
1204 |
5.55 |
|
|
|
| 12 |
A' |
1192 |
1080 |
81.68 |
|
|
|
| 13 |
A' |
1071 |
971 |
4.64 |
|
|
|
| 14 |
A' |
942 |
854 |
17.91 |
|
|
|
| 15 |
A' |
695 |
630 |
10.87 |
|
|
|
| 16 |
A' |
455 |
412 |
7.74 |
|
|
|
| 17 |
A' |
303 |
274 |
13.88 |
|
|
|
| 18 |
A" |
3272 |
2966 |
30.30 |
|
|
|
| 19 |
A" |
3249 |
2945 |
11.08 |
|
|
|
| 20 |
A" |
1613 |
1462 |
7.64 |
|
|
|
| 21 |
A" |
1597 |
1447 |
6.44 |
|
|
|
| 22 |
A" |
1291 |
1170 |
3.39 |
|
|
|
| 23 |
A" |
1174 |
1064 |
9.04 |
|
|
|
| 24 |
A" |
672 |
609 |
11.75 |
|
|
|
| 25 |
A" |
189 |
171 |
5.17 |
|
|
|
| 26 |
A" |
161 |
146 |
2.67 |
|
|
|
| 27 |
A" |
90 |
81 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21007.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19041.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.197 |
|
|
|
| 2 |
C |
0.522 |
|
|
|
| 3 |
O |
-0.463 |
|
|
|
| 4 |
O |
-0.377 |
|
|
|
| 5 |
C |
0.013 |
|
|
|
| 6 |
H |
0.102 |
|
|
|
| 7 |
H |
0.079 |
|
|
|
| 8 |
H |
0.079 |
|
|
|
| 9 |
H |
0.086 |
|
|
|
| 10 |
H |
0.078 |
|
|
|
| 11 |
H |
0.078 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.796 |
-1.048 |
0.000 |
2.079 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.929 |
4.955 |
0.000 |
| y |
4.955 |
-26.631 |
0.000 |
| z |
0.000 |
0.000 |
-29.069 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.079 |
4.955 |
0.000 |
| y |
4.955 |
4.368 |
0.000 |
| z |
0.000 |
0.000 |
0.711 |
|
| Polar |
|---|
| 3z2-r2 | 1.421 |
|---|
| x2-y2 | -6.298 |
|---|
| xy | 4.955 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.157 |
0.274 |
0.000 |
| y |
0.274 |
6.662 |
0.000 |
| z |
0.000 |
0.000 |
4.838 |
<r2> (average value of r
2) Å
2
| <r2> |
118.311 |
| (<r2>)1/2 |
10.877 |