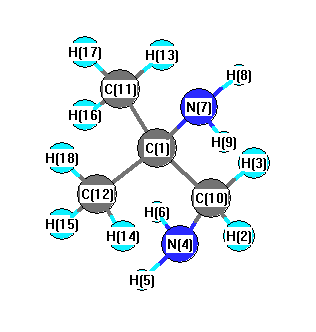
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3749 |
3398 |
1.48 |
|
|
|
| 2 |
A' |
3717 |
3369 |
0.52 |
|
|
|
| 3 |
A' |
3241 |
2938 |
67.47 |
|
|
|
| 4 |
A' |
3216 |
2915 |
76.81 |
|
|
|
| 5 |
A' |
3162 |
2866 |
5.88 |
|
|
|
| 6 |
A' |
3151 |
2856 |
44.90 |
|
|
|
| 7 |
A' |
1796 |
1628 |
24.69 |
|
|
|
| 8 |
A' |
1791 |
1624 |
35.62 |
|
|
|
| 9 |
A' |
1629 |
1476 |
6.22 |
|
|
|
| 10 |
A' |
1622 |
1470 |
6.26 |
|
|
|
| 11 |
A' |
1606 |
1455 |
2.13 |
|
|
|
| 12 |
A' |
1547 |
1402 |
10.33 |
|
|
|
| 13 |
A' |
1501 |
1361 |
4.32 |
|
|
|
| 14 |
A' |
1366 |
1238 |
9.28 |
|
|
|
| 15 |
A' |
1330 |
1205 |
27.57 |
|
|
|
| 16 |
A' |
1175 |
1065 |
26.94 |
|
|
|
| 17 |
A' |
1095 |
992 |
1.79 |
|
|
|
| 18 |
A' |
1000 |
906 |
11.94 |
|
|
|
| 19 |
A' |
932 |
845 |
277.06 |
|
|
|
| 20 |
A' |
896 |
812 |
111.64 |
|
|
|
| 21 |
A' |
770 |
698 |
16.37 |
|
|
|
| 22 |
A' |
550 |
499 |
11.87 |
|
|
|
| 23 |
A' |
457 |
414 |
9.91 |
|
|
|
| 24 |
A' |
388 |
351 |
0.06 |
|
|
|
| 25 |
A' |
285 |
259 |
0.30 |
|
|
|
| 26 |
A' |
275 |
249 |
4.81 |
|
|
|
| 27 |
A" |
3831 |
3473 |
2.93 |
|
|
|
| 28 |
A" |
3795 |
3440 |
0.49 |
|
|
|
| 29 |
A" |
3233 |
2931 |
0.49 |
|
|
|
| 30 |
A" |
3212 |
2912 |
21.18 |
|
|
|
| 31 |
A" |
3185 |
2887 |
51.36 |
|
|
|
| 32 |
A" |
3153 |
2858 |
56.62 |
|
|
|
| 33 |
A" |
1614 |
1463 |
0.50 |
|
|
|
| 34 |
A" |
1602 |
1452 |
0.02 |
|
|
|
| 35 |
A" |
1535 |
1392 |
10.74 |
|
|
|
| 36 |
A" |
1510 |
1368 |
0.74 |
|
|
|
| 37 |
A" |
1451 |
1315 |
1.04 |
|
|
|
| 38 |
A" |
1272 |
1153 |
0.19 |
|
|
|
| 39 |
A" |
1129 |
1023 |
0.74 |
|
|
|
| 40 |
A" |
1084 |
983 |
0.64 |
|
|
|
| 41 |
A" |
1014 |
919 |
0.03 |
|
|
|
| 42 |
A" |
921 |
835 |
0.01 |
|
|
|
| 43 |
A" |
481 |
436 |
2.06 |
|
|
|
| 44 |
A" |
371 |
336 |
10.75 |
|
|
|
| 45 |
A" |
312 |
283 |
1.69 |
|
|
|
| 46 |
A" |
279 |
253 |
75.56 |
|
|
|
| 47 |
A" |
234 |
212 |
0.06 |
|
|
|
| 48 |
A" |
122 |
111 |
1.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38791.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 35160.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.429 |
|
|
|
| 2 |
H |
0.043 |
|
|
|
| 3 |
H |
0.043 |
|
|
|
| 4 |
N |
-0.409 |
|
|
|
| 5 |
H |
0.132 |
|
|
|
| 6 |
H |
0.132 |
|
|
|
| 7 |
N |
-0.455 |
|
|
|
| 8 |
H |
0.128 |
|
|
|
| 9 |
H |
0.128 |
|
|
|
| 10 |
C |
0.005 |
|
|
|
| 11 |
C |
-0.263 |
|
|
|
| 12 |
C |
-0.263 |
|
|
|
| 13 |
H |
0.055 |
|
|
|
| 14 |
H |
0.055 |
|
|
|
| 15 |
H |
0.042 |
|
|
|
| 16 |
H |
0.042 |
|
|
|
| 17 |
H |
0.079 |
|
|
|
| 18 |
H |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.069 |
-0.268 |
0.000 |
0.277 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.461 |
2.474 |
0.000 |
| y |
2.474 |
-36.782 |
0.000 |
| z |
0.000 |
0.000 |
-37.495 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.323 |
2.474 |
0.000 |
| y |
2.474 |
6.697 |
0.000 |
| z |
0.000 |
0.000 |
5.626 |
|
| Polar |
|---|
| 3z2-r2 | 11.252 |
|---|
| x2-y2 | -12.680 |
|---|
| xy | 2.474 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.138 |
-0.243 |
0.000 |
| y |
-0.243 |
9.273 |
0.000 |
| z |
0.000 |
0.000 |
9.141 |
<r2> (average value of r
2) Å
2
| <r2> |
182.326 |
| (<r2>)1/2 |
13.503 |