Jump to
S1C2
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -217.063418 |
| Energy at 298.15K | |
| HF Energy | -216.822109 |
| Nuclear repulsion energy | 116.554523 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3270 |
3270 |
4.00 |
49.89 |
0.71 |
0.83 |
| 2 |
A' |
3182 |
3182 |
3.91 |
164.46 |
0.12 |
0.21 |
| 3 |
A' |
3167 |
3167 |
9.44 |
36.00 |
0.58 |
0.73 |
| 4 |
A' |
3049 |
3049 |
33.53 |
175.21 |
0.10 |
0.18 |
| 5 |
A' |
1715 |
1715 |
4.98 |
21.72 |
0.07 |
0.14 |
| 6 |
A' |
1511 |
1511 |
3.29 |
12.93 |
0.55 |
0.71 |
| 7 |
A' |
1462 |
1462 |
5.79 |
8.79 |
0.42 |
0.59 |
| 8 |
A' |
1427 |
1427 |
11.70 |
4.42 |
0.75 |
0.86 |
| 9 |
A' |
1324 |
1324 |
0.17 |
17.91 |
0.26 |
0.41 |
| 10 |
A' |
1137 |
1137 |
48.61 |
1.61 |
0.72 |
0.84 |
| 11 |
A' |
1015 |
1015 |
42.79 |
5.00 |
0.67 |
0.80 |
| 12 |
A' |
915 |
915 |
2.34 |
5.03 |
0.07 |
0.12 |
| 13 |
A' |
613 |
613 |
6.98 |
1.72 |
0.73 |
0.85 |
| 14 |
A' |
266 |
266 |
2.27 |
1.22 |
0.51 |
0.68 |
| 15 |
A" |
3087 |
3087 |
22.49 |
84.06 |
0.75 |
0.86 |
| 16 |
A" |
1277 |
1277 |
0.00 |
5.79 |
0.75 |
0.86 |
| 17 |
A" |
1055 |
1055 |
12.65 |
0.89 |
0.75 |
0.86 |
| 18 |
A" |
1027 |
1027 |
10.87 |
0.10 |
0.75 |
0.86 |
| 19 |
A" |
962 |
962 |
43.95 |
1.69 |
0.75 |
0.86 |
| 20 |
A" |
565 |
565 |
10.68 |
3.31 |
0.75 |
0.86 |
| 21 |
A" |
175 |
175 |
2.98 |
1.87 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16100.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16100.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.942 |
-0.206 |
0.000 |
| C2 |
0.000 |
0.951 |
0.000 |
| C3 |
1.324 |
0.852 |
0.000 |
| F4 |
-0.267 |
-1.418 |
0.000 |
| H5 |
1.947 |
1.733 |
0.000 |
| H6 |
1.812 |
-0.111 |
0.000 |
| H7 |
-0.480 |
1.923 |
0.000 |
| H8 |
-1.582 |
-0.184 |
0.884 |
| H9 |
-1.582 |
-0.184 |
-0.884 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4918 | 2.5004 | 1.3877 | 3.4791 | 2.7555 | 2.1786 | 1.0918 | 1.0918 |
C2 | 1.4918 | | 1.3275 | 2.3843 | 2.0980 | 2.1001 | 1.0842 | 2.1387 | 2.1387 | C3 | 2.5004 | 1.3275 | | 2.7720 | 1.0794 | 1.0794 | 2.0978 | 3.2093 | 3.2093 | F4 | 1.3877 | 2.3843 | 2.7720 | | 3.8514 | 2.4559 | 3.3483 | 2.0085 | 2.0085 | H5 | 3.4791 | 2.0980 | 1.0794 | 3.8514 | | 1.8491 | 2.4339 | 4.1123 | 4.1123 | H6 | 2.7555 | 2.1001 | 1.0794 | 2.4559 | 1.8491 | | 3.0642 | 3.5081 | 3.5081 | H7 | 2.1786 | 1.0842 | 2.0978 | 3.3483 | 2.4339 | 3.0642 | | 2.5374 | 2.5374 | H8 | 1.0918 | 2.1387 | 3.2093 | 2.0085 | 4.1123 | 3.5081 | 2.5374 | | 1.7684 | H9 | 1.0918 | 2.1387 | 3.2093 | 2.0085 | 4.1123 | 3.5081 | 2.5374 | 1.7684 | |
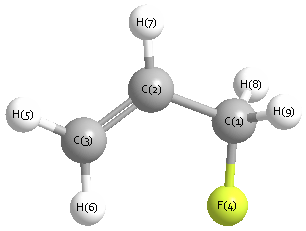 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.871 |
|
C1 |
C2 |
H7 |
114.579 |
| C2 |
C1 |
F4 |
111.742 |
|
C2 |
C1 |
H8 |
110.791 |
| C2 |
C1 |
H9 |
110.791 |
|
C2 |
C3 |
H5 |
120.962 |
| C2 |
C3 |
H6 |
121.172 |
|
C3 |
C2 |
H7 |
120.551 |
| F4 |
C1 |
H8 |
107.599 |
|
F4 |
C1 |
H9 |
107.599 |
| H5 |
C3 |
H6 |
117.866 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.142 |
|
|
|
| 2 |
C |
-0.140 |
|
|
|
| 3 |
C |
-0.207 |
|
|
|
| 4 |
F |
-0.253 |
|
|
|
| 5 |
H |
0.098 |
|
|
|
| 6 |
H |
0.107 |
|
|
|
| 7 |
H |
0.092 |
|
|
|
| 8 |
H |
0.080 |
|
|
|
| 9 |
H |
0.080 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.898 |
1.505 |
0.000 |
1.752 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.454 |
-0.326 |
0.003 |
| y |
-0.326 |
-25.676 |
-0.009 |
| z |
0.003 |
-0.009 |
-25.424 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.097 |
-0.326 |
0.003 |
| y |
-0.326 |
-2.237 |
-0.009 |
| z |
0.003 |
-0.009 |
-1.859 |
|
| Polar |
|---|
| 3z2-r2 | -3.719 |
|---|
| x2-y2 | 4.223 |
|---|
| xy | -0.326 |
|---|
| xz | 0.003 |
|---|
| yz | -0.009 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.184 |
0.294 |
-0.001 |
| y |
0.294 |
5.254 |
0.000 |
| z |
-0.001 |
0.000 |
4.160 |
<r2> (average value of r
2) Å
2
| <r2> |
80.379 |
| (<r2>)1/2 |
8.965 |
Jump to
S1C1
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -217.063268 |
| Energy at 298.15K | |
| HF Energy | -216.821831 |
| Nuclear repulsion energy | 114.312309 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3253 |
3253 |
9.50 |
59.05 |
0.59 |
0.74 |
| 2 |
A |
3182 |
3182 |
5.70 |
112.61 |
0.23 |
0.37 |
| 3 |
A |
3161 |
3161 |
8.01 |
77.82 |
0.12 |
0.22 |
| 4 |
A |
3117 |
3117 |
20.27 |
66.04 |
0.72 |
0.84 |
| 5 |
A |
3063 |
3063 |
31.63 |
130.58 |
0.08 |
0.15 |
| 6 |
A |
1711 |
1711 |
0.49 |
22.98 |
0.09 |
0.16 |
| 7 |
A |
1520 |
1520 |
3.04 |
4.58 |
0.64 |
0.78 |
| 8 |
A |
1475 |
1475 |
16.68 |
9.42 |
0.53 |
0.69 |
| 9 |
A |
1404 |
1404 |
13.02 |
3.49 |
0.33 |
0.50 |
| 10 |
A |
1326 |
1326 |
0.05 |
13.82 |
0.32 |
0.49 |
| 11 |
A |
1279 |
1279 |
4.21 |
9.29 |
0.66 |
0.80 |
| 12 |
A |
1192 |
1192 |
2.88 |
1.20 |
0.68 |
0.81 |
| 13 |
A |
1041 |
1041 |
66.20 |
2.86 |
0.57 |
0.73 |
| 14 |
A |
1035 |
1035 |
81.61 |
3.86 |
0.63 |
0.77 |
| 15 |
A |
996 |
996 |
3.36 |
1.78 |
0.10 |
0.19 |
| 16 |
A |
975 |
975 |
47.80 |
1.35 |
0.64 |
0.78 |
| 17 |
A |
938 |
938 |
3.60 |
3.24 |
0.13 |
0.23 |
| 18 |
A |
665 |
665 |
7.20 |
2.22 |
0.47 |
0.64 |
| 19 |
A |
437 |
437 |
2.17 |
3.50 |
0.41 |
0.58 |
| 20 |
A |
336 |
336 |
7.38 |
1.89 |
0.75 |
0.86 |
| 21 |
A |
115 |
115 |
1.02 |
3.69 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16110.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16110.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.605 |
0.427 |
0.329 |
| C2 |
0.646 |
-0.379 |
0.253 |
| C3 |
1.793 |
0.094 |
-0.222 |
| F4 |
-1.637 |
-0.223 |
-0.354 |
| H5 |
2.691 |
-0.506 |
-0.236 |
| H6 |
1.869 |
1.103 |
-0.607 |
| H7 |
0.583 |
-1.393 |
0.628 |
| H8 |
-0.940 |
0.545 |
1.360 |
| H9 |
-0.471 |
1.409 |
-0.121 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4896 | 2.4829 | 1.3983 | 3.4712 | 2.7297 | 2.1934 | 1.0907 | 1.0892 |
C2 | 1.4896 | | 1.3289 | 2.3675 | 2.1063 | 2.1047 | 1.0831 | 2.1432 | 2.1414 | C3 | 2.4829 | 1.3289 | | 3.4475 | 1.0799 | 1.0819 | 2.0977 | 3.1897 | 2.6203 | F4 | 1.3983 | 2.3675 | 3.4475 | | 4.3386 | 3.7565 | 2.6946 | 2.0047 | 2.0197 | H5 | 3.4712 | 2.1063 | 1.0799 | 4.3386 | | 1.8442 | 2.4448 | 4.1025 | 3.6984 | H6 | 2.7297 | 2.1047 | 1.0819 | 3.7565 | 1.8442 | | 3.0671 | 3.4736 | 2.4092 | H7 | 2.1934 | 1.0831 | 2.0977 | 2.6946 | 2.4448 | 3.0671 | | 2.5712 | 3.0863 | H8 | 1.0907 | 2.1432 | 3.1897 | 2.0047 | 4.1025 | 3.4736 | 2.5712 | | 1.7779 | H9 | 1.0892 | 2.1414 | 2.6203 | 2.0197 | 3.6984 | 2.4092 | 3.0863 | 1.7779 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.404 |
|
C1 |
C2 |
H7 |
116.087 |
| C2 |
C1 |
F4 |
110.092 |
|
C2 |
C1 |
H8 |
111.386 |
| C2 |
C1 |
H9 |
111.326 |
|
C2 |
C3 |
H5 |
121.610 |
| C2 |
C3 |
H6 |
121.286 |
|
C3 |
C2 |
H7 |
120.502 |
| F4 |
C1 |
H8 |
106.649 |
|
F4 |
C1 |
H9 |
107.926 |
| H5 |
C3 |
H6 |
117.103 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.133 |
|
|
|
| 2 |
C |
-0.120 |
|
|
|
| 3 |
C |
-0.187 |
|
|
|
| 4 |
F |
-0.254 |
|
|
|
| 5 |
H |
0.101 |
|
|
|
| 6 |
H |
0.086 |
|
|
|
| 7 |
H |
0.099 |
|
|
|
| 8 |
H |
0.073 |
|
|
|
| 9 |
H |
0.069 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.525 |
0.736 |
0.879 |
1.908 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.989 |
-1.142 |
-1.986 |
| y |
-1.142 |
-22.776 |
-1.002 |
| z |
-1.986 |
-1.002 |
-25.017 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.093 |
-1.142 |
-1.986 |
| y |
-1.142 |
2.727 |
-1.002 |
| z |
-1.986 |
-1.002 |
-0.634 |
|
| Polar |
|---|
| 3z2-r2 | -1.269 |
|---|
| x2-y2 | -3.213 |
|---|
| xy | -1.142 |
|---|
| xz | -1.986 |
|---|
| yz | -1.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.079 |
0.250 |
-0.710 |
| y |
0.250 |
5.096 |
-0.327 |
| z |
-0.710 |
-0.327 |
4.530 |
<r2> (average value of r
2) Å
2
| <r2> |
88.943 |
| (<r2>)1/2 |
9.431 |
