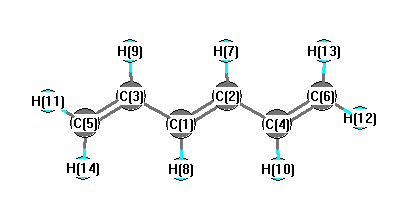
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3369 |
3054 |
0.00 |
|
|
|
| 2 |
Ag |
3298 |
2989 |
0.00 |
|
|
|
| 3 |
Ag |
3290 |
2982 |
0.00 |
|
|
|
| 4 |
Ag |
3282 |
2975 |
0.00 |
|
|
|
| 5 |
Ag |
1869 |
1694 |
0.00 |
|
|
|
| 6 |
Ag |
1774 |
1608 |
0.00 |
|
|
|
| 7 |
Ag |
1551 |
1405 |
0.00 |
|
|
|
| 8 |
Ag |
1442 |
1307 |
0.00 |
|
|
|
| 9 |
Ag |
1421 |
1288 |
0.00 |
|
|
|
| 10 |
Ag |
1300 |
1179 |
0.00 |
|
|
|
| 11 |
Ag |
1012 |
918 |
0.00 |
|
|
|
| 12 |
Ag |
469 |
425 |
0.00 |
|
|
|
| 13 |
Ag |
377 |
342 |
0.00 |
|
|
|
| 14 |
Au |
1150 |
1042 |
66.60 |
|
|
|
| 15 |
Au |
1085 |
983 |
53.00 |
|
|
|
| 16 |
Au |
1056 |
957 |
63.68 |
|
|
|
| 17 |
Au |
777 |
704 |
16.03 |
|
|
|
| 18 |
Au |
271 |
246 |
2.27 |
|
|
|
| 19 |
Au |
100 |
91 |
0.40 |
|
|
|
| 20 |
Bg |
1126 |
1021 |
0.00 |
|
|
|
| 21 |
Bg |
1064 |
964 |
0.00 |
|
|
|
| 22 |
Bg |
1005 |
911 |
0.00 |
|
|
|
| 23 |
Bg |
671 |
608 |
0.00 |
|
|
|
| 24 |
Bg |
217 |
197 |
0.00 |
|
|
|
| 25 |
Bu |
3369 |
3054 |
32.18 |
|
|
|
| 26 |
Bu |
3302 |
2993 |
37.78 |
|
|
|
| 27 |
Bu |
3291 |
2983 |
12.29 |
|
|
|
| 28 |
Bu |
3281 |
2974 |
8.07 |
|
|
|
| 29 |
Bu |
1831 |
1660 |
45.27 |
|
|
|
| 30 |
Bu |
1583 |
1435 |
3.74 |
|
|
|
| 31 |
Bu |
1430 |
1296 |
3.83 |
|
|
|
| 32 |
Bu |
1386 |
1256 |
4.62 |
|
|
|
| 33 |
Bu |
1217 |
1103 |
6.82 |
|
|
|
| 34 |
Bu |
1037 |
940 |
1.24 |
|
|
|
| 35 |
Bu |
581 |
526 |
6.16 |
|
|
|
| 36 |
Bu |
159 |
145 |
3.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27720.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 25125.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.003 |
|
|
|
| 2 |
C |
-0.003 |
|
|
|
| 3 |
C |
-0.105 |
|
|
|
| 4 |
C |
-0.105 |
|
|
|
| 5 |
C |
-0.131 |
|
|
|
| 6 |
C |
-0.131 |
|
|
|
| 7 |
H |
0.045 |
|
|
|
| 8 |
H |
0.045 |
|
|
|
| 9 |
H |
0.063 |
|
|
|
| 10 |
H |
0.063 |
|
|
|
| 11 |
H |
0.075 |
|
|
|
| 12 |
H |
0.075 |
|
|
|
| 13 |
H |
0.056 |
|
|
|
| 14 |
H |
0.056 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.964 |
-0.627 |
0.000 |
| y |
-0.627 |
-34.237 |
0.000 |
| z |
0.000 |
0.000 |
-42.903 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.606 |
-0.627 |
0.000 |
| y |
-0.627 |
4.197 |
0.000 |
| z |
0.000 |
0.000 |
-8.803 |
|
| Polar |
|---|
| 3z2-r2 | -17.605 |
|---|
| x2-y2 | 0.273 |
|---|
| xy | -0.627 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.611 |
3.778 |
0.000 |
| y |
3.778 |
21.845 |
0.000 |
| z |
0.000 |
0.000 |
6.492 |
<r2> (average value of r
2) Å
2
| <r2> |
254.006 |
| (<r2>)1/2 |
15.938 |