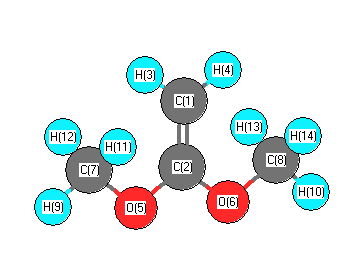
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3356 |
3042 |
1.12 |
|
|
|
| 2 |
A1 |
3281 |
2974 |
36.75 |
|
|
|
| 3 |
A1 |
3173 |
2876 |
8.34 |
|
|
|
| 4 |
A1 |
1803 |
1634 |
393.90 |
|
|
|
| 5 |
A1 |
1636 |
1483 |
5.50 |
|
|
|
| 6 |
A1 |
1617 |
1465 |
11.64 |
|
|
|
| 7 |
A1 |
1545 |
1401 |
42.94 |
|
|
|
| 8 |
A1 |
1344 |
1218 |
5.96 |
|
|
|
| 9 |
A1 |
1296 |
1175 |
0.33 |
|
|
|
| 10 |
A1 |
968 |
877 |
47.69 |
|
|
|
| 11 |
A1 |
539 |
489 |
0.61 |
|
|
|
| 12 |
A1 |
284 |
257 |
5.96 |
|
|
|
| 13 |
A2 |
3233 |
2931 |
0.00 |
|
|
|
| 14 |
A2 |
1620 |
1469 |
0.00 |
|
|
|
| 15 |
A2 |
1287 |
1167 |
0.00 |
|
|
|
| 16 |
A2 |
744 |
675 |
0.00 |
|
|
|
| 17 |
A2 |
252 |
228 |
0.00 |
|
|
|
| 18 |
A2 |
149 |
135 |
0.00 |
|
|
|
| 19 |
B1 |
3234 |
2931 |
80.45 |
|
|
|
| 20 |
B1 |
1620 |
1468 |
16.09 |
|
|
|
| 21 |
B1 |
1289 |
1169 |
5.64 |
|
|
|
| 22 |
B1 |
850 |
771 |
105.15 |
|
|
|
| 23 |
B1 |
764 |
693 |
2.59 |
|
|
|
| 24 |
B1 |
266 |
241 |
0.37 |
|
|
|
| 25 |
B1 |
104 |
94 |
10.28 |
|
|
|
| 26 |
B2 |
3439 |
3117 |
7.06 |
|
|
|
| 27 |
B2 |
3280 |
2973 |
26.29 |
|
|
|
| 28 |
B2 |
3173 |
2876 |
69.12 |
|
|
|
| 29 |
B2 |
1631 |
1478 |
32.37 |
|
|
|
| 30 |
B2 |
1627 |
1474 |
43.55 |
|
|
|
| 31 |
B2 |
1483 |
1344 |
597.75 |
|
|
|
| 32 |
B2 |
1333 |
1208 |
8.10 |
|
|
|
| 33 |
B2 |
1167 |
1058 |
156.48 |
|
|
|
| 34 |
B2 |
1028 |
932 |
0.02 |
|
|
|
| 35 |
B2 |
607 |
550 |
0.00 |
|
|
|
| 36 |
B2 |
401 |
364 |
7.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27711.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 25117.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.323 |
|
|
|
| 2 |
C |
0.477 |
|
|
|
| 3 |
H |
0.051 |
|
|
|
| 4 |
H |
0.051 |
|
|
|
| 5 |
O |
-0.359 |
|
|
|
| 6 |
O |
-0.359 |
|
|
|
| 7 |
C |
-0.000 |
|
|
|
| 8 |
C |
-0.000 |
|
|
|
| 9 |
H |
0.088 |
|
|
|
| 10 |
H |
0.088 |
|
|
|
| 11 |
H |
0.071 |
|
|
|
| 12 |
H |
0.071 |
|
|
|
| 13 |
H |
0.071 |
|
|
|
| 14 |
H |
0.071 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.052 |
1.052 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.586 |
0.000 |
0.000 |
| y |
0.000 |
-26.655 |
0.000 |
| z |
0.000 |
0.000 |
-39.632 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.442 |
0.000 |
0.000 |
| y |
0.000 |
12.454 |
0.000 |
| z |
0.000 |
0.000 |
-7.012 |
|
| Polar |
|---|
| 3z2-r2 | -14.024 |
|---|
| x2-y2 | -11.931 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.321 |
0.000 |
0.000 |
| y |
0.000 |
9.180 |
0.000 |
| z |
0.000 |
0.000 |
8.300 |
<r2> (average value of r
2) Å
2
| <r2> |
184.744 |
| (<r2>)1/2 |
13.592 |