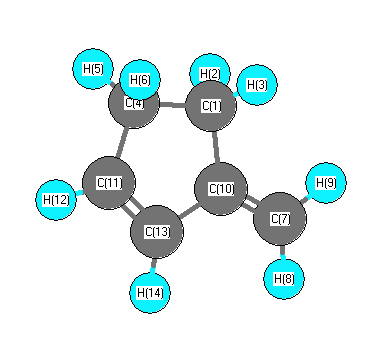
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3358 |
3044 |
21.08 |
|
|
|
| 2 |
A |
3353 |
3040 |
12.69 |
|
|
|
| 3 |
A |
3327 |
3016 |
5.30 |
|
|
|
| 4 |
A |
3281 |
2974 |
7.49 |
|
|
|
| 5 |
A |
3227 |
2925 |
31.88 |
|
|
|
| 6 |
A |
3195 |
2896 |
36.67 |
|
|
|
| 7 |
A |
3182 |
2884 |
17.19 |
|
|
|
| 8 |
A |
3164 |
2868 |
52.93 |
|
|
|
| 9 |
A |
1851 |
1678 |
57.51 |
|
|
|
| 10 |
A |
1787 |
1619 |
2.74 |
|
|
|
| 11 |
A |
1624 |
1472 |
2.37 |
|
|
|
| 12 |
A |
1596 |
1446 |
1.84 |
|
|
|
| 13 |
A |
1572 |
1425 |
0.49 |
|
|
|
| 14 |
A |
1486 |
1347 |
0.41 |
|
|
|
| 15 |
A |
1457 |
1321 |
1.35 |
|
|
|
| 16 |
A |
1398 |
1268 |
3.79 |
|
|
|
| 17 |
A |
1376 |
1247 |
1.57 |
|
|
|
| 18 |
A |
1355 |
1228 |
0.15 |
|
|
|
| 19 |
A |
1271 |
1152 |
2.56 |
|
|
|
| 20 |
A |
1213 |
1099 |
2.64 |
|
|
|
| 21 |
A |
1139 |
1032 |
4.68 |
|
|
|
| 22 |
A |
1094 |
992 |
5.74 |
|
|
|
| 23 |
A |
1080 |
979 |
2.95 |
|
|
|
| 24 |
A |
1021 |
926 |
2.45 |
|
|
|
| 25 |
A |
1014 |
919 |
50.36 |
|
|
|
| 26 |
A |
968 |
878 |
6.87 |
|
|
|
| 27 |
A |
919 |
833 |
19.86 |
|
|
|
| 28 |
A |
902 |
818 |
0.06 |
|
|
|
| 29 |
A |
862 |
781 |
4.88 |
|
|
|
| 30 |
A |
828 |
751 |
2.13 |
|
|
|
| 31 |
A |
745 |
675 |
13.07 |
|
|
|
| 32 |
A |
670 |
607 |
0.74 |
|
|
|
| 33 |
A |
560 |
508 |
16.24 |
|
|
|
| 34 |
A |
386 |
350 |
1.18 |
|
|
|
| 35 |
A |
317 |
287 |
1.79 |
|
|
|
| 36 |
A |
22i |
20i |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28278.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 25631.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.061 |
|
|
|
| 2 |
H |
0.057 |
|
|
|
| 3 |
H |
0.057 |
|
|
|
| 4 |
C |
0.001 |
|
|
|
| 5 |
H |
0.047 |
|
|
|
| 6 |
H |
0.047 |
|
|
|
| 7 |
C |
-0.160 |
|
|
|
| 8 |
H |
0.056 |
|
|
|
| 9 |
H |
0.059 |
|
|
|
| 10 |
C |
-0.022 |
|
|
|
| 11 |
C |
-0.109 |
|
|
|
| 12 |
H |
0.084 |
|
|
|
| 13 |
C |
-0.141 |
|
|
|
| 14 |
H |
0.086 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.891 |
-0.221 |
-0.000 |
0.918 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.946 |
-0.059 |
0.001 |
| y |
-0.059 |
-34.945 |
0.001 |
| z |
0.001 |
0.001 |
-39.876 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.465 |
-0.059 |
0.001 |
| y |
-0.059 |
2.966 |
0.001 |
| z |
0.001 |
0.001 |
-4.431 |
|
| Polar |
|---|
| 3z2-r2 | -8.861 |
|---|
| x2-y2 | -1.001 |
|---|
| xy | -0.059 |
|---|
| xz | 0.001 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.764 |
-0.079 |
0.003 |
| y |
-0.079 |
9.087 |
0.001 |
| z |
0.003 |
0.001 |
6.810 |
<r2> (average value of r
2) Å
2
| <r2> |
151.805 |
| (<r2>)1/2 |
12.321 |