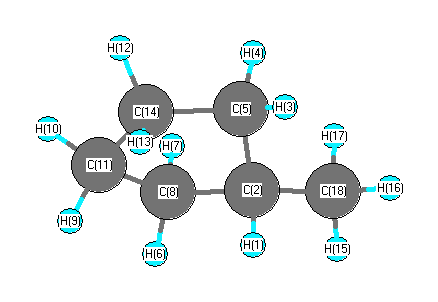
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3226 |
2924 |
157.20 |
|
|
|
| 2 |
A |
3207 |
2907 |
39.01 |
|
|
|
| 3 |
A |
3205 |
2905 |
78.84 |
|
|
|
| 4 |
A |
3203 |
2903 |
2.56 |
|
|
|
| 5 |
A |
3199 |
2900 |
26.29 |
|
|
|
| 6 |
A |
3190 |
2892 |
73.85 |
|
|
|
| 7 |
A |
3182 |
2884 |
47.58 |
|
|
|
| 8 |
A |
3166 |
2869 |
15.11 |
|
|
|
| 9 |
A |
3155 |
2859 |
98.55 |
|
|
|
| 10 |
A |
3146 |
2852 |
13.15 |
|
|
|
| 11 |
A |
3144 |
2850 |
20.98 |
|
|
|
| 12 |
A |
3117 |
2826 |
13.35 |
|
|
|
| 13 |
A |
1645 |
1491 |
1.01 |
|
|
|
| 14 |
A |
1620 |
1468 |
7.02 |
|
|
|
| 15 |
A |
1619 |
1468 |
1.52 |
|
|
|
| 16 |
A |
1614 |
1463 |
3.03 |
|
|
|
| 17 |
A |
1613 |
1462 |
2.98 |
|
|
|
| 18 |
A |
1606 |
1455 |
0.00 |
|
|
|
| 19 |
A |
1538 |
1394 |
3.28 |
|
|
|
| 20 |
A |
1518 |
1376 |
0.96 |
|
|
|
| 21 |
A |
1475 |
1337 |
0.97 |
|
|
|
| 22 |
A |
1460 |
1323 |
0.03 |
|
|
|
| 23 |
A |
1450 |
1314 |
0.20 |
|
|
|
| 24 |
A |
1432 |
1298 |
1.32 |
|
|
|
| 25 |
A |
1417 |
1284 |
0.00 |
|
|
|
| 26 |
A |
1370 |
1242 |
0.25 |
|
|
|
| 27 |
A |
1354 |
1227 |
0.46 |
|
|
|
| 28 |
A |
1323 |
1199 |
0.18 |
|
|
|
| 29 |
A |
1310 |
1188 |
0.43 |
|
|
|
| 30 |
A |
1255 |
1138 |
0.59 |
|
|
|
| 31 |
A |
1174 |
1064 |
0.19 |
|
|
|
| 32 |
A |
1121 |
1017 |
0.15 |
|
|
|
| 33 |
A |
1087 |
985 |
0.39 |
|
|
|
| 34 |
A |
1065 |
965 |
1.78 |
|
|
|
| 35 |
A |
1053 |
954 |
0.05 |
|
|
|
| 36 |
A |
999 |
906 |
0.01 |
|
|
|
| 37 |
A |
978 |
886 |
0.53 |
|
|
|
| 38 |
A |
944 |
855 |
0.84 |
|
|
|
| 39 |
A |
906 |
821 |
0.48 |
|
|
|
| 40 |
A |
877 |
795 |
0.09 |
|
|
|
| 41 |
A |
810 |
734 |
0.39 |
|
|
|
| 42 |
A |
674 |
611 |
0.10 |
|
|
|
| 43 |
A |
563 |
510 |
0.29 |
|
|
|
| 44 |
A |
457 |
414 |
0.12 |
|
|
|
| 45 |
A |
323 |
293 |
0.01 |
|
|
|
| 46 |
A |
252 |
229 |
0.00 |
|
|
|
| 47 |
A |
189 |
171 |
0.00 |
|
|
|
| 48 |
A |
37 |
33 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 39132.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 35469.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.013 |
|
|
|
| 2 |
C |
0.119 |
|
|
|
| 3 |
H |
0.050 |
|
|
|
| 4 |
H |
0.037 |
|
|
|
| 5 |
C |
-0.128 |
|
|
|
| 6 |
H |
0.050 |
|
|
|
| 7 |
H |
0.038 |
|
|
|
| 8 |
C |
-0.128 |
|
|
|
| 9 |
H |
0.047 |
|
|
|
| 10 |
H |
0.052 |
|
|
|
| 11 |
C |
-0.094 |
|
|
|
| 12 |
H |
0.052 |
|
|
|
| 13 |
H |
0.047 |
|
|
|
| 14 |
C |
-0.094 |
|
|
|
| 15 |
H |
0.060 |
|
|
|
| 16 |
H |
0.060 |
|
|
|
| 17 |
H |
0.044 |
|
|
|
| 18 |
C |
-0.223 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.022 |
-0.000 |
-0.086 |
0.089 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.392 |
-0.000 |
-0.277 |
| y |
-0.000 |
-40.742 |
-0.001 |
| z |
-0.277 |
-0.001 |
-39.465 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.289 |
-0.000 |
-0.277 |
| y |
-0.000 |
-0.313 |
-0.001 |
| z |
-0.277 |
-0.001 |
1.602 |
|
| Polar |
|---|
| 3z2-r2 | 3.204 |
|---|
| x2-y2 | -0.650 |
|---|
| xy | -0.000 |
|---|
| xz | -0.277 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.789 |
0.000 |
0.036 |
| y |
0.000 |
9.806 |
0.000 |
| z |
0.036 |
0.000 |
8.756 |
<r2> (average value of r
2) Å
2
| <r2> |
178.344 |
| (<r2>)1/2 |
13.355 |