Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -340.678142 |
| Energy at 298.15K | |
| HF Energy | -340.678142 |
| Nuclear repulsion energy | 249.972884 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3232 |
2929 |
56.53 |
|
|
|
| 2 |
A1 |
2056 |
1864 |
894.62 |
|
|
|
| 3 |
A1 |
1675 |
1518 |
1.64 |
|
|
|
| 4 |
A1 |
1530 |
1386 |
0.01 |
|
|
|
| 5 |
A1 |
1264 |
1145 |
308.73 |
|
|
|
| 6 |
A1 |
1056 |
957 |
21.60 |
|
|
|
| 7 |
A1 |
991 |
898 |
0.51 |
|
|
|
| 8 |
A1 |
791 |
717 |
8.18 |
|
|
|
| 9 |
A2 |
3258 |
2953 |
0.00 |
|
|
|
| 10 |
A2 |
1358 |
1230 |
0.00 |
|
|
|
| 11 |
A2 |
1266 |
1148 |
0.00 |
|
|
|
| 12 |
A2 |
92i |
84i |
0.00 |
|
|
|
| 13 |
B1 |
3283 |
2976 |
44.90 |
|
|
|
| 14 |
B1 |
1353 |
1227 |
4.19 |
|
|
|
| 15 |
B1 |
920 |
834 |
4.84 |
|
|
|
| 16 |
B1 |
881 |
799 |
40.29 |
|
|
|
| 17 |
B1 |
194 |
175 |
1.28 |
|
|
|
| 18 |
B2 |
3221 |
2920 |
25.02 |
|
|
|
| 19 |
B2 |
1661 |
1506 |
12.39 |
|
|
|
| 20 |
B2 |
1564 |
1417 |
84.02 |
|
|
|
| 21 |
B2 |
1308 |
1185 |
267.91 |
|
|
|
| 22 |
B2 |
1194 |
1082 |
15.41 |
|
|
|
| 23 |
B2 |
826 |
749 |
2.76 |
|
|
|
| 24 |
B2 |
579 |
525 |
1.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17683.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 16028.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.848 |
| O2 |
0.000 |
0.000 |
2.007 |
| O3 |
0.000 |
1.089 |
0.079 |
| O4 |
0.000 |
-1.089 |
0.079 |
| C5 |
0.000 |
0.768 |
-1.281 |
| C6 |
0.000 |
-0.768 |
-1.281 |
| H7 |
-0.878 |
1.180 |
-1.758 |
| H8 |
0.878 |
1.180 |
-1.758 |
| H9 |
0.878 |
-1.180 |
-1.758 |
| H10 |
-0.878 |
-1.180 |
-1.758 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1590 | 1.3333 | 1.3333 | 2.2635 | 2.2635 | 2.9924 | 2.9924 | 2.9924 | 2.9924 |
O2 | 1.1590 | | 2.2146 | 2.2146 | 3.3768 | 3.3768 | 4.0421 | 4.0421 | 4.0421 | 4.0421 | O3 | 1.3333 | 2.2146 | | 2.1778 | 1.3973 | 2.3015 | 2.0379 | 2.0379 | 3.0481 | 3.0481 | O4 | 1.3333 | 2.2146 | 2.1778 | | 2.3015 | 1.3973 | 3.0481 | 3.0481 | 2.0379 | 2.0379 | C5 | 2.2635 | 3.3768 | 1.3973 | 2.3015 | | 1.5357 | 1.0809 | 1.0809 | 2.1889 | 2.1889 | C6 | 2.2635 | 3.3768 | 2.3015 | 1.3973 | 1.5357 | | 2.1889 | 2.1889 | 1.0809 | 1.0809 | H7 | 2.9924 | 4.0421 | 2.0379 | 3.0481 | 1.0809 | 2.1889 | | 1.7567 | 2.9414 | 2.3592 | H8 | 2.9924 | 4.0421 | 2.0379 | 3.0481 | 1.0809 | 2.1889 | 1.7567 | | 2.3592 | 2.9414 | H9 | 2.9924 | 4.0421 | 3.0481 | 2.0379 | 2.1889 | 1.0809 | 2.9414 | 2.3592 | | 1.7567 | H10 | 2.9924 | 4.0421 | 3.0481 | 2.0379 | 2.1889 | 1.0809 | 2.3592 | 2.9414 | 1.7567 | |
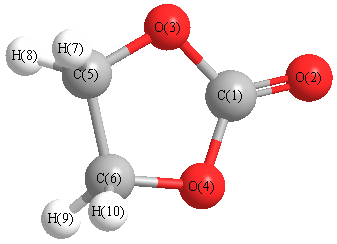 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
111.962 |
|
C1 |
O4 |
C6 |
111.962 |
| O2 |
C1 |
O3 |
125.245 |
|
O2 |
C1 |
O4 |
125.245 |
| O3 |
C1 |
O4 |
109.510 |
|
O3 |
C5 |
C6 |
103.283 |
| O3 |
C5 |
H7 |
109.982 |
|
O3 |
C5 |
H8 |
109.982 |
| O4 |
C6 |
C5 |
103.283 |
|
O4 |
C6 |
H9 |
109.982 |
| O4 |
C6 |
H10 |
109.982 |
|
C5 |
C6 |
H9 |
112.392 |
| C5 |
C6 |
H10 |
112.392 |
|
C6 |
C5 |
H7 |
112.392 |
| C6 |
C5 |
H8 |
112.392 |
|
H7 |
C5 |
H8 |
108.706 |
| H9 |
C6 |
H10 |
108.706 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.617 |
|
|
|
| 2 |
O |
-0.410 |
|
|
|
| 3 |
O |
-0.350 |
|
|
|
| 4 |
O |
-0.350 |
|
|
|
| 5 |
C |
0.122 |
|
|
|
| 6 |
C |
0.122 |
|
|
|
| 7 |
H |
0.062 |
|
|
|
| 8 |
H |
0.062 |
|
|
|
| 9 |
H |
0.062 |
|
|
|
| 10 |
H |
0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-5.893 |
5.893 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.776 |
0.000 |
0.000 |
| y |
0.000 |
-35.977 |
0.000 |
| z |
0.000 |
0.000 |
-36.894 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.659 |
0.000 |
0.000 |
| y |
0.000 |
-1.642 |
0.000 |
| z |
0.000 |
0.000 |
-3.017 |
|
| Polar |
|---|
| 3z2-r2 | -6.034 |
|---|
| x2-y2 | 4.201 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.497 |
0.000 |
0.000 |
| y |
0.000 |
5.331 |
0.000 |
| z |
0.000 |
0.000 |
7.166 |
<r2> (average value of r
2) Å
2
| <r2> |
125.232 |
| (<r2>)1/2 |
11.191 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -340.678332 |
| Energy at 298.15K | -340.685062 |
| HF Energy | -340.678332 |
| Nuclear repulsion energy | 250.257777 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3270 |
2964 |
15.28 |
|
|
|
| 2 |
A |
3219 |
2918 |
36.84 |
|
|
|
| 3 |
A |
2059 |
1866 |
886.41 |
|
|
|
| 4 |
A |
1669 |
1513 |
1.74 |
|
|
|
| 5 |
A |
1531 |
1388 |
0.65 |
|
|
|
| 6 |
A |
1360 |
1232 |
13.96 |
|
|
|
| 7 |
A |
1271 |
1152 |
62.48 |
|
|
|
| 8 |
A |
1250 |
1133 |
233.15 |
|
|
|
| 9 |
A |
1049 |
950 |
18.99 |
|
|
|
| 10 |
A |
995 |
902 |
0.11 |
|
|
|
| 11 |
A |
789 |
715 |
8.19 |
|
|
|
| 12 |
A |
129 |
117 |
0.19 |
|
|
|
| 13 |
B |
3288 |
2980 |
36.49 |
|
|
|
| 14 |
B |
3217 |
2916 |
34.60 |
|
|
|
| 15 |
B |
1657 |
1502 |
12.73 |
|
|
|
| 16 |
B |
1558 |
1412 |
79.25 |
|
|
|
| 17 |
B |
1355 |
1228 |
21.48 |
|
|
|
| 18 |
B |
1299 |
1178 |
251.06 |
|
|
|
| 19 |
B |
1188 |
1077 |
14.49 |
|
|
|
| 20 |
B |
953 |
864 |
3.30 |
|
|
|
| 21 |
B |
885 |
802 |
42.17 |
|
|
|
| 22 |
B |
781 |
708 |
2.63 |
|
|
|
| 23 |
B |
579 |
525 |
1.30 |
|
|
|
| 24 |
B |
196 |
178 |
1.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17773.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 16109.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.839 |
| O2 |
0.000 |
0.000 |
1.997 |
| O3 |
0.000 |
1.093 |
0.068 |
| O4 |
0.000 |
-1.093 |
0.068 |
| C5 |
0.228 |
0.721 |
-1.271 |
| C6 |
-0.228 |
-0.721 |
-1.271 |
| H7 |
-0.341 |
1.359 |
-1.928 |
| H8 |
1.283 |
0.814 |
-1.494 |
| H9 |
0.341 |
-1.359 |
-1.928 |
| H10 |
-1.283 |
-0.814 |
-1.494 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1577 | 1.3375 | 1.3375 | 2.2413 | 2.2413 | 3.1012 | 2.7844 | 3.1012 | 2.7844 |
O2 | 1.1577 | | 2.2171 | 2.2171 | 3.3539 | 3.3539 | 4.1670 | 3.8075 | 4.1670 | 3.8075 | O3 | 1.3375 | 2.2171 | | 2.1853 | 1.4074 | 2.2657 | 2.0416 | 2.0405 | 3.1793 | 2.7785 | O4 | 1.3375 | 2.2171 | 2.1853 | | 2.2657 | 1.4074 | 3.1793 | 2.7785 | 2.0416 | 2.0405 | C5 | 2.2413 | 3.3539 | 1.4074 | 2.2657 | | 1.5129 | 1.0779 | 1.0824 | 2.1846 | 2.1653 | C6 | 2.2413 | 3.3539 | 2.2657 | 1.4074 | 1.5129 | | 2.1846 | 2.1653 | 1.0779 | 1.0824 | H7 | 3.1012 | 4.1670 | 2.0416 | 3.1793 | 1.0779 | 2.1846 | | 1.7668 | 2.8024 | 2.4074 | H8 | 2.7844 | 3.8075 | 2.0405 | 2.7785 | 1.0824 | 2.1653 | 1.7668 | | 2.4074 | 3.0382 | H9 | 3.1012 | 4.1670 | 3.1793 | 2.0416 | 2.1846 | 1.0779 | 2.8024 | 2.4074 | | 1.7668 | H10 | 2.7844 | 3.8075 | 2.7785 | 2.0405 | 2.1653 | 1.0824 | 2.4074 | 3.0382 | 1.7668 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
109.445 |
|
C1 |
O4 |
C6 |
109.445 |
| O2 |
C1 |
O3 |
125.223 |
|
O2 |
C1 |
O4 |
125.223 |
| O3 |
C1 |
O4 |
109.554 |
|
O3 |
C5 |
C6 |
101.703 |
| O3 |
C5 |
H7 |
109.754 |
|
O3 |
C5 |
H8 |
109.379 |
| O4 |
C6 |
C5 |
101.703 |
|
O4 |
C6 |
H9 |
109.754 |
| O4 |
C6 |
H10 |
109.379 |
|
C5 |
C6 |
H9 |
113.910 |
| C5 |
C6 |
H10 |
112.022 |
|
C6 |
C5 |
H7 |
113.910 |
| C6 |
C5 |
H8 |
112.022 |
|
H7 |
C5 |
H8 |
109.744 |
| H9 |
C6 |
H10 |
109.744 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.618 |
|
|
|
| 2 |
O |
-0.408 |
|
|
|
| 3 |
O |
-0.353 |
|
|
|
| 4 |
O |
-0.353 |
|
|
|
| 5 |
C |
0.122 |
|
|
|
| 6 |
C |
0.122 |
|
|
|
| 7 |
H |
0.066 |
|
|
|
| 8 |
H |
0.060 |
|
|
|
| 9 |
H |
0.066 |
|
|
|
| 10 |
H |
0.060 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-5.867 |
5.867 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.806 |
0.226 |
0.000 |
| y |
0.226 |
-36.000 |
0.000 |
| z |
0.000 |
0.000 |
-36.786 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.587 |
0.226 |
0.000 |
| y |
0.226 |
-1.703 |
0.000 |
| z |
0.000 |
0.000 |
-2.883 |
|
| Polar |
|---|
| 3z2-r2 | -5.767 |
|---|
| x2-y2 | 4.193 |
|---|
| xy | 0.226 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.537 |
0.096 |
0.000 |
| y |
0.096 |
5.303 |
0.000 |
| z |
0.000 |
0.000 |
7.158 |
<r2> (average value of r
2) Å
2
| <r2> |
124.718 |
| (<r2>)1/2 |
11.168 |
