Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -341.491947 |
| Energy at 298.15K | -341.497147 |
| HF Energy | -341.491947 |
| Nuclear repulsion energy | 274.868574 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3432 |
3111 |
0.72 |
125.81 |
0.13 |
0.23 |
| 2 |
A' |
3409 |
3090 |
0.26 |
48.01 |
0.34 |
0.51 |
| 3 |
A' |
3397 |
3079 |
2.61 |
72.38 |
0.66 |
0.80 |
| 4 |
A' |
3124 |
2831 |
82.06 |
110.58 |
0.30 |
0.46 |
| 5 |
A' |
1974 |
1789 |
460.97 |
170.88 |
0.31 |
0.47 |
| 6 |
A' |
1763 |
1598 |
7.86 |
36.77 |
0.41 |
0.58 |
| 7 |
A' |
1648 |
1494 |
55.91 |
149.30 |
0.22 |
0.36 |
| 8 |
A' |
1552 |
1407 |
48.10 |
69.37 |
0.34 |
0.51 |
| 9 |
A' |
1522 |
1379 |
2.04 |
18.78 |
0.09 |
0.17 |
| 10 |
A' |
1393 |
1262 |
50.62 |
4.21 |
0.71 |
0.83 |
| 11 |
A' |
1340 |
1214 |
18.15 |
22.60 |
0.36 |
0.53 |
| 12 |
A' |
1282 |
1162 |
21.28 |
7.64 |
0.18 |
0.31 |
| 13 |
A' |
1202 |
1089 |
6.54 |
15.69 |
0.44 |
0.62 |
| 14 |
A' |
1092 |
990 |
37.65 |
16.91 |
0.49 |
0.66 |
| 15 |
A' |
1031 |
935 |
27.43 |
3.96 |
0.06 |
0.12 |
| 16 |
A' |
972 |
881 |
14.89 |
9.05 |
0.69 |
0.82 |
| 17 |
A' |
821 |
745 |
85.03 |
2.25 |
0.70 |
0.82 |
| 18 |
A' |
534 |
484 |
1.39 |
5.02 |
0.22 |
0.36 |
| 19 |
A' |
221 |
201 |
8.33 |
1.03 |
0.64 |
0.78 |
| 20 |
A" |
1132 |
1026 |
0.05 |
3.93 |
0.75 |
0.86 |
| 21 |
A" |
1030 |
934 |
1.57 |
3.47 |
0.75 |
0.86 |
| 22 |
A" |
976 |
884 |
5.82 |
1.38 |
0.75 |
0.86 |
| 23 |
A" |
874 |
792 |
74.19 |
0.27 |
0.75 |
0.86 |
| 24 |
A" |
710 |
644 |
1.02 |
2.18 |
0.75 |
0.86 |
| 25 |
A" |
656 |
594 |
11.09 |
0.64 |
0.75 |
0.86 |
| 26 |
A" |
311 |
282 |
18.88 |
0.51 |
0.75 |
0.86 |
| 27 |
A" |
142 |
129 |
3.05 |
0.83 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 18769.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 17012.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.494 |
-0.893 |
0.000 |
| C2 |
-0.540 |
-1.727 |
0.000 |
| C3 |
-1.705 |
-1.061 |
0.000 |
| C4 |
0.000 |
0.354 |
0.000 |
| C5 |
-1.344 |
0.323 |
0.000 |
| C6 |
0.910 |
1.499 |
0.000 |
| O7 |
2.090 |
1.430 |
0.000 |
| H8 |
-0.311 |
-2.769 |
0.000 |
| H9 |
-2.684 |
-1.485 |
0.000 |
| H10 |
-1.996 |
1.169 |
0.000 |
| H11 |
0.392 |
2.461 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3284 | 2.2056 | 1.3415 | 2.2037 | 2.4277 | 2.8188 | 2.0415 | 3.2332 | 3.2332 | 3.3558 |
C2 | 1.3284 | | 1.3416 | 2.1498 | 2.2013 | 3.5364 | 4.1092 | 1.0671 | 2.1578 | 3.2412 | 4.2903 | C3 | 2.2056 | 1.3416 | | 2.2161 | 1.4303 | 3.6597 | 4.5401 | 2.2042 | 1.0672 | 2.2493 | 4.0994 | C4 | 1.3415 | 2.1498 | 2.2161 | | 1.3443 | 1.4623 | 2.3510 | 3.1386 | 3.2542 | 2.1560 | 2.1431 | C5 | 2.2037 | 2.2013 | 1.4303 | 1.3443 | | 2.5425 | 3.6084 | 3.2595 | 2.2506 | 1.0685 | 2.7543 | C6 | 2.4277 | 3.5364 | 3.6597 | 1.4623 | 2.5425 | | 1.1819 | 4.4390 | 4.6717 | 2.9249 | 1.0932 | O7 | 2.8188 | 4.1092 | 4.5401 | 2.3510 | 3.6084 | 1.1819 | | 4.8376 | 5.5944 | 4.0946 | 1.9866 | H8 | 2.0415 | 1.0671 | 2.2042 | 3.1386 | 3.2595 | 4.4390 | 4.8376 | | 2.6980 | 4.2832 | 5.2772 | H9 | 3.2332 | 2.1578 | 1.0672 | 3.2542 | 2.2506 | 4.6717 | 5.5944 | 2.6980 | | 2.7421 | 5.0038 | H10 | 3.2332 | 3.2412 | 2.2493 | 2.1560 | 1.0685 | 2.9249 | 4.0946 | 4.2832 | 2.7421 | | 2.7151 | H11 | 3.3558 | 4.2903 | 4.0994 | 2.1431 | 2.7543 | 1.0932 | 1.9866 | 5.2772 | 5.0038 | 2.7151 | |
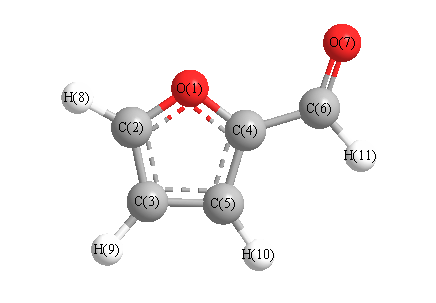 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.396 |
|
O1 |
C2 |
H8 |
116.493 |
| O1 |
C4 |
C5 |
110.267 |
|
O1 |
C4 |
C6 |
119.894 |
| C2 |
O1 |
C4 |
107.259 |
|
C2 |
C3 |
C5 |
105.110 |
| C2 |
C3 |
H9 |
126.860 |
|
C3 |
C2 |
H8 |
132.111 |
| C3 |
C5 |
C4 |
105.969 |
|
C3 |
C5 |
H10 |
127.762 |
| C4 |
C5 |
H10 |
126.269 |
|
C4 |
C6 |
O7 |
125.182 |
| C4 |
C6 |
H11 |
113.202 |
|
C5 |
C3 |
H9 |
128.030 |
| C5 |
C4 |
C6 |
129.839 |
|
O7 |
C6 |
H11 |
121.615 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.334 |
|
|
|
| 2 |
C |
0.144 |
|
|
|
| 3 |
C |
-0.159 |
|
|
|
| 4 |
C |
0.197 |
|
|
|
| 5 |
C |
-0.091 |
|
|
|
| 6 |
C |
0.230 |
|
|
|
| 7 |
O |
-0.387 |
|
|
|
| 8 |
H |
0.119 |
|
|
|
| 9 |
H |
0.114 |
|
|
|
| 10 |
H |
0.120 |
|
|
|
| 11 |
H |
0.049 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.201 |
-0.799 |
0.000 |
4.276 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.948 |
-3.246 |
0.000 |
| y |
-3.246 |
-33.388 |
0.000 |
| z |
0.000 |
0.000 |
-41.549 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.480 |
-3.246 |
0.000 |
| y |
-3.246 |
9.861 |
0.000 |
| z |
0.000 |
0.000 |
-2.380 |
|
| Polar |
|---|
| 3z2-r2 | -4.761 |
|---|
| x2-y2 | -11.560 |
|---|
| xy | -3.246 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.797 |
1.482 |
0.000 |
| y |
1.482 |
10.020 |
0.000 |
| z |
0.000 |
0.000 |
5.253 |
<r2> (average value of r
2) Å
2
| <r2> |
187.852 |
| (<r2>)1/2 |
13.706 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -341.492536 |
| Energy at 298.15K | -341.497687 |
| HF Energy | -341.492536 |
| Nuclear repulsion energy | 273.957841 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3432 |
3111 |
0.86 |
132.19 |
0.13 |
0.23 |
| 2 |
A' |
3418 |
3099 |
0.09 |
30.57 |
0.29 |
0.45 |
| 3 |
A' |
3402 |
3083 |
1.72 |
73.61 |
0.71 |
0.83 |
| 4 |
A' |
3136 |
2843 |
79.77 |
123.29 |
0.29 |
0.45 |
| 5 |
A' |
1969 |
1785 |
379.76 |
110.44 |
0.34 |
0.51 |
| 6 |
A' |
1772 |
1606 |
55.99 |
61.10 |
0.38 |
0.55 |
| 7 |
A' |
1636 |
1483 |
117.92 |
259.15 |
0.23 |
0.37 |
| 8 |
A' |
1566 |
1419 |
13.72 |
21.00 |
0.55 |
0.71 |
| 9 |
A' |
1512 |
1371 |
13.57 |
30.72 |
0.36 |
0.52 |
| 10 |
A' |
1365 |
1237 |
22.55 |
6.54 |
0.57 |
0.72 |
| 11 |
A' |
1325 |
1201 |
18.61 |
6.09 |
0.19 |
0.31 |
| 12 |
A' |
1266 |
1147 |
30.22 |
13.14 |
0.10 |
0.18 |
| 13 |
A' |
1202 |
1089 |
23.92 |
19.07 |
0.56 |
0.72 |
| 14 |
A' |
1089 |
987 |
30.72 |
18.98 |
0.44 |
0.61 |
| 15 |
A' |
1046 |
948 |
17.53 |
5.00 |
0.22 |
0.36 |
| 16 |
A' |
971 |
880 |
10.75 |
7.44 |
0.74 |
0.85 |
| 17 |
A' |
811 |
735 |
110.18 |
3.39 |
0.73 |
0.84 |
| 18 |
A' |
528 |
478 |
1.32 |
3.44 |
0.25 |
0.40 |
| 19 |
A' |
226 |
204 |
7.60 |
0.47 |
0.38 |
0.55 |
| 20 |
A" |
1139 |
1032 |
0.18 |
5.47 |
0.75 |
0.86 |
| 21 |
A" |
1032 |
935 |
2.13 |
3.61 |
0.75 |
0.86 |
| 22 |
A" |
986 |
893 |
5.21 |
0.83 |
0.75 |
0.86 |
| 23 |
A" |
876 |
794 |
71.48 |
0.40 |
0.75 |
0.86 |
| 24 |
A" |
698 |
633 |
1.36 |
1.45 |
0.75 |
0.86 |
| 25 |
A" |
656 |
595 |
13.51 |
0.67 |
0.75 |
0.86 |
| 26 |
A" |
258 |
234 |
15.81 |
1.43 |
0.75 |
0.86 |
| 27 |
A" |
155 |
141 |
7.42 |
0.50 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 18735.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 16982.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.218 |
-0.305 |
0.000 |
| C2 |
1.027 |
-1.620 |
0.000 |
| C3 |
-0.281 |
-1.920 |
0.000 |
| C4 |
0.000 |
0.275 |
0.000 |
| C5 |
-0.961 |
-0.662 |
0.000 |
| C6 |
-0.040 |
1.739 |
0.000 |
| O7 |
-1.042 |
2.369 |
0.000 |
| H8 |
1.903 |
-2.230 |
0.000 |
| H9 |
-0.711 |
-2.897 |
0.000 |
| H10 |
-2.012 |
-0.479 |
0.000 |
| H11 |
0.941 |
2.219 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3293 | 2.2034 | 1.3485 | 2.2079 | 2.3998 | 3.5009 | 2.0435 | 3.2310 | 3.2348 | 2.5383 |
C2 | 1.3293 | | 1.3418 | 2.1551 | 2.2066 | 3.5246 | 4.4940 | 1.0669 | 2.1564 | 3.2467 | 3.8397 | C3 | 2.2034 | 1.3418 | | 2.2123 | 1.4296 | 3.6667 | 4.3563 | 2.2053 | 1.0674 | 2.2529 | 4.3152 | C4 | 1.3485 | 2.1551 | 2.2123 | | 1.3420 | 1.4650 | 2.3397 | 3.1451 | 3.2500 | 2.1486 | 2.1600 | C5 | 2.2079 | 2.2066 | 1.4296 | 1.3420 | | 2.5717 | 3.0328 | 3.2644 | 2.2485 | 1.0674 | 3.4522 | C6 | 2.3998 | 3.5246 | 3.6667 | 1.4650 | 2.5717 | | 1.1834 | 4.4189 | 4.6841 | 2.9675 | 1.0925 | O7 | 3.5009 | 4.4940 | 4.3563 | 2.3397 | 3.0328 | 1.1834 | | 5.4610 | 5.2767 | 3.0088 | 1.9887 | H8 | 2.0435 | 1.0669 | 2.2053 | 3.1451 | 3.2644 | 4.4189 | 5.4610 | | 2.6970 | 4.2888 | 4.5512 | H9 | 3.2310 | 2.1564 | 1.0674 | 3.2500 | 2.2485 | 4.6841 | 5.2767 | 2.6970 | | 2.7464 | 5.3756 | H10 | 3.2348 | 3.2467 | 2.2529 | 2.1486 | 1.0674 | 2.9675 | 3.0088 | 4.2888 | 2.7464 | | 3.9998 | H11 | 2.5383 | 3.8397 | 4.3152 | 2.1600 | 3.4522 | 1.0925 | 1.9887 | 4.5512 | 5.3756 | 3.9998 | |
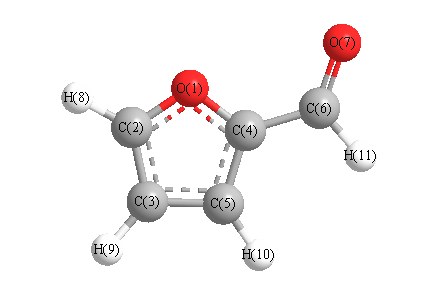 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.161 |
|
O1 |
C2 |
H8 |
116.605 |
| O1 |
C4 |
C5 |
110.295 |
|
O1 |
C4 |
C6 |
117.008 |
| C2 |
O1 |
C4 |
107.179 |
|
C2 |
C3 |
C5 |
105.494 |
| C2 |
C3 |
H9 |
126.658 |
|
C3 |
C2 |
H8 |
132.234 |
| C3 |
C5 |
C4 |
105.870 |
|
C3 |
C5 |
H10 |
128.318 |
| C4 |
C5 |
H10 |
125.811 |
|
C4 |
C6 |
O7 |
123.770 |
| C4 |
C6 |
H11 |
114.469 |
|
C5 |
C3 |
H9 |
127.848 |
| C5 |
C4 |
C6 |
132.697 |
|
O7 |
C6 |
H11 |
121.761 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.361 |
|
|
|
| 2 |
C |
0.137 |
|
|
|
| 3 |
C |
-0.142 |
|
|
|
| 4 |
C |
0.164 |
|
|
|
| 5 |
C |
-0.066 |
|
|
|
| 6 |
C |
0.246 |
|
|
|
| 7 |
O |
-0.402 |
|
|
|
| 8 |
H |
0.119 |
|
|
|
| 9 |
H |
0.113 |
|
|
|
| 10 |
H |
0.134 |
|
|
|
| 11 |
H |
0.059 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.569 |
-3.115 |
0.000 |
3.488 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.729 |
4.683 |
0.000 |
| y |
4.683 |
-40.634 |
0.000 |
| z |
0.000 |
0.000 |
-41.491 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.333 |
4.683 |
0.000 |
| y |
4.683 |
-0.523 |
0.000 |
| z |
0.000 |
0.000 |
-1.810 |
|
| Polar |
|---|
| 3z2-r2 | -3.620 |
|---|
| x2-y2 | 1.904 |
|---|
| xy | 4.683 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.009 |
-0.772 |
0.000 |
| y |
-0.772 |
11.644 |
0.000 |
| z |
0.000 |
0.000 |
5.217 |
<r2> (average value of r
2) Å
2
| <r2> |
190.384 |
| (<r2>)1/2 |
13.798 |
