Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -382.618683 |
| Energy at 298.15K | -382.627522 |
| HF Energy | -382.618683 |
| Nuclear repulsion energy | 404.532760 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3371 |
3056 |
3.86 |
|
|
|
| 2 |
A |
3360 |
3045 |
10.65 |
|
|
|
| 3 |
A |
3345 |
3032 |
24.00 |
|
|
|
| 4 |
A |
3334 |
3022 |
11.16 |
|
|
|
| 5 |
A |
3320 |
3009 |
0.15 |
|
|
|
| 6 |
A |
3284 |
2976 |
18.52 |
|
|
|
| 7 |
A |
3234 |
2931 |
13.69 |
|
|
|
| 8 |
A |
3177 |
2880 |
4.34 |
|
|
|
| 9 |
A |
1953 |
1770 |
284.50 |
|
|
|
| 10 |
A |
1785 |
1618 |
17.23 |
|
|
|
| 11 |
A |
1761 |
1596 |
7.26 |
|
|
|
| 12 |
A |
1653 |
1498 |
0.48 |
|
|
|
| 13 |
A |
1602 |
1452 |
9.51 |
|
|
|
| 14 |
A |
1598 |
1449 |
20.72 |
|
|
|
| 15 |
A |
1590 |
1441 |
9.56 |
|
|
|
| 16 |
A |
1523 |
1380 |
42.12 |
|
|
|
| 17 |
A |
1462 |
1325 |
5.32 |
|
|
|
| 18 |
A |
1378 |
1249 |
172.15 |
|
|
|
| 19 |
A |
1334 |
1209 |
3.09 |
|
|
|
| 20 |
A |
1290 |
1169 |
10.39 |
|
|
|
| 21 |
A |
1211 |
1098 |
6.86 |
|
|
|
| 22 |
A |
1184 |
1073 |
3.71 |
|
|
|
| 23 |
A |
1166 |
1057 |
0.21 |
|
|
|
| 24 |
A |
1152 |
1044 |
0.87 |
|
|
|
| 25 |
A |
1130 |
1024 |
0.17 |
|
|
|
| 26 |
A |
1116 |
1011 |
2.80 |
|
|
|
| 27 |
A |
1115 |
1010 |
0.22 |
|
|
|
| 28 |
A |
1086 |
985 |
0.67 |
|
|
|
| 29 |
A |
1057 |
958 |
2.47 |
|
|
|
| 30 |
A |
1035 |
938 |
30.08 |
|
|
|
| 31 |
A |
957 |
868 |
0.14 |
|
|
|
| 32 |
A |
851 |
772 |
47.54 |
|
|
|
| 33 |
A |
790 |
717 |
0.92 |
|
|
|
| 34 |
A |
764 |
692 |
41.04 |
|
|
|
| 35 |
A |
675 |
612 |
1.02 |
|
|
|
| 36 |
A |
657 |
596 |
17.19 |
|
|
|
| 37 |
A |
641 |
581 |
36.59 |
|
|
|
| 38 |
A |
494 |
448 |
1.24 |
|
|
|
| 39 |
A |
472 |
427 |
0.15 |
|
|
|
| 40 |
A |
453 |
411 |
0.00 |
|
|
|
| 41 |
A |
388 |
352 |
0.85 |
|
|
|
| 42 |
A |
234 |
212 |
5.79 |
|
|
|
| 43 |
A |
185 |
167 |
0.01 |
|
|
|
| 44 |
A |
169 |
153 |
0.01 |
|
|
|
| 45 |
A |
55 |
50 |
4.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32194.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29180.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.489 |
0.790 |
0.746 |
| C2 |
1.697 |
-0.136 |
-0.136 |
| C3 |
0.201 |
-0.048 |
-0.062 |
| O4 |
2.225 |
-0.950 |
-0.821 |
| C5 |
-0.458 |
1.141 |
-0.320 |
| C6 |
-0.532 |
-1.177 |
0.254 |
| C7 |
-1.836 |
1.198 |
-0.271 |
| C8 |
-1.909 |
-1.113 |
0.327 |
| C9 |
-2.562 |
0.073 |
0.061 |
| H10 |
3.548 |
0.627 |
0.611 |
| H11 |
2.255 |
1.824 |
0.524 |
| H12 |
2.224 |
0.618 |
1.783 |
| H13 |
0.098 |
2.020 |
-0.579 |
| H14 |
-0.025 |
-2.102 |
0.436 |
| H15 |
-2.341 |
2.118 |
-0.487 |
| H16 |
-2.470 |
-1.991 |
0.579 |
| H17 |
-3.632 |
0.119 |
0.109 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
C5 |
C6 |
C7 |
C8 |
C9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.5038 | 2.5681 | 2.3560 | 3.1545 | 3.6392 | 4.4619 | 4.8106 | 5.1478 | 1.0795 | 1.0830 | 1.0839 | 2.9983 | 3.8448 | 5.1590 | 5.6882 | 6.1905 |
C2 | 1.5038 | | 1.5011 | 1.1877 | 2.5124 | 2.4916 | 3.7790 | 3.7646 | 4.2690 | 2.1363 | 2.1413 | 2.1272 | 2.7208 | 2.6758 | 4.6379 | 4.6172 | 5.3406 | C3 | 2.5681 | 1.5011 | | 2.3425 | 1.3841 | 1.3824 | 2.3966 | 2.3947 | 2.7680 | 3.4806 | 2.8403 | 2.8182 | 2.1342 | 2.1256 | 3.3663 | 3.3640 | 3.8396 | O4 | 2.3560 | 1.1877 | 2.3425 | | 3.4387 | 2.9685 | 4.6269 | 4.2933 | 4.9740 | 2.5070 | 3.0824 | 3.0391 | 3.6610 | 2.8235 | 5.5111 | 5.0090 | 6.0258 | C5 | 3.1545 | 2.5124 | 1.3841 | 3.4387 | | 2.3892 | 1.3795 | 2.7576 | 2.3900 | 4.1455 | 2.9223 | 3.4484 | 1.0716 | 3.3586 | 2.1273 | 3.8292 | 3.3611 | C6 | 3.6392 | 2.4916 | 1.3824 | 2.9685 | 2.3892 | | 2.7592 | 1.3795 | 2.3910 | 4.4756 | 4.1043 | 3.6274 | 3.3630 | 1.0712 | 3.8309 | 2.1265 | 3.3623 | C7 | 4.4619 | 3.7790 | 2.3966 | 4.6269 | 1.3795 | 2.7592 | | 2.3882 | 1.3796 | 5.4856 | 4.2138 | 4.5864 | 2.1235 | 3.8302 | 1.0717 | 3.3600 | 2.1287 | C8 | 4.8106 | 3.7646 | 2.3947 | 4.2933 | 2.7576 | 1.3795 | 2.3882 | | 1.3799 | 5.7344 | 5.0988 | 4.7113 | 3.8291 | 2.1304 | 3.3602 | 1.0717 | 2.1297 | C9 | 5.1478 | 4.2690 | 2.7680 | 4.9740 | 2.3900 | 2.3910 | 1.3796 | 1.3799 | | 6.1596 | 5.1459 | 5.1153 | 3.3578 | 3.3628 | 2.1291 | 2.1292 | 1.0717 | H10 | 1.0795 | 2.1363 | 3.4806 | 2.5070 | 4.1455 | 4.4756 | 5.4856 | 5.7344 | 6.1596 | | 1.7646 | 1.7677 | 3.9070 | 4.4990 | 6.1733 | 6.5626 | 7.2149 | H11 | 1.0830 | 2.1413 | 2.8403 | 3.0824 | 2.9223 | 4.1043 | 4.2138 | 5.0988 | 5.1459 | 1.7646 | | 1.7432 | 2.4307 | 4.5405 | 4.7145 | 6.0724 | 6.1420 | H12 | 1.0839 | 2.1272 | 2.8182 | 3.0391 | 3.4484 | 3.6274 | 4.5864 | 4.7113 | 5.1153 | 1.7677 | 1.7432 | | 3.4734 | 3.7775 | 5.3139 | 5.5034 | 6.1104 | H13 | 2.9983 | 2.7208 | 2.1342 | 3.6610 | 1.0716 | 3.3630 | 2.1235 | 3.8291 | 3.3578 | 3.9070 | 2.4307 | 3.4734 | | 4.2471 | 2.4421 | 4.9008 | 4.2417 | H14 | 3.8448 | 2.6758 | 2.1256 | 2.8235 | 3.3586 | 1.0712 | 3.8302 | 2.1304 | 3.3628 | 4.4990 | 4.5405 | 3.7775 | 4.2471 | | 4.9018 | 2.4520 | 4.2487 | H15 | 5.1590 | 4.6379 | 3.3663 | 5.5111 | 2.1273 | 3.8309 | 1.0717 | 3.3602 | 2.1291 | 6.1733 | 4.7145 | 5.3139 | 2.4421 | 4.9018 | | 4.2466 | 2.4528 | H16 | 5.6882 | 4.6172 | 3.3640 | 5.0090 | 3.8292 | 2.1265 | 3.3600 | 1.0717 | 2.1292 | 6.5626 | 6.0724 | 5.5034 | 4.9008 | 2.4520 | 4.2466 | | 2.4539 | H17 | 6.1905 | 5.3406 | 3.8396 | 6.0258 | 3.3611 | 3.3623 | 2.1287 | 2.1297 | 1.0717 | 7.2149 | 6.1420 | 6.1104 | 4.2417 | 4.2487 | 2.4528 | 2.4539 | |
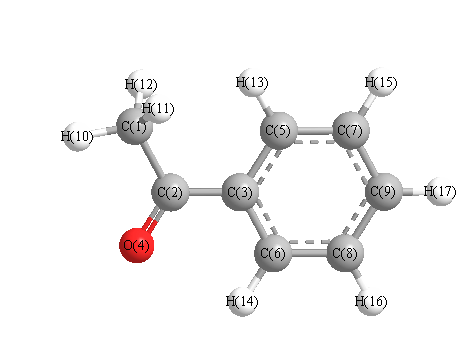 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
117.436 |
|
C1 |
C2 |
O4 |
121.730 |
| C2 |
C1 |
H10 |
110.502 |
|
C2 |
C1 |
H11 |
110.685 |
| C2 |
C1 |
H12 |
109.512 |
|
C2 |
C3 |
C5 |
121.050 |
| C2 |
C3 |
C6 |
119.495 |
|
C3 |
C2 |
O4 |
120.747 |
| C3 |
C5 |
C7 |
120.269 |
|
C3 |
C5 |
H13 |
120.171 |
| C3 |
C6 |
C8 |
120.231 |
|
C3 |
C6 |
H14 |
119.530 |
| C5 |
C3 |
C6 |
119.451 |
|
C5 |
C7 |
C9 |
120.042 |
| C5 |
C7 |
H15 |
119.895 |
|
C6 |
C8 |
C9 |
120.116 |
| C6 |
C8 |
H16 |
119.831 |
|
C7 |
C5 |
H13 |
119.549 |
| C7 |
C9 |
C8 |
119.875 |
|
C7 |
C9 |
H17 |
120.030 |
| C8 |
C6 |
H14 |
120.238 |
|
C8 |
C9 |
H17 |
120.095 |
| C9 |
C7 |
H15 |
120.062 |
|
C9 |
C8 |
H16 |
120.049 |
| H10 |
C1 |
H11 |
109.366 |
|
H10 |
C1 |
H12 |
109.594 |
| H11 |
C1 |
H12 |
107.115 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.206 |
|
|
|
| 2 |
C |
0.374 |
|
|
|
| 3 |
C |
-0.050 |
|
|
|
| 4 |
O |
-0.430 |
|
|
|
| 5 |
C |
-0.106 |
|
|
|
| 6 |
C |
-0.069 |
|
|
|
| 7 |
C |
-0.123 |
|
|
|
| 8 |
C |
-0.099 |
|
|
|
| 9 |
C |
-0.063 |
|
|
|
| 10 |
H |
0.114 |
|
|
|
| 11 |
H |
0.071 |
|
|
|
| 12 |
H |
0.071 |
|
|
|
| 13 |
H |
0.092 |
|
|
|
| 14 |
H |
0.128 |
|
|
|
| 15 |
H |
0.098 |
|
|
|
| 16 |
H |
0.100 |
|
|
|
| 17 |
H |
0.099 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.871 |
2.647 |
0.003 |
3.241 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.377 |
7.072 |
0.007 |
| y |
7.072 |
-51.177 |
-0.009 |
| z |
0.007 |
-0.009 |
-56.103 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.264 |
7.072 |
0.007 |
| y |
7.072 |
2.563 |
-0.009 |
| z |
0.007 |
-0.009 |
-4.826 |
|
| Polar |
|---|
| 3z2-r2 | -9.653 |
|---|
| x2-y2 | -0.199 |
|---|
| xy | 7.072 |
|---|
| xz | 0.007 |
|---|
| yz | -0.009 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
16.852 |
-0.189 |
-0.000 |
| y |
-0.189 |
14.147 |
-0.001 |
| z |
-0.000 |
-0.001 |
8.003 |
<r2> (average value of r
2) Å
2
| <r2> |
342.046 |
| (<r2>)1/2 |
18.494 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -382.618683 |
| Energy at 298.15K | -382.627518 |
| HF Energy | -382.618683 |
| Nuclear repulsion energy | 404.543038 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3372 |
3056 |
3.85 |
|
|
|
| 2 |
A' |
3360 |
3046 |
10.62 |
|
|
|
| 3 |
A' |
3346 |
3033 |
23.93 |
|
|
|
| 4 |
A' |
3334 |
3022 |
11.25 |
|
|
|
| 5 |
A' |
3320 |
3010 |
0.15 |
|
|
|
| 6 |
A' |
3285 |
2977 |
18.44 |
|
|
|
| 7 |
A' |
3177 |
2880 |
4.37 |
|
|
|
| 8 |
A' |
1952 |
1769 |
284.18 |
|
|
|
| 9 |
A' |
1785 |
1618 |
17.43 |
|
|
|
| 10 |
A' |
1761 |
1596 |
7.28 |
|
|
|
| 11 |
A' |
1653 |
1498 |
0.49 |
|
|
|
| 12 |
A' |
1598 |
1449 |
20.74 |
|
|
|
| 13 |
A' |
1590 |
1441 |
9.59 |
|
|
|
| 14 |
A' |
1523 |
1380 |
42.30 |
|
|
|
| 15 |
A' |
1462 |
1325 |
5.39 |
|
|
|
| 16 |
A' |
1378 |
1249 |
171.94 |
|
|
|
| 17 |
A' |
1333 |
1209 |
3.12 |
|
|
|
| 18 |
A' |
1290 |
1169 |
10.39 |
|
|
|
| 19 |
A' |
1211 |
1098 |
6.90 |
|
|
|
| 20 |
A' |
1184 |
1073 |
3.72 |
|
|
|
| 21 |
A' |
1166 |
1057 |
0.21 |
|
|
|
| 22 |
A' |
1116 |
1011 |
2.81 |
|
|
|
| 23 |
A' |
1086 |
985 |
0.65 |
|
|
|
| 24 |
A' |
1035 |
938 |
29.99 |
|
|
|
| 25 |
A' |
791 |
717 |
0.91 |
|
|
|
| 26 |
A' |
675 |
612 |
1.03 |
|
|
|
| 27 |
A' |
641 |
581 |
36.59 |
|
|
|
| 28 |
A' |
494 |
448 |
1.25 |
|
|
|
| 29 |
A' |
388 |
352 |
0.85 |
|
|
|
| 30 |
A' |
234 |
212 |
5.77 |
|
|
|
| 31 |
A" |
3234 |
2931 |
13.71 |
|
|
|
| 32 |
A" |
1602 |
1452 |
9.51 |
|
|
|
| 33 |
A" |
1152 |
1044 |
0.87 |
|
|
|
| 34 |
A" |
1129 |
1024 |
0.17 |
|
|
|
| 35 |
A" |
1115 |
1010 |
0.23 |
|
|
|
| 36 |
A" |
1057 |
958 |
2.46 |
|
|
|
| 37 |
A" |
957 |
868 |
0.15 |
|
|
|
| 38 |
A" |
851 |
772 |
47.43 |
|
|
|
| 39 |
A" |
764 |
692 |
41.10 |
|
|
|
| 40 |
A" |
657 |
596 |
17.22 |
|
|
|
| 41 |
A" |
472 |
427 |
0.15 |
|
|
|
| 42 |
A" |
453 |
411 |
0.00 |
|
|
|
| 43 |
A" |
183 |
166 |
0.01 |
|
|
|
| 44 |
A" |
169 |
153 |
0.01 |
|
|
|
| 45 |
A" |
55 |
50 |
4.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32194.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29181.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.737 |
2.134 |
0.000 |
| C2 |
0.298 |
1.676 |
0.000 |
| C3 |
0.000 |
0.207 |
0.000 |
| O4 |
-0.586 |
2.472 |
0.000 |
| C5 |
0.999 |
-0.753 |
0.000 |
| C6 |
-1.328 |
-0.200 |
0.000 |
| C7 |
0.681 |
-2.097 |
0.000 |
| C8 |
-1.647 |
-1.540 |
0.000 |
| C9 |
-0.642 |
-2.491 |
0.000 |
| H10 |
1.755 |
3.212 |
0.000 |
| H11 |
2.256 |
1.764 |
0.876 |
| H12 |
2.256 |
1.764 |
-0.876 |
| H13 |
2.031 |
-0.464 |
0.000 |
| H14 |
-2.096 |
0.546 |
0.000 |
| H15 |
1.462 |
-2.832 |
0.000 |
| H16 |
-2.675 |
-1.845 |
0.000 |
| H17 |
-0.890 |
-3.535 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
C5 |
C6 |
C7 |
C8 |
C9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.5102 | 2.5947 | 2.3473 | 2.9797 | 3.8526 | 4.3614 | 4.9951 | 5.2008 | 1.0783 | 1.0833 | 1.0833 | 2.6148 | 4.1487 | 4.9738 | 5.9418 | 6.2481 |
C2 | 1.5102 | | 1.4994 | 1.1893 | 2.5281 | 2.4827 | 3.7929 | 3.7585 | 4.2716 | 2.1172 | 2.1469 | 2.1469 | 2.7540 | 2.6470 | 4.6561 | 4.6088 | 5.3447 | C3 | 2.5947 | 1.4994 | | 2.3401 | 1.3852 | 1.3887 | 2.4024 | 2.4007 | 2.7728 | 3.4805 | 2.8777 | 2.8777 | 2.1388 | 2.1229 | 3.3721 | 3.3717 | 3.8459 | O4 | 2.3473 | 1.1893 | 2.3401 | | 3.5934 | 2.7736 | 4.7418 | 4.1501 | 4.9634 | 2.4549 | 3.0570 | 3.0570 | 3.9331 | 2.4476 | 5.6858 | 4.7966 | 6.0147 | C5 | 2.9797 | 2.5281 | 1.3852 | 3.5934 | | 2.3916 | 1.3819 | 2.7609 | 2.3903 | 4.0364 | 2.9460 | 2.9460 | 1.0714 | 3.3562 | 2.1302 | 3.8335 | 3.3629 | C6 | 3.8526 | 2.4827 | 1.3887 | 2.7736 | 2.3916 | | 2.7626 | 1.3772 | 2.3911 | 4.5988 | 4.1795 | 4.1795 | 3.3691 | 1.0706 | 3.8352 | 2.1265 | 3.3631 | C7 | 4.3614 | 3.7929 | 2.4024 | 4.7418 | 1.3819 | 2.7626 | | 2.3934 | 1.3794 | 5.4173 | 4.2612 | 4.2612 | 2.1193 | 3.8331 | 1.0726 | 3.3653 | 2.1290 | C8 | 4.9951 | 3.7585 | 2.4007 | 4.1501 | 2.7609 | 1.3772 | 2.3934 | | 1.3839 | 5.8445 | 5.1880 | 5.1880 | 3.8322 | 2.1334 | 3.3669 | 1.0727 | 2.1336 | C9 | 5.2008 | 4.2716 | 2.7728 | 4.9634 | 2.3903 | 2.3911 | 1.3794 | 1.3839 | | 6.1862 | 5.2216 | 5.2216 | 3.3542 | 3.3668 | 2.1311 | 2.1337 | 1.0731 | H10 | 1.0783 | 2.1172 | 3.4805 | 2.4549 | 4.0364 | 4.5988 | 5.4173 | 5.8445 | 6.1862 | | 1.7654 | 1.7654 | 3.6868 | 4.6837 | 6.0514 | 6.7236 | 7.2470 | H11 | 1.0833 | 2.1469 | 2.8777 | 3.0570 | 2.9460 | 4.1795 | 4.2612 | 5.1880 | 5.2216 | 1.7654 | | 1.7513 | 2.4043 | 4.6030 | 4.7453 | 6.1734 | 6.2241 | H12 | 1.0833 | 2.1469 | 2.8777 | 3.0570 | 2.9460 | 4.1795 | 4.2612 | 5.1880 | 5.2216 | 1.7654 | 1.7513 | | 2.4043 | 4.6030 | 4.7453 | 6.1734 | 6.2241 | H13 | 2.6148 | 2.7540 | 2.1388 | 3.9331 | 1.0714 | 3.3691 | 2.1193 | 3.8322 | 3.3542 | 3.6868 | 2.4043 | 2.4043 | | 4.2484 | 2.4353 | 4.9048 | 4.2381 | H14 | 4.1487 | 2.6470 | 2.1229 | 2.4476 | 3.3562 | 1.0706 | 3.8331 | 2.1334 | 3.3668 | 4.6837 | 4.6030 | 4.6030 | 4.2484 | | 4.9057 | 2.4604 | 4.2549 | H15 | 4.9738 | 4.6561 | 3.3721 | 5.6858 | 2.1302 | 3.8352 | 1.0726 | 3.3669 | 2.1311 | 6.0514 | 4.7453 | 4.7453 | 2.4353 | 4.9057 | | 4.2534 | 2.4548 | H16 | 5.9418 | 4.6088 | 3.3717 | 4.7966 | 3.8335 | 2.1265 | 3.3653 | 1.0727 | 2.1337 | 6.7236 | 6.1734 | 6.1734 | 4.9048 | 2.4604 | 4.2534 | | 2.4579 | H17 | 6.2481 | 5.3447 | 3.8459 | 6.0147 | 3.3629 | 3.3631 | 2.1290 | 2.1336 | 1.0731 | 7.2470 | 6.2241 | 6.2241 | 4.2381 | 4.2549 | 2.4548 | 2.4579 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
119.109 |
|
C1 |
C2 |
O4 |
120.336 |
| C2 |
C1 |
H10 |
108.603 |
|
C2 |
C1 |
H11 |
110.667 |
| C2 |
C1 |
H12 |
110.667 |
|
C2 |
C3 |
C5 |
122.379 |
| C2 |
C3 |
C6 |
118.498 |
|
C3 |
C2 |
O4 |
120.555 |
| C3 |
C5 |
C7 |
120.502 |
|
C3 |
C5 |
H13 |
120.535 |
| C3 |
C6 |
C8 |
120.448 |
|
C3 |
C6 |
H14 |
118.784 |
| C5 |
C3 |
C6 |
119.123 |
|
C5 |
C7 |
C9 |
119.907 |
| C5 |
C7 |
H15 |
119.894 |
|
C6 |
C8 |
C9 |
119.996 |
| C6 |
C8 |
H16 |
119.949 |
|
C7 |
C5 |
H13 |
118.963 |
| C7 |
C9 |
C8 |
120.024 |
|
C7 |
C9 |
H17 |
119.959 |
| C8 |
C6 |
H14 |
120.767 |
|
C8 |
C9 |
H17 |
120.017 |
| C9 |
C7 |
H15 |
120.199 |
|
C9 |
C8 |
H16 |
120.056 |
| H10 |
C1 |
H11 |
109.509 |
|
H10 |
C1 |
H12 |
109.509 |
| H11 |
C1 |
H12 |
107.873 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.206 |
|
|
|
| 2 |
C |
0.374 |
|
|
|
| 3 |
C |
-0.050 |
|
|
|
| 4 |
O |
-0.430 |
|
|
|
| 5 |
C |
-0.106 |
|
|
|
| 6 |
C |
-0.069 |
|
|
|
| 7 |
C |
-0.123 |
|
|
|
| 8 |
C |
-0.099 |
|
|
|
| 9 |
C |
-0.062 |
|
|
|
| 10 |
H |
0.114 |
|
|
|
| 11 |
H |
0.071 |
|
|
|
| 12 |
H |
0.071 |
|
|
|
| 13 |
H |
0.092 |
|
|
|
| 14 |
H |
0.128 |
|
|
|
| 15 |
H |
0.098 |
|
|
|
| 16 |
H |
0.100 |
|
|
|
| 17 |
H |
0.099 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.985 |
-2.564 |
0.000 |
3.243 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.242 |
5.812 |
0.000 |
| y |
5.812 |
-55.309 |
0.000 |
| z |
0.000 |
0.000 |
-56.102 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.464 |
5.812 |
0.000 |
| y |
5.812 |
-3.637 |
0.000 |
| z |
0.000 |
0.000 |
-4.827 |
|
| Polar |
|---|
| 3z2-r2 | -9.654 |
|---|
| x2-y2 | 8.067 |
|---|
| xy | 5.812 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.271 |
0.599 |
0.000 |
| y |
0.599 |
16.728 |
0.000 |
| z |
0.000 |
0.000 |
8.002 |
<r2> (average value of r
2) Å
2
| <r2> |
342.036 |
| (<r2>)1/2 |
18.494 |
