return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2CH3 (Ethylbenzene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-309.933844
Energy at 298.15K-309.943977
HF Energy-308.798219
Nuclear repulsion energy340.308321
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3274 3075 15.88      
2 A' 3252 3054 2.67      
3 A' 3234 3037 9.73      
4 A' 3210 3015 37.88      
5 A' 3115 2926 32.05      
6 A' 3113 2924 29.91      
7 A' 1675 1574 3.64      
8 A' 1553 1458 11.22      
9 A' 1544 1450 1.52      
10 A' 1530 1437 1.24      
11 A' 1448 1360 3.27      
12 A' 1386 1302 2.12      
13 A' 1254 1178 0.23      
14 A' 1223 1149 0.12      
15 A' 1110 1043 6.39      
16 A' 1064 999 3.13      
17 A' 1022 960 0.29      
18 A' 1002 941 2.33      
19 A' 904 849 0.08      
20 A' 872 819 0.58      
21 A' 790 742 0.65      
22 A' 733 689 85.96      
23 A' 573 538 11.94      
24 A' 494 464 6.62      
25 A' 429 403 0.69      
26 A' 295 277 0.14      
27 A' 128 120 1.03      
28 A" 3261 3063 29.52      
29 A" 3235 3038 8.80      
30 A" 3215 3020 25.03      
31 A" 3170 2977 9.14      
32 A" 1653 1553 0.15      
33 A" 1541 1447 9.99      
34 A" 1504 1413 4.48      
35 A" 1473 1384 0.58      
36 A" 1374 1291 0.59      
37 A" 1301 1222 0.02      
38 A" 1203 1130 0.01      
39 A" 1142 1072 3.26      
40 A" 1075 1010 3.22      
41 A" 912 856 0.00      
42 A" 841 790 0.04      
43 A" 803 754 0.22      
44 A" 633 595 0.02      
45 A" 390 366 0.00      
46 A" 361 339 0.06      
47 A" 224 211 0.01      
48 A" 38 36 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 34786.9 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 32671.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.15032 0.04919 0.04077

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.244 0.504 0.000
C2 -0.246 -0.214 1.204
C3 -0.246 -1.610 1.207
C4 -0.245 -2.313 0.000
C5 -0.246 -1.610 -1.207
C6 -0.246 -0.214 -1.204
C7 -0.174 2.007 0.000
C8 1.272 2.509 0.000
H9 -0.253 0.327 2.144
H10 -0.252 -2.147 2.148
H11 -0.248 -3.395 0.000
H12 -0.252 -2.147 -2.148
H13 -0.253 0.327 -2.144
H14 -0.696 2.394 -0.877
H15 -0.696 2.394 0.877
H16 1.801 2.147 0.881
H17 1.801 2.147 -0.881
H18 1.310 3.598 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.40112.43422.81622.43421.40111.50502.51382.15123.41123.89843.41122.15122.13202.13202.76802.76803.4629
C21.40111.39652.41972.78632.40742.52683.34131.08442.15133.40133.86883.39093.36602.66563.14173.75634.2894
C32.43421.39651.39702.41482.78633.81384.55262.15121.08262.15483.39763.87074.53624.04224.29134.76115.5680
C42.81622.41971.39701.39702.41974.32015.05443.40032.15391.08222.15393.40034.80844.80844.98534.98536.1118
C52.43422.78632.41481.39701.39653.81384.55263.87073.39762.15481.08262.15124.04224.53624.76114.29135.5680
C61.40112.40742.78632.41971.39652.52683.34133.39093.86883.40132.15131.08442.66563.36603.75633.14174.2894
C71.50502.52683.81384.32013.81382.52681.53012.72514.67675.40224.67672.72511.09181.09182.16712.16712.1757
C82.51383.34134.55265.05444.55263.34131.53013.41795.34876.09615.34873.41792.15742.15741.08941.08941.0898
H92.15121.08442.15123.40033.87073.39092.72513.41792.47344.29484.95324.28793.68752.46433.02154.08424.2120
H103.41122.15131.08262.15393.39763.86884.67675.34872.47342.48384.29504.95325.47374.73554.92555.64146.3289
H113.89843.40132.15481.08222.15483.40135.40226.09614.29482.48382.48384.29485.87165.87165.97415.97417.1643
H123.41123.86883.39762.15391.08262.15134.67675.34874.95324.29502.48382.47344.73555.47375.64144.92556.3289
H132.15123.39093.87073.40032.15121.08442.72513.41794.28794.95324.29482.47342.46433.68754.08423.02154.2120
H142.13203.36604.53624.80844.04222.66561.09182.15743.68755.47375.87164.73552.46431.75483.06392.50942.4990
H152.13202.66564.04224.80844.53623.36601.09182.15742.46434.73555.87165.47373.68751.75482.50943.06392.4990
H162.76803.14174.29134.98534.76113.75632.16711.08943.02154.92555.97415.64144.08423.06392.50941.76121.7667
H172.76803.75634.76114.98534.29133.14172.16711.08944.08425.64145.97414.92553.02152.50943.06391.76121.7667
H183.46294.28945.56806.11185.56804.28942.17571.08984.21206.32897.16436.32894.21202.49902.49901.76671.7667

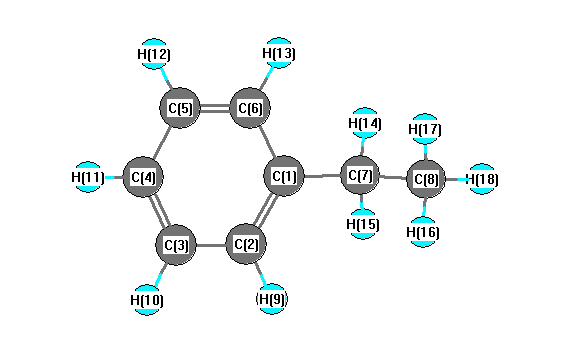
picture of Ethylbenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.939 C1 C2 H9 119.338
C1 C6 C5 120.939 C1 C6 H13 119.338
C1 C7 C8 111.839 C1 C7 H14 109.349
C1 C7 H15 109.349 C2 C1 C6 118.427
C2 C1 C7 120.753 C2 C3 C4 120.043
C2 C3 H10 119.879 C3 C2 H9 119.722
C3 C4 C5 119.609 C3 C4 H11 120.195
C4 C3 H10 120.077 C4 C5 C6 120.043
C4 C5 H12 120.077 C5 C4 H11 120.195
C5 C6 H13 119.722 C6 C1 C7 120.753
C6 C5 H12 119.879 C7 C8 H16 110.514
C7 C8 H17 110.514 C7 C8 H18 111.177
C8 C7 H14 109.609 C8 C7 H15 109.609
H14 C7 H15 106.961 H16 C8 H17 107.866
H16 C8 H18 108.332 H17 C8 H18 108.332
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability