return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CHO (benzaldehyde)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-344.593875
Energy at 298.15K 
HF Energy-343.451418
Nuclear repulsion energy320.485022
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3280 3081 10.85      
2 A' 3275 3075 9.48      
3 A' 3265 3067 8.89      
4 A' 3255 3057 0.68      
5 A' 3241 3044 4.53      
6 A' 3012 2829 102.48      
7 A' 1741 1635 183.45      
8 A' 1673 1571 15.41      
9 A' 1654 1554 12.81      
10 A' 1548 1454 0.95      
11 A' 1507 1416 10.21      
12 A' 1483 1393 3.03      
13 A' 1439 1351 6.12      
14 A' 1358 1276 13.41      
15 A' 1257 1180 58.62      
16 A' 1214 1140 8.25      
17 A' 1207 1133 1.60      
18 A' 1118 1050 4.14      
19 A' 1058 994 2.35      
20 A' 1019 957 0.71      
21 A' 846 795 31.58      
22 A' 659 619 20.04      
23 A' 627 589 0.67      
24 A' 443 416 0.29      
25 A' 220 206 7.84      
26 A" 998 937 1.18      
27 A" 923 867 0.07      
28 A" 909 853 0.20      
29 A" 884 830 0.66      
30 A" 844 793 0.02      
31 A" 729 685 100.23      
32 A" 410 385 4.41      
33 A" 378 355 0.01      
34 A" 324 304 0.06      
35 A" 198 186 10.17      
36 A" 95 89 4.37      

Unscaled Zero Point Vibrational Energy (zpe) 24044.8 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 22582.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.17441 0.05203 0.04007

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.572 0.000
C2 -1.043 -0.364 0.000
C3 -0.745 -1.724 0.000
C4 0.587 -2.154 0.000
C5 1.626 -1.220 0.000
C6 1.333 0.144 0.000
C7 -0.292 2.021 0.000
O8 -1.423 2.503 0.000
H9 0.594 2.680 0.000
H10 -2.065 -0.009 0.000
H11 -1.546 -2.452 0.000
H12 0.813 -3.212 0.000
H13 2.656 -1.554 0.000
H14 2.132 0.877 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 H9 H10 H11 H12 H13 H14
C11.40162.41412.78822.42001.40001.47832.39862.18992.14573.39653.87043.40182.1539
C21.40161.39232.42112.80352.43012.50062.89223.45631.08212.14793.39973.88553.4093
C32.41411.39231.40032.42482.79483.77264.28124.60322.16421.08212.15493.40523.8789
C42.78822.42111.40031.39692.41584.26645.07214.83343.41122.15401.08222.15333.4015
C52.42002.80352.42481.39691.39543.76644.81244.03413.88543.40322.15151.08202.1570
C61.40002.43012.79482.41581.39542.48273.62752.64103.40203.87693.39622.15251.0841
C71.47832.50063.77264.26643.76642.48271.22911.10432.69574.64585.34864.63362.6808
O82.39862.89224.28125.07214.81243.62751.22912.02472.59324.95706.13705.75263.9092
H92.18993.45634.60324.83344.03412.64101.10432.02473.78215.56045.89584.70892.3698
H102.14571.08212.16423.41123.88543.40202.69572.59323.78212.49794.30654.96744.2901
H113.39652.14791.08212.15403.40323.87694.64584.95705.56042.49792.47854.29634.9610
H123.87043.39972.15491.08222.15153.39625.34866.13705.89584.30652.47852.47844.2962
H133.40183.88553.40522.15331.08202.15254.63365.75264.70894.96744.29632.47842.4865
H142.15393.40933.87893.40152.15701.08412.68083.90922.36984.29014.96104.29622.4865

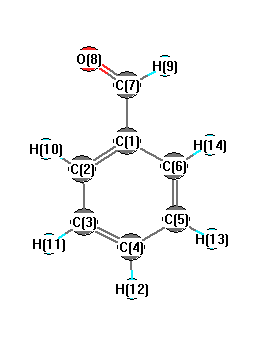
picture of benzaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.548 C1 C2 H10 118.953
C1 C6 C5 119.926 C1 C6 H14 119.697
C1 C7 O8 124.488 C1 C7 H9 115.221
C2 C1 C6 120.310 C2 C1 C7 120.503
C2 C3 C4 120.213 C2 C3 H11 119.942
C3 C2 H10 121.499 C3 C4 C5 120.198
C3 C4 H12 119.915 C4 C3 H11 119.845
C4 C5 C6 119.805 C4 C5 H13 120.077
C5 C4 H12 119.887 C5 C6 H14 120.377
C6 C1 C7 119.187 C6 C5 H13 120.117
O8 C7 H9 120.292
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability