return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4O2 (parabenzoquinone)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-380.418812
Energy at 298.15K 
Nuclear repulsion energy321.635147
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3285 3085 0.00      
2 Ag 1704 1600 0.00      
3 Ag 1640 1540 0.00      
4 Ag 1185 1113 0.00      
5 Ag 793 745 0.00      
6 Ag 453 425 0.00      
7 Au 955 897 0.00      
8 Au 299 281 0.00      
9 B1g 737 692 0.00      
10 B1u 3267 3068 0.14      
11 B1u 1688 1585 205.06      
12 B1u 1405 1320 4.00      
13 B1u 958 900 12.20      
14 B1u 765 718 0.12      
15 B2g 953 895 0.00      
16 B2g 567 532 0.00      
17 B2g 35i 33i 0.00      
18 B2u 3284 3084 2.50      
19 B2u 1645 1545 20.72      
20 B2u 1358 1276 73.37      
21 B2u 1105 1038 37.72      
22 B2u 409 384 24.24      
23 B3g 3267 3068 0.00      
24 B3g 1438 1350 0.00      
25 B3g 1271 1194 0.00      
26 B3g 606 570 0.00      
27 B3g 452 425 0.00      
28 B3u 860 808 88.17      
29 B3u 495 465 4.52      
30 B3u 81 76 14.07      

Unscaled Zero Point Vibrational Energy (zpe) 18443.7 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 17322.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.17540 0.05485 0.04178

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.438
C2 0.000 0.000 -1.438
C3 0.000 1.266 0.675
C4 0.000 -1.266 0.675
C5 0.000 -1.266 -0.675
C6 0.000 1.266 -0.675
O7 0.000 0.000 2.678
O8 0.000 0.000 -2.678
H9 0.000 2.180 1.255
H10 0.000 -2.180 1.255
H11 0.000 -2.180 -1.255
H12 0.000 2.180 -1.255

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 O8 H9 H10 H11 H12
C12.87661.47851.47852.46372.46371.24004.11662.18732.18733.46503.4650
C22.87662.46372.46371.47851.47854.11661.24003.46503.46502.18732.1873
C31.47852.46372.53242.86991.35022.36993.58451.08213.49433.94972.1356
C41.47852.46372.53241.35022.86992.36993.58453.49431.08212.13563.9497
C52.46371.47852.86991.35022.53243.58452.36993.94972.13561.08213.4943
C62.46371.47851.35022.86992.53243.58452.36992.13563.94973.49431.0821
O71.24004.11662.36992.36993.58453.58455.35662.60302.60304.49714.4971
O84.11661.24003.58453.58452.36992.36995.35664.49714.49712.60302.6030
H92.18733.46501.08213.49433.94972.13562.60304.49714.35925.03052.5107
H102.18733.46503.49431.08212.13563.94972.60304.49714.35922.51075.0305
H113.46502.18733.94972.13561.08213.49434.49712.60305.03052.51074.3592
H123.46502.18732.13563.94973.49431.08214.49712.60302.51075.03054.3592

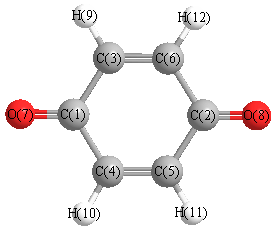
picture of parabenzoquinone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 121.081 C1 C3 H9 116.493
C1 C4 C5 121.081 C1 C4 H10 116.493
C2 C5 C4 121.081 C2 C5 H11 116.493
C2 C6 C3 121.081 C2 C6 H12 116.493
C3 C1 C4 117.839 C3 C1 O7 121.081
C3 C6 H12 122.427 C4 C1 O7 121.081
C4 C5 H11 122.427 C5 C2 C6 117.839
C5 C2 O8 121.081 C5 C4 H10 122.427
C6 C2 O8 121.081 C6 C3 H9 122.427
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability