Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -5218.468329 |
| Energy at 298.15K | -5218.478615 |
| HF Energy | -5217.886411 |
| Nuclear repulsion energy | 418.591820 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3188 |
2994 |
0.00 |
|
|
|
| 2 |
Ag |
1533 |
1440 |
0.00 |
|
|
|
| 3 |
Ag |
1351 |
1269 |
0.00 |
|
|
|
| 4 |
Ag |
1109 |
1042 |
0.00 |
|
|
|
| 5 |
Ag |
711 |
668 |
0.00 |
|
|
|
| 6 |
Ag |
199 |
187 |
0.00 |
|
|
|
| 7 |
Au |
3283 |
3083 |
0.00 |
|
|
|
| 8 |
Au |
1142 |
1073 |
4.33 |
|
|
|
| 9 |
Au |
783 |
735 |
3.05 |
|
|
|
| 10 |
Au |
133 |
125 |
4.78 |
|
|
|
| 11 |
Bg |
3262 |
3064 |
0.00 |
|
|
|
| 12 |
Bg |
1325 |
1245 |
0.00 |
|
|
|
| 13 |
Bg |
991 |
931 |
0.00 |
|
|
|
| 14 |
Bu |
3195 |
3001 |
5.41 |
|
|
|
| 15 |
Bu |
1528 |
1435 |
6.75 |
|
|
|
| 16 |
Bu |
1261 |
1184 |
65.69 |
|
|
|
| 17 |
Bu |
633 |
595 |
54.72 |
|
|
|
| 18 |
Bu |
180 |
169 |
6.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12903.7 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12119.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.490 |
0.571 |
0.000 |
| C2 |
-0.490 |
-0.571 |
0.000 |
| Br3 |
-0.490 |
2.258 |
0.000 |
| Br4 |
0.490 |
-2.258 |
0.000 |
| H5 |
1.112 |
0.571 |
0.889 |
| H6 |
1.112 |
0.571 |
-0.889 |
| H7 |
-1.112 |
-0.571 |
0.889 |
| H8 |
-1.112 |
-0.571 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5059 | 1.9507 | 2.8291 | 1.0849 | 1.0849 | 2.1596 | 2.1596 |
C2 | 1.5059 | | 2.8291 | 1.9507 | 2.1596 | 2.1596 | 1.0849 | 1.0849 | Br3 | 1.9507 | 2.8291 | | 4.6206 | 2.4905 | 2.4905 | 3.0298 | 3.0298 | Br4 | 2.8291 | 1.9507 | 4.6206 | | 3.0298 | 3.0298 | 2.4905 | 2.4905 | H5 | 1.0849 | 2.1596 | 2.4905 | 3.0298 | | 1.7780 | 2.5005 | 3.0682 | H6 | 1.0849 | 2.1596 | 2.4905 | 3.0298 | 1.7780 | | 3.0682 | 2.5005 | H7 | 2.1596 | 1.0849 | 3.0298 | 2.4905 | 2.5005 | 3.0682 | | 1.7780 | H8 | 2.1596 | 1.0849 | 3.0298 | 2.4905 | 3.0682 | 2.5005 | 1.7780 | |
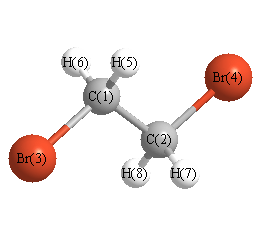 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
109.189 |
|
C1 |
C2 |
H7 |
111.906 |
| C1 |
C2 |
H8 |
111.906 |
|
C2 |
C1 |
Br3 |
109.189 |
| C2 |
C1 |
H5 |
111.906 |
|
C2 |
C1 |
H6 |
111.906 |
| Br3 |
C1 |
H5 |
106.758 |
|
Br3 |
C1 |
H6 |
106.758 |
| Br4 |
C2 |
H7 |
106.758 |
|
Br4 |
C2 |
H8 |
106.758 |
| H5 |
C1 |
H6 |
110.055 |
|
H7 |
C2 |
H8 |
110.055 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -5218.465744 |
| Energy at 298.15K | |
| HF Energy | -5217.882674 |
| Nuclear repulsion energy | 453.923213 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3246 |
3048 |
0.00 |
84.89 |
0.75 |
0.85 |
| 2 |
A |
3170 |
2978 |
13.47 |
194.33 |
0.02 |
0.04 |
| 3 |
A |
1519 |
1427 |
0.13 |
4.69 |
0.72 |
0.84 |
| 4 |
A |
1362 |
1279 |
25.12 |
4.88 |
0.38 |
0.55 |
| 5 |
A |
1237 |
1161 |
1.83 |
10.35 |
0.72 |
0.84 |
| 6 |
A |
1082 |
1016 |
0.03 |
2.96 |
0.72 |
0.84 |
| 7 |
A |
951 |
893 |
5.94 |
6.31 |
0.24 |
0.38 |
| 8 |
A |
583 |
548 |
7.43 |
20.54 |
0.06 |
0.12 |
| 9 |
A |
248 |
233 |
0.98 |
1.36 |
0.19 |
0.33 |
| 10 |
A |
76 |
71 |
0.22 |
0.64 |
0.73 |
0.84 |
| 11 |
B |
3256 |
3058 |
0.21 |
26.72 |
0.75 |
0.86 |
| 12 |
B |
3164 |
2972 |
2.23 |
38.59 |
0.75 |
0.86 |
| 13 |
B |
1513 |
1421 |
11.56 |
12.36 |
0.75 |
0.86 |
| 14 |
B |
1333 |
1252 |
69.22 |
1.42 |
0.75 |
0.86 |
| 15 |
B |
1160 |
1090 |
3.16 |
2.64 |
0.75 |
0.86 |
| 16 |
B |
886 |
832 |
15.25 |
0.89 |
0.75 |
0.86 |
| 17 |
B |
619 |
582 |
12.31 |
9.46 |
0.75 |
0.86 |
| 18 |
B |
377 |
354 |
5.36 |
2.12 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12891.1 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12107.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.317 |
0.683 |
1.182 |
| C2 |
-0.317 |
-0.683 |
1.182 |
| Br3 |
-0.317 |
1.783 |
-0.294 |
| Br4 |
0.317 |
-1.783 |
-0.294 |
| H5 |
0.042 |
1.213 |
2.091 |
| H6 |
1.398 |
0.627 |
1.099 |
| H7 |
-0.042 |
-1.213 |
2.091 |
| H8 |
-1.398 |
-0.627 |
1.099 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5066 | 1.9471 | 2.8741 | 1.0879 | 1.0856 | 2.1334 | 2.1601 |
C2 | 1.5066 | | 2.8741 | 1.9471 | 2.1334 | 2.1601 | 1.0879 | 1.0856 | Br3 | 1.9471 | 2.8741 | | 3.6225 | 2.4786 | 2.4944 | 3.8395 | 2.9858 | Br4 | 2.8741 | 1.9471 | 3.6225 | | 3.8395 | 2.9858 | 2.4786 | 2.4944 | H5 | 1.0879 | 2.1334 | 2.4786 | 3.8395 | | 1.7800 | 2.4272 | 2.5384 | H6 | 1.0856 | 2.1601 | 2.4944 | 2.9858 | 1.7800 | | 2.5384 | 3.0646 | H7 | 2.1334 | 1.0879 | 3.8395 | 2.4786 | 2.4272 | 2.5384 | | 1.7800 | H8 | 2.1601 | 1.0856 | 2.9858 | 2.4944 | 2.5384 | 3.0646 | 1.7800 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.018 |
|
C1 |
C2 |
H7 |
109.565 |
| C1 |
C2 |
H8 |
111.862 |
|
C2 |
C1 |
Br3 |
112.018 |
| C2 |
C1 |
H5 |
109.565 |
|
C2 |
C1 |
H6 |
111.862 |
| Br3 |
C1 |
H5 |
106.016 |
|
Br3 |
C1 |
H6 |
107.232 |
| Br4 |
C2 |
H7 |
106.016 |
|
Br4 |
C2 |
H8 |
107.232 |
| H5 |
C1 |
H6 |
109.961 |
|
H7 |
C2 |
H8 |
109.961 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
