return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2Br (n-propyl bromide)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-2688.203354
Energy at 298.15K-2688.213932
HF Energy-2687.603446
Nuclear repulsion energy236.865125
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3219 3023 23.21      
2 A' 3168 2976 17.09      
3 A' 3139 2948 8.92      
4 A' 3118 2929 24.95      
5 A' 1552 1458 6.89      
6 A' 1539 1445 1.43      
7 A' 1532 1439 1.70      
8 A' 1459 1370 2.75      
9 A' 1410 1324 9.76      
10 A' 1302 1223 43.64      
11 A' 1155 1085 1.19      
12 A' 1083 1017 1.37      
13 A' 938 881 8.09      
14 A' 695 653 19.12      
15 A' 323 303 1.42      
16 A' 214 201 1.36      
17 A" 3245 3048 9.07      
18 A" 3215 3020 30.13      
19 A" 3192 2998 0.47      
20 A" 1546 1452 9.28      
21 A" 1350 1268 0.03      
22 A" 1284 1206 0.80      
23 A" 1099 1032 2.39      
24 A" 885 831 0.02      
25 A" 768 722 2.63      
26 A" 247 232 0.01      
27 A" 157 147 1.00      

Unscaled Zero Point Vibrational Energy (zpe) 21417.1 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 20115.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.85615 0.05479 0.05302

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.826 0.000
C2 1.505 0.675 0.000
C3 2.173 2.046 0.000
Br4 -0.915 -0.896 0.000
H5 -0.351 1.348 0.886
H6 -0.351 1.348 -0.886
H7 1.811 0.104 0.877
H8 1.811 0.104 -0.877
H9 3.257 1.946 0.000
H10 1.891 2.620 -0.882
H11 1.891 2.620 0.882

Atom - Atom Distances (Å)
  C1 C2 C3 Br4 H5 H6 H7 H8 H9 H10 H11
C11.51232.49241.95001.08691.08692.13742.13743.44472.75192.7519
C21.51231.52522.88522.16422.16421.08991.08992.16522.17042.1704
C32.49241.52524.26542.76542.76542.16072.16071.08851.08961.0896
Br41.95002.88524.26542.47772.47773.03343.03345.04874.58444.5844
H51.08692.16422.76542.47771.77212.49453.05453.76393.12602.5782
H61.08692.16422.76542.47771.77213.05452.49453.76392.57823.1260
H72.13741.08992.16073.03342.49453.05451.75372.50063.07052.5170
H82.13741.08992.16073.03343.05452.49451.75372.50062.51703.0705
H93.44472.16521.08855.04873.76393.76392.50062.50061.76031.7603
H102.75192.17041.08964.58443.12602.57823.07052.51701.76031.7633
H112.75192.17041.08964.58442.57823.12602.51703.07051.76031.7633

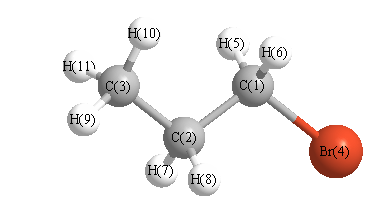
picture of n-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 110.276 C1 C2 H7 109.371
C1 C2 H8 109.371 C2 C1 Br4 112.262
C2 C1 H5 111.697 C2 C1 H6 111.697
C2 C3 H9 110.756 C2 C3 H10 111.116
C2 C3 H11 111.116 C3 C2 H7 110.319
C3 C2 H8 110.319 Br4 C1 H5 105.813
Br4 C1 H6 105.813 H5 C1 H6 109.220
H7 C2 H8 107.124 H9 C3 H10 107.840
H9 C3 H11 107.840 H10 C3 H11 108.031
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability