Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -3108.045379 |
| Energy at 298.15K | |
| HF Energy | -3107.463232 |
| Nuclear repulsion energy | 289.711909 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3193 |
2999 |
8.65 |
12.68 |
0.07 |
0.13 |
| 2 |
A' |
3184 |
2991 |
1.51 |
129.41 |
0.07 |
0.14 |
| 3 |
A' |
1535 |
1441 |
0.91 |
10.35 |
0.66 |
0.80 |
| 4 |
A' |
1532 |
1439 |
5.58 |
1.75 |
0.68 |
0.81 |
| 5 |
A' |
1389 |
1305 |
2.30 |
15.61 |
0.40 |
0.57 |
| 6 |
A' |
1286 |
1208 |
52.23 |
2.83 |
0.28 |
0.43 |
| 7 |
A' |
1110 |
1043 |
1.09 |
7.16 |
0.75 |
0.86 |
| 8 |
A' |
798 |
750 |
22.51 |
44.90 |
0.26 |
0.42 |
| 9 |
A' |
685 |
644 |
45.05 |
13.79 |
0.22 |
0.37 |
| 10 |
A' |
260 |
244 |
0.56 |
4.78 |
0.28 |
0.43 |
| 11 |
A' |
196 |
184 |
7.10 |
0.13 |
0.20 |
0.34 |
| 12 |
A" |
3278 |
3079 |
0.48 |
3.62 |
0.75 |
0.86 |
| 13 |
A" |
3256 |
3058 |
0.38 |
82.95 |
0.75 |
0.86 |
| 14 |
A" |
1332 |
1251 |
0.10 |
8.73 |
0.75 |
0.86 |
| 15 |
A" |
1170 |
1099 |
3.00 |
0.28 |
0.75 |
0.86 |
| 16 |
A" |
1026 |
964 |
0.32 |
4.44 |
0.75 |
0.86 |
| 17 |
A" |
794 |
746 |
2.29 |
0.03 |
0.75 |
0.86 |
| 18 |
A" |
145 |
136 |
5.70 |
0.01 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13085.0 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12289.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.632 |
0.000 |
| C2 |
1.242 |
-0.225 |
0.000 |
| Br3 |
-1.582 |
-0.501 |
0.000 |
| Cl4 |
2.674 |
0.841 |
0.000 |
| H5 |
-0.058 |
1.252 |
0.888 |
| H6 |
-0.058 |
1.252 |
-0.888 |
| H7 |
1.288 |
-0.849 |
0.887 |
| H8 |
1.288 |
-0.849 |
-0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5085 | 1.9463 | 2.6825 | 1.0850 | 1.0850 | 2.1539 | 2.1539 |
C2 | 1.5085 | | 2.8374 | 1.7855 | 2.1585 | 2.1585 | 1.0854 | 1.0854 | Br3 | 1.9463 | 2.8374 | | 4.4632 | 2.4878 | 2.4878 | 3.0244 | 3.0244 | Cl4 | 2.6825 | 1.7855 | 4.4632 | | 2.9021 | 2.9021 | 2.3586 | 2.3586 | H5 | 1.0850 | 2.1585 | 2.4878 | 2.9021 | | 1.7767 | 2.4955 | 3.0623 | H6 | 1.0850 | 2.1585 | 2.4878 | 2.9021 | 1.7767 | | 3.0623 | 2.4955 | H7 | 2.1539 | 1.0854 | 3.0244 | 2.3586 | 2.4955 | 3.0623 | | 1.7732 | H8 | 2.1539 | 1.0854 | 3.0244 | 2.3586 | 3.0623 | 2.4955 | 1.7732 | |
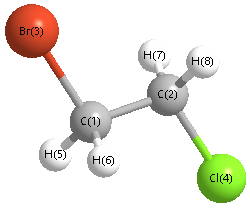 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
108.755 |
|
C1 |
C2 |
H7 |
111.227 |
| C1 |
C2 |
H8 |
111.227 |
|
C2 |
C1 |
Br3 |
109.781 |
| C2 |
C1 |
Cl4 |
39.071 |
|
C2 |
C1 |
H6 |
111.620 |
| Br3 |
C1 |
H5 |
106.840 |
|
Br3 |
C1 |
H6 |
106.840 |
| Cl4 |
C2 |
H7 |
107.988 |
|
Cl4 |
C2 |
H8 |
107.988 |
| H5 |
C1 |
H6 |
109.915 |
|
H7 |
C2 |
H8 |
109.540 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -3108.041806 |
| Energy at 298.15K | |
| HF Energy | -3107.458983 |
| Nuclear repulsion energy | 304.879190 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3255 |
3057 |
0.64 |
34.94 |
0.75 |
0.86 |
| 2 |
A |
3239 |
3042 |
1.04 |
77.80 |
0.75 |
0.86 |
| 3 |
A |
3173 |
2980 |
11.94 |
140.18 |
0.04 |
0.08 |
| 4 |
A |
3160 |
2967 |
9.40 |
89.77 |
0.20 |
0.33 |
| 5 |
A |
1518 |
1426 |
0.24 |
4.49 |
0.73 |
0.85 |
| 6 |
A |
1513 |
1421 |
12.31 |
12.82 |
0.73 |
0.85 |
| 7 |
A |
1395 |
1311 |
28.65 |
2.79 |
0.75 |
0.86 |
| 8 |
A |
1351 |
1269 |
52.75 |
2.58 |
0.27 |
0.43 |
| 9 |
A |
1259 |
1183 |
2.27 |
11.21 |
0.70 |
0.82 |
| 10 |
A |
1186 |
1114 |
2.40 |
3.09 |
0.72 |
0.83 |
| 11 |
A |
1093 |
1027 |
0.87 |
2.84 |
0.75 |
0.85 |
| 12 |
A |
974 |
915 |
7.58 |
6.26 |
0.21 |
0.35 |
| 13 |
A |
915 |
859 |
17.65 |
1.54 |
0.74 |
0.85 |
| 14 |
A |
722 |
678 |
16.18 |
9.14 |
0.37 |
0.54 |
| 15 |
A |
601 |
564 |
10.34 |
16.49 |
0.17 |
0.28 |
| 16 |
A |
403 |
379 |
5.72 |
1.90 |
0.70 |
0.82 |
| 17 |
A |
255 |
240 |
1.07 |
1.55 |
0.21 |
0.35 |
| 18 |
A |
94 |
88 |
0.63 |
0.99 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 13052.7 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12259.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.019 |
1.079 |
-0.394 |
| C2 |
1.278 |
0.881 |
0.413 |
| Br3 |
-1.363 |
-0.221 |
0.036 |
| Cl4 |
2.184 |
-0.565 |
-0.087 |
| H5 |
-0.407 |
2.053 |
-0.164 |
| H6 |
0.210 |
1.002 |
-1.459 |
| H7 |
1.930 |
1.738 |
0.253 |
| H8 |
1.055 |
0.793 |
1.472 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5084 | 1.9450 | 2.7358 | 1.0873 | 1.0856 | 2.1221 | 2.1535 |
C2 | 1.5084 | | 2.8859 | 1.7782 | 2.1322 | 2.1589 | 1.0886 | 1.0859 | Br3 | 1.9450 | 2.8859 | | 3.5650 | 2.4746 | 2.4909 | 3.8371 | 2.9897 | Cl4 | 2.7358 | 1.7782 | 3.5650 | | 3.6841 | 2.8696 | 2.3422 | 2.3557 | H5 | 1.0873 | 2.1322 | 2.4746 | 3.6841 | | 1.7784 | 2.3945 | 2.5308 | H6 | 1.0856 | 2.1589 | 2.4909 | 2.8696 | 1.7784 | | 2.5359 | 3.0582 | H7 | 2.1221 | 1.0886 | 3.8371 | 2.3422 | 2.3945 | 2.5359 | | 1.7731 | H8 | 2.1535 | 1.0859 | 2.9897 | 2.3557 | 2.5308 | 3.0582 | 1.7731 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.438 |
|
C1 |
C2 |
H7 |
108.523 |
| C1 |
C2 |
H8 |
111.174 |
|
C2 |
C1 |
Br3 |
112.767 |
| C2 |
C1 |
Cl4 |
36.925 |
|
C2 |
C1 |
H6 |
111.625 |
| Br3 |
C1 |
H5 |
105.901 |
|
Br3 |
C1 |
H6 |
107.117 |
| Cl4 |
C2 |
H7 |
107.107 |
|
Cl4 |
C2 |
H8 |
108.226 |
| H5 |
C1 |
H6 |
109.858 |
|
H7 |
C2 |
H8 |
109.257 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
