Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -997.620882 |
| Energy at 298.15K | -997.625668 |
| HF Energy | -997.038580 |
| Nuclear repulsion energy | 195.355225 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3182 |
2989 |
0.00 |
|
|
|
| 2 |
Ag |
1535 |
1442 |
0.00 |
|
|
|
| 3 |
Ag |
1413 |
1327 |
0.00 |
|
|
|
| 4 |
Ag |
1108 |
1041 |
0.00 |
|
|
|
| 5 |
Ag |
826 |
775 |
0.00 |
|
|
|
| 6 |
Ag |
314 |
295 |
0.00 |
|
|
|
| 7 |
Au |
3272 |
3073 |
2.80 |
|
|
|
| 8 |
Au |
1190 |
1117 |
2.45 |
|
|
|
| 9 |
Au |
802 |
754 |
1.57 |
|
|
|
| 10 |
Au |
126 |
119 |
6.73 |
|
|
|
| 11 |
Bg |
3252 |
3054 |
0.00 |
|
|
|
| 12 |
Bg |
1335 |
1254 |
0.00 |
|
|
|
| 13 |
Bg |
1064 |
999 |
0.00 |
|
|
|
| 14 |
Bu |
3189 |
2996 |
16.52 |
|
|
|
| 15 |
Bu |
1536 |
1442 |
6.53 |
|
|
|
| 16 |
Bu |
1321 |
1241 |
44.60 |
|
|
|
| 17 |
Bu |
780 |
733 |
77.75 |
|
|
|
| 18 |
Bu |
222 |
209 |
9.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13233.4 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12428.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.756 |
0.000 |
| C2 |
0.000 |
-0.756 |
0.000 |
| Cl3 |
-1.681 |
1.349 |
0.000 |
| Cl4 |
1.681 |
-1.349 |
0.000 |
| H5 |
0.493 |
1.144 |
0.886 |
| H6 |
0.493 |
1.144 |
-0.886 |
| H7 |
-0.493 |
-1.144 |
0.886 |
| H8 |
-0.493 |
-1.144 |
-0.886 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5119 | 1.7825 | 2.6940 | 1.0854 | 1.0854 | 2.1534 | 2.1534 |
C2 | 1.5119 | | 2.6940 | 1.7825 | 2.1534 | 2.1534 | 1.0854 | 1.0854 | Cl3 | 1.7825 | 2.6940 | | 4.3109 | 2.3562 | 2.3562 | 2.9005 | 2.9005 | Cl4 | 2.6940 | 1.7825 | 4.3109 | | 2.9005 | 2.9005 | 2.3562 | 2.3562 | H5 | 1.0854 | 2.1534 | 2.3562 | 2.9005 | | 1.7717 | 2.4912 | 3.0569 | H6 | 1.0854 | 2.1534 | 2.3562 | 2.9005 | 1.7717 | | 3.0569 | 2.4912 | H7 | 2.1534 | 1.0854 | 2.9005 | 2.3562 | 2.4912 | 3.0569 | | 1.7717 | H8 | 2.1534 | 1.0854 | 2.9005 | 2.3562 | 3.0569 | 2.4912 | 1.7717 | |
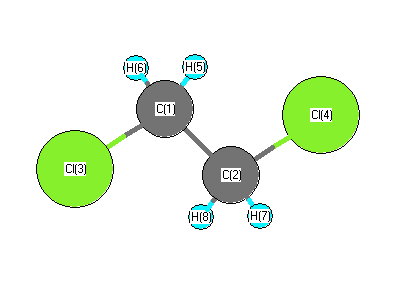 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.450 |
|
C1 |
C2 |
H7 |
110.943 |
| C1 |
C2 |
H8 |
110.943 |
|
C2 |
C1 |
Cl3 |
109.450 |
| C2 |
C1 |
H5 |
110.943 |
|
C2 |
C1 |
H6 |
110.943 |
| Cl3 |
C1 |
H5 |
108.007 |
|
Cl3 |
C1 |
H6 |
108.007 |
| Cl4 |
C2 |
H7 |
108.007 |
|
Cl4 |
C2 |
H8 |
108.007 |
| H5 |
C1 |
H6 |
109.397 |
|
H7 |
C2 |
H8 |
109.397 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -997.618574 |
| Energy at 298.15K | -997.623499 |
| HF Energy | -997.035454 |
| Nuclear repulsion energy | 203.130223 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3237 |
3040 |
0.99 |
|
|
|
| 2 |
A |
3169 |
2976 |
23.24 |
|
|
|
| 3 |
A |
1516 |
1424 |
0.04 |
|
|
|
| 4 |
A |
1404 |
1319 |
24.19 |
|
|
|
| 5 |
A |
1274 |
1197 |
1.30 |
|
|
|
| 6 |
A |
1096 |
1029 |
0.51 |
|
|
|
| 7 |
A |
1005 |
944 |
9.39 |
|
|
|
| 8 |
A |
709 |
666 |
16.46 |
|
|
|
| 9 |
A |
270 |
253 |
0.58 |
|
|
|
| 10 |
A |
123 |
115 |
0.80 |
|
|
|
| 11 |
B |
3248 |
3050 |
3.74 |
|
|
|
| 12 |
B |
3163 |
2971 |
4.43 |
|
|
|
| 13 |
B |
1510 |
1419 |
12.71 |
|
|
|
| 14 |
B |
1386 |
1302 |
40.82 |
|
|
|
| 15 |
B |
1206 |
1133 |
1.67 |
|
|
|
| 16 |
B |
936 |
879 |
17.46 |
|
|
|
| 17 |
B |
737 |
692 |
19.35 |
|
|
|
| 18 |
B |
425 |
399 |
6.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13206.2 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12403.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.290 |
0.697 |
0.894 |
| C2 |
-0.290 |
-0.697 |
0.894 |
| Cl3 |
-0.290 |
1.677 |
-0.472 |
| Cl4 |
0.290 |
-1.677 |
-0.472 |
| H5 |
-0.015 |
1.201 |
1.809 |
| H6 |
1.374 |
0.666 |
0.847 |
| H7 |
0.015 |
-1.201 |
1.809 |
| H8 |
-1.374 |
-0.666 |
0.847 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5102 | 1.7780 | 2.7390 | 1.0878 | 1.0862 | 2.1254 | 2.1519 |
C2 | 1.5102 | | 2.7390 | 1.7780 | 2.1254 | 2.1519 | 1.0878 | 1.0862 | Cl3 | 1.7780 | 2.7390 | | 3.4028 | 2.3461 | 2.3515 | 3.6849 | 2.8993 | Cl4 | 2.7390 | 1.7780 | 3.4028 | | 3.6849 | 2.8993 | 2.3461 | 2.3515 | H5 | 1.0878 | 2.1254 | 2.3461 | 3.6849 | | 1.7724 | 2.4029 | 2.5023 | H6 | 1.0862 | 2.1519 | 2.3515 | 2.8993 | 1.7724 | | 2.5023 | 3.0547 | H7 | 2.1254 | 1.0878 | 3.6849 | 2.3461 | 2.4029 | 2.5023 | | 1.7724 | H8 | 2.1519 | 1.0862 | 2.8993 | 2.3515 | 2.5023 | 3.0547 | 1.7724 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.562 |
|
C1 |
C2 |
H7 |
108.697 |
| C1 |
C2 |
H8 |
110.897 |
|
C2 |
C1 |
Cl3 |
112.562 |
| C2 |
C1 |
H5 |
108.697 |
|
C2 |
C1 |
H6 |
110.897 |
| Cl3 |
C1 |
H5 |
107.441 |
|
Cl3 |
C1 |
H6 |
107.921 |
| Cl4 |
C2 |
H7 |
107.441 |
|
Cl4 |
C2 |
H8 |
107.921 |
| H5 |
C1 |
H6 |
109.224 |
|
H7 |
C2 |
H8 |
109.224 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
