return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2CH3 (1-Propanamine)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-173.947888
Energy at 298.15K-173.958355
HF Energy-173.304673
Nuclear repulsion energy131.392907
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3574 3357 0.47      
2 A' 3215 3019 29.69      
3 A' 3119 2929 69.88      
4 A' 3115 2926 12.86      
5 A' 3105 2916 15.63      
6 A' 1699 1596 34.34      
7 A' 1554 1459 6.00      
8 A' 1541 1447 0.91      
9 A' 1531 1438 0.01      
10 A' 1457 1368 3.22      
11 A' 1438 1350 7.57      
12 A' 1358 1275 6.48      
13 A' 1169 1098 3.09      
14 A' 1132 1064 18.78      
15 A' 1073 1008 0.39      
16 A' 934 877 71.50      
17 A' 854 802 139.77      
18 A' 457 429 3.82      
19 A' 271 255 5.00      
20 A" 3684 3460 2.41      
21 A" 3209 3014 49.59      
22 A" 3181 2988 36.46      
23 A" 3153 2961 3.12      
24 A" 1548 1453 8.18      
25 A" 1411 1326 0.22      
26 A" 1355 1272 0.52      
27 A" 1276 1198 0.04      
28 A" 1050 987 0.12      
29 A" 881 827 1.33      
30 A" 762 715 0.89      
31 A" 281 264 47.82      
32 A" 234 220 9.84      
33 A" 137 128 0.46      

Unscaled Zero Point Vibrational Energy (zpe) 27377.9 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 25713.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.83360 0.12340 0.11639

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.428 1.284 0.000
C2 0.000 0.752 0.000
C3 -0.053 -0.774 0.000
N4 -1.386 -1.373 0.000
H5 1.448 2.373 0.000
H6 1.970 0.940 0.881
H7 1.970 0.940 -0.881
H8 -0.537 1.124 0.877
H9 -0.537 1.124 -0.877
H10 0.478 -1.153 -0.875
H11 0.478 -1.153 0.875
H12 -1.910 -1.079 -0.816
H13 -1.910 -1.079 0.816

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.52412.53553.87081.08871.08991.08992.15752.15752.75792.75794.17114.1711
C21.52411.52672.53752.17342.16602.16601.09291.09292.15012.15012.76932.7693
C32.53551.52671.46163.48612.79342.79342.14522.14521.09171.09172.05142.0514
N43.87082.53751.46164.69754.16984.16982.77952.77952.07132.07131.01351.0135
H51.08872.17343.48614.69751.76121.76122.50352.50353.75963.75964.88484.8848
H61.08992.16602.79344.16981.76121.76142.51333.06673.11202.56944.69154.3745
H71.08992.16602.79344.16981.76121.76143.06672.51332.56943.11204.37454.6915
H82.15751.09292.14522.77952.50352.51333.06671.75323.04642.49243.09932.5970
H92.15751.09292.14522.77952.50353.06672.51331.75322.49243.04642.59703.0993
H102.75792.15011.09172.07133.75963.11202.56943.04642.49241.75002.39052.9276
H112.75792.15011.09172.07133.75962.56943.11202.49243.04641.75002.92762.3905
H124.17112.76932.05141.01354.88484.69154.37453.09932.59702.39052.92761.6321
H134.17112.76932.05141.01354.88484.37454.69152.59703.09932.92762.39051.6321

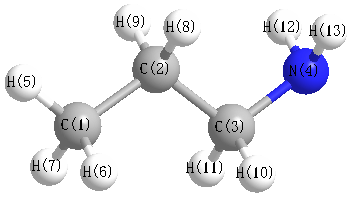
picture of 1-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.423 C1 C2 H8 109.961
C1 C2 H9 109.961 C2 C1 H5 111.482
C2 C1 H6 110.816 C2 C1 H7 110.816
C2 C3 N4 116.221 C2 C3 H10 109.274
C2 C3 H11 109.274 C3 C2 H8 108.828
C3 C2 H9 108.828 C3 N4 H12 110.648
C3 N4 H13 110.648 N4 C3 H10 107.547
N4 C3 H11 107.547 H5 C1 H6 107.882
H5 C1 H7 107.882 H6 C1 H7 107.815
H8 C2 H9 106.661 H10 C3 H11 106.546
H12 N4 H13 107.248
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability