return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H8N2 (Ethylenediamine)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-189.962381
Energy at 298.15K-189.972611
HF Energy-189.294571
Nuclear repulsion energy131.780506
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3578 3361 0.00      
2 Ag 3111 2922 0.00      
3 Ag 1700 1597 0.00      
4 Ag 1527 1434 0.00      
5 Ag 1436 1349 0.00      
6 Ag 1144 1075 0.00      
7 Ag 1095 1028 0.00      
8 Ag 838 787 0.00      
9 Ag 470 441 0.00      
10 Au 3688 3464 6.45      
11 Au 3188 2995 61.90      
12 Au 1420 1333 0.82      
13 Au 1086 1020 0.48      
14 Au 771 725 0.01      
15 Au 260 244 123.09      
16 Au 155 145 0.26      
17 Bg 3688 3464 0.00      
18 Bg 3163 2971 0.00      
19 Bg 1397 1312 0.00      
20 Bg 1332 1251 0.00      
21 Bg 960 901 0.00      
22 Bg 289 271 0.00      
23 Bu 3578 3360 0.74      
24 Bu 3118 2929 76.55      
25 Bu 1699 1596 69.64      
26 Bu 1544 1451 0.92      
27 Bu 1360 1278 20.69      
28 Bu 1115 1048 23.48      
29 Bu 871 818 416.07      
30 Bu 279 262 22.49      

Unscaled Zero Point Vibrational Energy (zpe) 24929.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 23413.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.88495 0.12486 0.11831

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.398 0.654 0.000
C2 -0.398 -0.654 0.000
N3 -0.398 1.878 0.000
N4 0.398 -1.878 0.000
H5 0.995 -1.923 0.817
H6 0.995 -1.923 -0.817
H7 -0.995 1.923 0.817
H8 -0.995 1.923 -0.817
H9 -1.049 -0.676 -0.876
H10 -1.049 -0.676 0.876
H11 1.049 0.676 -0.876
H12 1.049 0.676 0.876

Atom - Atom Distances (Å)
  C1 C2 N3 N4 H5 H6 H7 H8 H9 H10 H11 H12
C11.53091.46002.53252.76862.76862.05362.05362.15162.15161.09191.0919
C21.53092.53251.46002.05362.05362.76862.76861.09191.09192.15162.1516
N31.46002.53253.84024.13014.13011.01331.01332.77782.77782.07532.0753
N42.53251.46003.84021.01331.01334.13014.13012.07532.07532.77782.7778
H52.76862.05364.13011.01331.63454.33014.62842.93272.39533.10212.5999
H62.76862.05364.13011.01331.63454.62844.33012.39532.93272.59993.1021
H72.05362.76861.01334.13014.33014.62841.63453.10212.59992.93272.3953
H82.05362.76861.01334.13014.62844.33011.63452.59993.10212.39532.9327
H92.15161.09192.77782.07532.93272.39533.10212.59991.75192.49603.0495
H102.15161.09192.77782.07532.39532.93272.59993.10211.75193.04952.4960
H111.09192.15162.07532.77783.10212.59992.93272.39532.49603.04951.7519
H121.09192.15162.07532.77782.59993.10212.39532.93273.04952.49601.7519

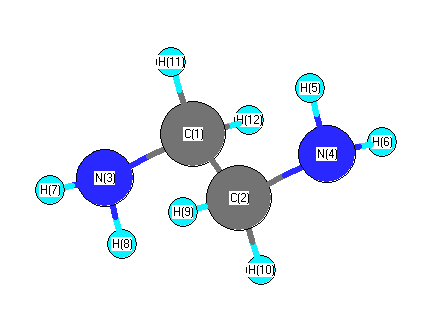
picture of Ethylenediamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N4 115.697 C1 C2 H9 109.088
C1 C2 H10 109.088 C1 N3 H7 110.971
C1 N3 H8 110.971 C2 C1 N3 115.697
C2 C1 H11 109.088 C2 C1 H12 109.088
C2 N4 H5 110.971 C2 N4 H6 110.971
N3 C1 H11 107.956 N3 C1 H12 107.956
N4 C2 H9 107.956 N4 C2 H10 107.956
H5 N4 H6 107.509 H7 N3 H8 107.509
H9 C2 H10 106.688 H11 C1 H12 106.688
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability