return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (2-Propen-1-ol)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-192.562283
Energy at 298.15K-192.568540
HF Energy-191.937885
Nuclear repulsion energy116.605427
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3875 3639 28.95      
2 A 3337 3134 11.89      
3 A 3257 3059 5.43      
4 A 3233 3037 8.39      
5 A 3139 2949 35.85      
6 A 3083 2896 50.28      
7 A 1731 1626 0.44      
8 A 1563 1468 1.41      
9 A 1508 1416 16.07      
10 A 1446 1358 7.94      
11 A 1336 1255 0.12      
12 A 1299 1220 10.87      
13 A 1242 1167 51.04      
14 A 1187 1115 25.95      
15 A 1071 1006 91.69      
16 A 1030 967 32.01      
17 A 985 925 6.80      
18 A 945 888 5.53      
19 A 941 884 38.38      
20 A 664 623 5.59      
21 A 448 421 3.63      
22 A 345 324 12.83      
23 A 264 248 142.35      
24 A 113 106 2.04      

Unscaled Zero Point Vibrational Energy (zpe) 19020.7 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 17864.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.89077 0.14490 0.14019

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.585 0.452 0.311
C2 -0.666 -0.361 0.281
C3 -1.819 0.080 -0.237
O4 1.606 -0.300 -0.361
H5 0.418 1.413 -0.184
H6 0.880 0.644 1.347
H7 -0.600 -1.353 0.712
H8 -2.711 -0.528 -0.227
H9 -1.895 1.063 -0.683
H10 2.456 0.130 -0.203

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.49152.49301.43521.09341.09442.19553.48022.74011.9670
C21.49151.33872.36132.12972.12971.08352.11412.11333.1966
C32.49301.33873.44822.60433.17972.10701.08001.08184.2753
O41.43522.36133.44822.09192.08192.66904.32523.77040.9655
H51.09342.12972.60432.09191.77413.07993.68252.39182.4083
H61.09442.12973.17972.08191.77412.56524.09243.46352.2688
H72.19551.08352.10702.66903.07992.56522.45333.07523.5170
H83.48022.11411.08004.32523.68254.09242.45331.84545.2088
H92.74012.11331.08183.77042.39183.46353.07521.84544.4755
H101.96703.19664.27530.96552.40832.26883.51705.20884.4755

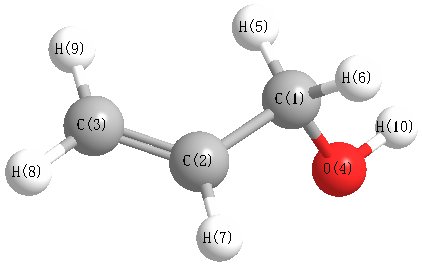
picture of 2-Propen-1-ol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.394 C1 C2 H7 116.090
C1 O4 H10 108.450 C2 C1 O4 107.556
C2 C1 H5 109.998 C2 C1 H6 109.943
C2 C3 H8 121.501 C2 C3 H9 121.271
C3 C2 H7 120.516 O4 C1 H5 110.925
O4 C1 H6 110.053 H5 C1 H6 108.366
H8 C3 H9 117.227
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability