return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H2O3 (Maleic Anhydride)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-378.314435
Energy at 298.15K-378.317601
HF Energy-377.235415
Nuclear repulsion energy273.603012
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3346 3143 1.35      
2 A1 1875 1761 40.11      
3 A1 1653 1552 3.13      
4 A1 1281 1203 144.47      
5 A1 1100 1033 24.36      
6 A1 882 828 8.40      
7 A1 628 590 0.62      
8 A1 397 373 9.62      
9 A2 930 874 0.00      
10 A2 702 660 0.00      
11 A2 239 224 0.00      
12 B1 825 775 77.99      
13 B1 627 589 0.14      
14 B1 157 148 2.27      
15 B2 3326 3124 1.66      
16 B2 1821 1710 529.80      
17 B2 1362 1280 0.06      
18 B2 1088 1022 94.03      
19 B2 905 850 189.32      
20 B2 708 665 42.98      
21 B2 557 523 7.86      

Unscaled Zero Point Vibrational Energy (zpe) 12204.5 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 11462.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.22849 0.08097 0.05978

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 0.976
C2 0.000 1.133 0.156
C3 0.000 -1.133 0.156
O4 0.000 2.257 0.599
O5 0.000 -2.257 0.599
C6 0.000 0.670 -1.257
C7 0.000 -0.670 -1.257
H8 0.000 1.357 -2.087
H9 0.000 -1.357 -2.087

Atom - Atom Distances (Å)
  O1 C2 C3 O4 O5 C6 C7 H8 H9
O11.39891.39892.28792.28792.33102.33103.35033.3503
C21.39892.26611.20773.41841.48592.29052.25383.3511
C31.39892.26613.41841.20772.29051.48593.35112.2538
O42.28791.20773.41844.51302.44073.46542.83234.5023
O52.28793.41841.20774.51303.46542.44074.50232.8323
C62.33101.48592.29052.44073.46541.34101.07762.1911
C72.33102.29051.48593.46542.44071.34102.19111.0776
H83.35032.25383.35112.83234.50231.07762.19112.7141
H93.35033.35112.25384.50232.83232.19111.07762.7141

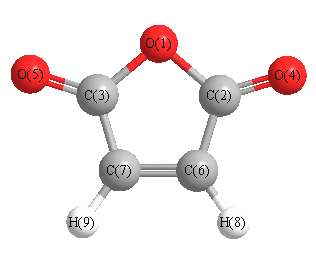
picture of Maleic Anhydride state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 O4 122.568 O1 C2 C6 107.770
O1 C3 O5 122.568 O1 C3 C7 107.770
C2 O1 C3 108.183 C2 C6 C7 108.138
C2 C6 H8 122.285 C3 C7 C6 108.138
C3 C7 H9 122.285 O4 C2 C6 129.662
O5 C3 C7 129.662 C6 C7 H9 129.577
C7 C6 H8 129.577
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability