return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H13N (cyclohexanamine)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-290.338079
Energy at 298.15K-290.354594
HF Energy-289.258313
Nuclear repulsion energy331.155788
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3658 3436 1.16      
2 A 3548 3333 0.75      
3 A 3173 2980 44.26      
4 A 3166 2974 51.49      
5 A 3160 2968 30.65      
6 A 3157 2965 43.52      
7 A 3146 2955 49.90      
8 A 3101 2912 55.82      
9 A 3098 2910 23.65      
10 A 3098 2909 28.35      
11 A 3095 2907 28.52      
12 A 3081 2894 20.79      
13 A 3081 2893 10.18      
14 A 1688 1585 40.31      
15 A 1544 1450 1.52      
16 A 1532 1439 13.08      
17 A 1527 1434 4.68      
18 A 1525 1432 1.45      
19 A 1518 1426 0.37      
20 A 1450 1361 5.03      
21 A 1416 1330 2.12      
22 A 1414 1328 2.06      
23 A 1408 1323 3.78      
24 A 1398 1313 0.11      
25 A 1388 1304 2.07      
26 A 1361 1278 0.54      
27 A 1324 1243 0.89      
28 A 1311 1231 2.25      
29 A 1309 1230 1.31      
30 A 1251 1175 3.37      
31 A 1224 1150 0.96      
32 A 1159 1089 12.33      
33 A 1155 1085 3.88      
34 A 1121 1053 1.42      
35 A 1093 1027 0.41      
36 A 1076 1011 4.48      
37 A 1068 1003 0.07      
38 A 1010 948 1.44      
39 A 961 902 11.29      
40 A 928 872 35.28      
41 A 913 857 8.27      
42 A 879 825 104.25      
43 A 877 823 15.74      
44 A 811 762 0.36      
45 A 808 759 5.79      
46 A 562 528 0.85      
47 A 469 441 1.70      
48 A 456 428 2.56      
49 A 412 387 0.16      
50 A 347 326 12.24      
51 A 342 321 2.00      
52 A 256 240 33.74      
53 A 229 215 12.43      
54 A 164 154 0.49      

Unscaled Zero Point Vibrational Energy (zpe) 42121.8 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 39560.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.14321 0.07396 0.05378

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.872 0.009 0.292
C2 1.174 -1.249 -0.219
C3 -0.306 -1.258 0.159
C4 -1.024 -0.007 -0.331
C5 -0.315 1.249 0.181
C6 1.163 1.265 -0.207
N7 -2.441 -0.097 0.030
H8 2.918 0.015 -0.019
H9 1.866 0.004 1.387
H10 1.265 -1.293 -1.309
H11 1.665 -2.143 0.172
H12 -0.810 -2.137 -0.244
H13 -0.400 -1.306 1.251
H14 -0.984 0.001 -1.426
H15 -0.407 1.274 1.273
H16 -0.823 2.138 -0.201
H17 1.647 2.160 0.189
H18 1.248 1.318 -1.297
H19 -2.944 0.723 -0.291
H20 -2.538 -0.126 1.041

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.52672.52272.96242.51591.52654.32181.09191.09452.15042.16463.47612.79473.33332.78523.46972.16582.15124.90374.4746
C21.52671.52732.52762.93522.51483.80212.16362.15131.09441.09172.17352.15452.77113.33093.93223.46652.78554.56704.0774
C32.52271.52731.52402.50672.94272.43373.47092.79572.15052.16001.09081.09662.13542.76843.45393.93703.34253.33052.6533
C42.96242.52761.52401.53042.53351.46473.95523.36222.80213.47082.14322.14011.09602.14382.15803.47902.80232.05472.0463
C52.51592.93522.50671.53041.52792.52083.46632.78503.34293.92663.44832.77082.14251.09611.09342.16322.15232.72302.7515
C61.52652.51482.94272.53351.52793.86012.16362.15022.78733.46553.93353.34382.77462.15772.16951.09181.09464.14394.1459
N74.32183.80212.43371.46472.52083.86015.36074.51654.11804.58922.62652.66742.06212.74992.76864.67244.16821.01521.0159
H81.09192.16363.47093.95523.46632.16365.36071.75572.47122.50244.31063.79084.14903.78354.30592.50282.47425.91185.5598
H91.09452.15132.79573.36222.78502.15024.51651.75573.05092.47473.79532.62144.00472.60643.78212.47593.05105.14524.4191
H102.15041.09442.15052.80213.34292.78734.11802.47123.05091.75352.48053.05372.59784.00674.16643.78352.61114.77724.6200
H112.16461.09172.16003.47083.92663.46554.58922.50242.47471.75352.50922.47543.76414.14494.96514.30313.78265.44744.7414
H123.47612.17351.09082.14323.44833.93352.62654.31063.79532.48052.50921.75852.44973.75554.27554.96934.15753.56992.9465
H132.79472.15451.09662.14012.77083.34382.66743.79082.62143.05372.47541.75853.03582.58053.76124.16354.01183.60142.4504
H143.33332.77112.13541.09602.14252.77462.06214.14904.00472.59783.76412.44973.03583.04022.46883.76812.59562.37732.9184
H152.78523.33092.76842.14381.09612.15772.74993.78352.60644.00674.14493.75552.58053.04021.75832.48573.05733.03142.5602
H163.46973.93223.45392.15801.09342.16952.76864.30593.78214.16644.96514.27553.76122.46881.75832.50092.48282.55103.0996
H172.16583.46653.93703.47902.16321.09184.67242.50282.47593.78354.30314.96934.16353.76812.48572.50091.75414.83484.8440
H182.15122.78553.34252.80232.15231.09464.16822.47423.05102.61113.78264.15754.01182.59563.05732.48281.75414.35234.6781
H194.90374.56703.33052.05472.72304.14391.01525.91185.14524.77725.44743.56993.60142.37733.03142.55104.83484.35231.6320
H204.47464.07742.65332.04632.75154.14591.01595.55984.41914.62004.74142.94652.45042.91842.56023.09964.84404.67811.6320

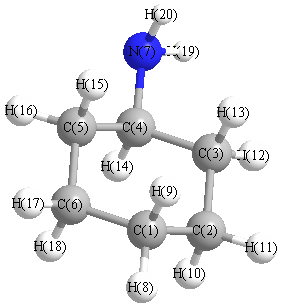
picture of cyclohexanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.383 C1 C2 H10 109.143
C1 C2 H11 110.414 C1 C6 C5 110.910
C1 C6 H17 110.516 C1 C6 H18 109.204
C2 C1 C6 110.902 C2 C1 H8 110.329
C2 C1 H9 109.206 C2 C3 C4 111.863
C2 C3 H12 111.140 C2 C3 H13 109.289
C3 C2 H10 109.103 C3 C2 H11 110.009
C3 C4 C5 110.309 C3 C4 N7 109.024
C3 C4 H14 108.069 C4 C3 H12 108.973
C4 C3 H13 108.399 C4 C5 C6 111.867
C4 C5 H15 108.286 C4 C5 H16 109.538
C4 N7 H19 110.587 C4 N7 H20 109.839
C5 C4 N7 114.610 C5 C4 H14 108.183
C5 C6 H17 110.211 C5 C6 H18 109.197
C6 C1 H8 110.337 C6 C1 H9 109.133
C6 C5 H15 109.533 C6 C5 H16 110.614
N7 C4 H14 106.388 H8 C1 H9 106.834
H10 C2 H11 106.661 H12 C3 H13 107.021
H15 C5 H16 106.849 H17 C6 H18 106.698
H19 N7 H20 106.931
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability