return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H13N (2-Methylpiperidine)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-290.329932
Energy at 298.15K-290.346467
HF Energy-289.250646
Nuclear repulsion energy333.383763
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3551 3335 0.21      
2 A 3225 3029 18.25      
3 A 3200 3006 35.66      
4 A 3174 2981 43.90      
5 A 3163 2971 45.75      
6 A 3155 2963 44.77      
7 A 3148 2957 41.25      
8 A 3109 2920 29.60      
9 A 3102 2914 40.15      
10 A 3098 2910 27.45      
11 A 3095 2907 30.28      
12 A 3087 2899 35.12      
13 A 3078 2891 11.17      
14 A 1544 1450 2.78      
15 A 1540 1446 8.91      
16 A 1538 1445 5.80      
17 A 1526 1433 9.95      
18 A 1523 1430 3.39      
19 A 1519 1427 0.89      
20 A 1497 1406 21.26      
21 A 1449 1361 10.85      
22 A 1420 1334 1.00      
23 A 1419 1333 8.34      
24 A 1410 1324 0.11      
25 A 1397 1312 0.71      
26 A 1386 1302 2.66      
27 A 1359 1276 0.42      
28 A 1329 1249 3.25      
29 A 1310 1230 5.55      
30 A 1274 1197 6.65      
31 A 1224 1150 3.08      
32 A 1206 1133 9.61      
33 A 1174 1103 21.51      
34 A 1143 1073 2.70      
35 A 1126 1057 3.07      
36 A 1096 1030 5.19      
37 A 1018 956 4.23      
38 A 1009 948 0.36      
39 A 989 929 1.77      
40 A 946 889 10.21      
41 A 908 853 1.73      
42 A 881 828 0.71      
43 A 865 812 36.18      
44 A 828 777 11.71      
45 A 768 721 75.19      
46 A 573 538 2.06      
47 A 485 456 6.49      
48 A 461 433 1.22      
49 A 431 405 1.17      
50 A 344 323 0.74      
51 A 322 302 0.97      
52 A 255 240 2.47      
53 A 232 218 1.22      
54 A 156 147 0.98      

Unscaled Zero Point Vibrational Energy (zpe) 42032.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 39476.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.14478 0.07601 0.05502

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.865 -0.056 0.297
C2 -1.216 1.233 -0.200
C3 0.267 1.259 0.167
C4 0.986 0.005 -0.334
N5 0.339 -1.236 0.106
C6 -1.089 -1.269 -0.212
C7 2.446 -0.021 0.082
H8 -2.907 -0.114 -0.024
H9 -1.864 -0.066 1.392
H10 -1.316 1.288 -1.288
H11 -1.727 2.106 0.209
H12 0.748 2.151 -0.242
H13 0.374 1.307 1.257
H14 0.931 -0.001 -1.429
H15 0.464 -1.333 1.112
H16 -1.504 -2.195 0.189
H17 -1.176 -1.324 -1.301
H18 2.935 -0.912 -0.307
H19 2.529 -0.028 1.170
H20 2.969 0.861 -0.287

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.52612.50782.91992.50721.52744.31611.09271.09452.14992.16793.46272.79093.28682.77862.17202.15334.91254.47934.9544
C21.52611.52722.52402.93332.50523.88022.16952.15401.09461.09162.16852.15722.76483.33603.46182.78384.67294.18174.2018
C32.50781.52721.53032.49682.89412.52883.46372.79212.15062.16651.09331.09602.14002.76683.88183.30283.47242.78942.7690
C42.91992.52401.53031.46652.43781.51853.90703.33242.80303.47402.16152.14491.09682.03823.36322.71522.15422.15522.1608
N52.50722.93332.49681.46651.46392.43203.43742.80663.32533.93033.42942.79112.05771.01842.07972.07002.64802.71713.3862
C61.52742.50522.89412.43781.46393.76062.16222.14982.78423.46063.88303.30682.67852.04241.09071.09414.04104.06684.5839
C74.31613.88022.52881.51852.43203.76065.35514.50514.21254.68542.77572.72722.13942.59074.51014.08991.08811.09171.0895
H81.09272.16953.46373.90703.43742.16225.35511.75952.46892.52504.30623.79834.08943.76132.51892.46805.90315.56635.9624
H91.09452.15402.79213.33242.80662.14984.50511.75953.05232.47633.79602.62893.97242.66612.47223.05105.16064.39865.1997
H102.14991.09462.15062.80303.32532.78424.21252.46893.05231.75502.47023.05532.59523.97593.78832.61584.88644.75004.4214
H112.16791.09162.16653.47403.93033.46064.68542.52502.47631.75502.51672.47993.76744.17704.30673.78775.57744.85704.8836
H123.46272.16851.09332.16153.42943.88302.77574.30623.79602.47022.51671.76082.46493.74964.91424.11093.76413.14902.5687
H132.79092.15721.09602.14492.79113.30682.72723.79832.62893.05532.47991.76083.03942.64594.11483.98313.73192.53653.0523
H143.28682.76482.14001.09682.05772.67852.13944.08943.97242.59523.76742.46493.03942.90763.65572.49142.47053.05122.4899
H152.77863.33602.76682.03821.01842.04242.59073.76132.66613.97594.17703.74962.64592.90762.33882.91822.88002.44293.6120
H162.17203.46183.88183.36322.07971.09074.51012.51892.47223.78834.30674.91424.11483.65572.33881.75704.64714.68195.4382
H172.15332.78383.30282.71522.07001.09414.08992.46803.05102.61583.78774.11093.98312.49142.91821.75704.24934.63784.7939
H184.91254.67293.47242.15422.64804.04101.08815.90315.16064.88645.57743.76413.73192.47052.88004.64714.24931.76831.7734
H194.47934.18172.78942.15522.71714.06681.09175.56634.39864.75004.85703.14902.53653.05122.44294.68194.63781.76831.7631
H204.95444.20182.76902.16083.38624.58391.08955.96245.19974.42144.88362.56873.05232.48993.61205.43824.79391.77341.7631

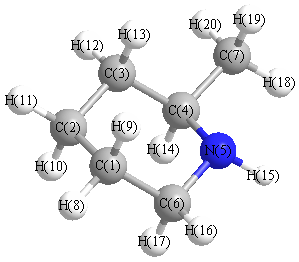
picture of 2-Methylpiperidine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 110.441 C1 C2 H10 109.128
C1 C2 H11 110.731 C1 C6 N5 113.873
C1 C6 H16 111.020 C1 C6 H17 109.340
C2 C1 C6 110.258 C2 C1 H8 110.789
C2 C1 H9 109.452 C2 C3 C4 111.285
C2 C3 H12 110.597 C2 C3 H13 109.546
C3 C2 H10 109.113 C3 C2 H11 110.546
C3 C4 N5 112.838 C3 C4 C7 112.084
C3 C4 H14 107.957 C4 C3 H12 109.834
C4 C3 H13 108.379 C4 N5 C6 112.590
C4 N5 H15 108.886 C4 C7 H18 110.378
C4 C7 H19 110.237 C4 C7 H20 110.822
N5 C4 C7 109.110 N5 C4 H14 105.883
N5 C6 H16 108.102 N5 C6 H17 107.153
C6 C1 H8 110.119 C6 C1 H9 109.039
C6 N5 H15 109.416 C7 C4 H14 108.703
H8 C1 H9 107.114 H10 C2 H11 106.790
H12 C3 H13 107.082 H16 C6 H17 107.065
H18 C7 H19 108.423 H18 C7 H20 109.050
H19 C7 H20 107.857
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability