return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12 (Pentane)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-197.122089
Energy at 298.15K-197.135044
HF Energy-196.352644
Nuclear repulsion energy185.445773
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3210 3015 42.16      
2 A1 3113 2924 19.85      
3 A1 3109 2920 84.56      
4 A1 3089 2901 10.53      
5 A1 1556 1461 8.47      
6 A1 1542 1449 0.06      
7 A1 1531 1438 0.02      
8 A1 1459 1370 4.18      
9 A1 1403 1317 0.14      
10 A1 1198 1125 0.60      
11 A1 1090 1024 0.91      
12 A1 902 847 0.68      
13 A1 409 384 0.00      
14 A1 181 170 0.00      
15 A2 3207 3012 0.00      
16 A2 3156 2964 0.00      
17 A2 1547 1453 0.00      
18 A2 1355 1273 0.00      
19 A2 1300 1221 0.00      
20 A2 1014 952 0.00      
21 A2 785 737 0.00      
22 A2 262 246 0.00      
23 A2 105 99 0.00      
24 B1 3208 3013 95.33      
25 B1 3168 2976 54.46      
26 B1 3135 2945 1.20      
27 B1 1546 1452 15.93      
28 B1 1354 1272 1.37      
29 B1 1245 1170 0.00      
30 B1 888 834 0.65      
31 B1 747 701 2.22      
32 B1 251 235 0.00      
33 B1 117 110 0.00      
34 B2 3210 3015 27.65      
35 B2 3112 2923 46.76      
36 B2 3104 2915 0.04      
37 B2 1550 1456 2.51      
38 B2 1534 1441 0.71      
39 B2 1455 1367 2.40      
40 B2 1445 1358 1.26      
41 B2 1317 1237 1.60      
42 B2 1122 1054 0.52      
43 B2 1074 1009 1.24      
44 B2 959 901 2.39      
45 B2 405 381 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 36234.7 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 34031.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.57133 0.06520 0.06186

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.311
C2 0.000 1.275 -0.524
C3 0.000 -1.275 -0.524
C4 0.000 2.539 0.326
C5 0.000 -2.539 0.326
H6 0.876 0.000 0.967
H7 -0.876 0.000 0.967
H8 0.876 1.273 -1.178
H9 -0.876 1.273 -1.178
H10 -0.876 -1.273 -1.178
H11 0.876 -1.273 -1.178
H12 0.000 3.436 -0.291
H13 0.000 -3.436 -0.291
H14 -0.881 2.573 0.968
H15 0.881 2.573 0.968
H16 0.881 -2.573 0.968
H17 -0.881 -2.573 0.968

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.52411.52412.53932.53931.09441.09442.14572.14572.14572.14573.48883.48882.79802.79802.79802.7980
C21.52412.54951.52413.90772.14852.14851.09291.09292.77192.77192.17434.71702.16532.16534.22014.2201
C31.52412.54953.90771.52412.14852.14852.77192.77191.09291.09294.71702.17434.22014.22012.16532.1653
C42.53931.52413.90775.07852.76152.76152.15302.15304.19064.19061.08926.00761.08991.08995.22735.2273
C52.53933.90771.52415.07852.76152.76154.19064.19062.15302.15306.00761.08925.22735.22731.08991.0899
H61.09442.14852.14852.76152.76151.75282.49393.04783.04782.49393.76293.76293.11592.57332.57333.1159
H71.09442.14852.14852.76152.76151.75283.04782.49392.49393.04783.76293.76292.57333.11593.11592.5733
H82.14571.09292.77192.15304.19062.49393.04781.75143.08952.54522.49734.87123.06302.50944.40414.7414
H92.14571.09292.77192.15304.19063.04782.49391.75142.54523.08952.49734.87122.50943.06304.74144.4041
H102.14572.77191.09294.19062.15303.04782.49393.08952.54521.75144.87122.49734.40414.74143.06302.5094
H112.14572.77191.09294.19062.15302.49393.04782.54523.08951.75144.87122.49734.74144.40412.50943.0630
H123.48882.17434.71701.08926.00763.76293.76292.49732.49734.87124.87126.87301.76221.76226.20316.2031
H133.48884.71702.17436.00761.08923.76293.76294.87124.87122.49732.49736.87306.20316.20311.76221.7622
H142.79802.16534.22011.08995.22733.11592.57333.06302.50944.40414.74141.76226.20311.76125.43965.1466
H152.79802.16534.22011.08995.22732.57333.11592.50943.06304.74144.40411.76226.20311.76125.14665.4396
H162.79804.22012.16535.22731.08992.57333.11594.40414.74143.06302.50946.20311.76225.43965.14661.7612
H172.79804.22012.16535.22731.08993.11592.57334.74144.40412.50943.06306.20311.76225.14665.43961.7612

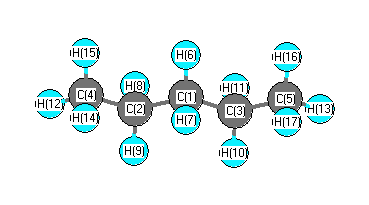
picture of Pentane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 112.825 C1 C2 H8 109.043
C1 C2 H9 109.043 C1 C3 C5 112.825
C1 C3 H10 109.043 C1 C3 H11 109.043
C2 C1 C3 113.519 C2 C1 H6 109.167
C2 C1 H7 109.167 C2 C4 H12 111.527
C2 C4 H14 110.759 C2 C4 H15 110.759
C3 C1 H6 109.167 C3 C1 H7 109.167
C3 C5 H13 111.527 C3 C5 H16 110.759
C3 C5 H17 110.759 C4 C2 H8 109.611
C4 C2 H9 109.611 C5 C3 H10 109.611
C5 C3 H11 109.611 H6 C1 H7 106.407
H8 C2 H9 106.508 H10 C3 H11 106.508
H12 C4 H14 107.932 H12 C4 H15 107.932
H13 C5 H16 107.932 H13 C5 H17 107.932
H14 C4 H15 107.787 H16 C5 H17 107.787
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability