return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(NH2)H2CH2CH2CH3 (1-Butanamine)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-213.136408
Energy at 298.15K-213.149110
Nuclear repulsion energy185.909376
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3573 3356 0.60      
2 A' 3211 3016 33.99      
3 A' 3119 2929 60.23      
4 A' 3114 2925 33.23      
5 A' 3107 2918 22.00      
6 A' 3089 2902 15.10      
7 A' 1698 1595 33.68      
8 A' 1554 1459 5.80      
9 A' 1545 1451 0.72      
10 A' 1535 1442 0.32      
11 A' 1528 1435 0.17      
12 A' 1458 1369 2.54      
13 A' 1442 1354 9.74      
14 A' 1411 1325 4.36      
15 A' 1314 1234 2.84      
16 A' 1173 1102 4.87      
17 A' 1127 1059 18.55      
18 A' 1106 1039 1.01      
19 A' 1050 986 14.97      
20 A' 938 881 17.82      
21 A' 867 814 181.32      
22 A' 437 411 6.23      
23 A' 400 376 0.49      
24 A' 183 172 2.92      
25 A" 3682 3458 2.33      
26 A" 3209 3014 47.83      
27 A" 3182 2988 55.45      
28 A" 3161 2969 12.23      
29 A" 3138 2947 2.91      
30 A" 1547 1453 8.05      
31 A" 1413 1327 0.72      
32 A" 1357 1274 0.28      
33 A" 1337 1256 0.23      
34 A" 1264 1187 0.01      
35 A" 1062 998 0.58      
36 A" 940 883 0.06      
37 A" 805 756 0.55      
38 A" 751 705 1.47      
39 A" 278 261 55.26      
40 A" 258 242 0.88      
41 A" 125 117 0.01      
42 A" 106 100 3.69      

Unscaled Zero Point Vibrational Energy (zpe) 33796.3 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 31741.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.60222 0.06505 0.06196

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 2.536 0.384 0.000
C2 1.324 -0.433 0.000
C3 0.000 0.328 0.000
C4 -1.218 -0.589 0.000
C5 -2.535 0.178 0.000
H6 2.560 0.985 0.816
H7 2.560 0.985 -0.816
H8 1.366 -1.084 0.875
H9 1.366 -1.084 -0.875
H10 -0.035 0.981 -0.877
H11 -0.035 0.981 0.877
H12 -1.174 -1.241 0.876
H13 -1.174 -1.241 -0.876
H14 -3.390 -0.498 0.000
H15 -2.611 0.815 0.881
H16 -2.611 0.815 -0.881

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.46242.53693.87815.07561.01371.01372.07162.07162.78152.78154.14424.14425.99115.23995.2399
C21.46241.52682.54613.90672.05102.05101.09161.09162.14832.14832.76722.76724.71364.22084.2208
C32.53691.52681.52442.53962.76622.76622.15092.15091.09441.09442.14652.14653.48862.79832.7983
C43.87812.54611.52441.52424.17334.17332.77262.77262.15272.15271.09281.09282.17372.16532.1653
C55.07563.90672.53961.52425.22295.22294.19264.19262.76892.76892.15252.15251.08911.08981.0898
H61.01372.05102.76624.17335.22291.63142.39022.92723.09852.59574.34804.66506.18575.17445.4449
H71.01372.05102.76624.17335.22291.63142.92722.39022.59573.09854.66504.34806.18575.44495.1744
H82.07161.09162.15092.77264.19262.39022.92721.75033.04952.49562.54493.08894.87094.40754.7443
H92.07161.09162.15092.77264.19262.92722.39021.75032.49563.04953.08892.54494.87094.74434.4075
H102.78152.14831.09442.15272.76893.09852.59573.04952.49561.75503.05122.49733.76973.12342.5816
H112.78152.14831.09442.15272.76892.59573.09852.49563.04951.75502.49733.05123.76972.58163.1234
H124.14422.76722.14651.09282.15254.34804.66502.54493.08893.05122.49731.75122.49572.50903.0626
H134.14422.76722.14651.09282.15254.66504.34803.08892.54492.49733.05121.75122.49573.06262.5090
H145.99114.71363.48862.17371.08916.18576.18574.87094.87093.76973.76972.49572.49571.76211.7621
H155.23994.22082.79832.16531.08985.17445.44494.40754.74433.12342.58162.50903.06261.76211.7613
H165.23994.22082.79832.16531.08985.44495.17444.74434.40752.58163.12343.06262.50901.76211.7613

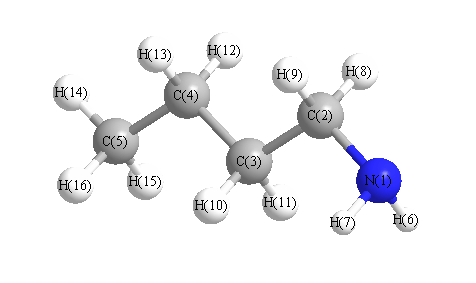
picture of 1-Butanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 116.123 N1 C2 H8 107.522
N1 C2 H9 107.522 C2 N1 H6 110.548
C2 N1 H7 110.548 C2 C3 C4 113.122
C2 C3 H10 108.970 C2 C3 H11 108.970
C3 C2 H8 109.335 C3 C2 H9 109.335
C3 C4 C5 112.824 C3 C4 H12 109.090
C3 C4 H13 109.090 C4 C3 H10 109.481
C4 C3 H11 109.481 C4 C5 H14 111.478
C4 C5 H15 110.761 C4 C5 H16 110.761
C5 C4 H12 109.571 C5 C4 H13 109.571
H6 N1 H7 107.163 H8 C2 H9 106.581
H10 C3 H11 106.607 H12 C4 H13 106.495
H14 C5 H15 107.943 H14 C5 H16 107.943
H15 C5 H16 107.812
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability