return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2CN (Butanenitrile)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-210.759783
Energy at 298.15K-210.767037
HF Energy-210.007754
Nuclear repulsion energy153.875839
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3222 3026 22.59      
2 A' 3140 2949 23.60      
3 A' 3129 2939 0.73      
4 A' 3119 2930 21.58      
5 A' 2209 2074 0.09      
6 A' 1552 1458 7.40      
7 A' 1540 1446 1.47      
8 A' 1518 1425 3.14      
9 A' 1462 1373 3.11      
10 A' 1428 1341 1.74      
11 A' 1334 1253 1.80      
12 A' 1147 1078 2.24      
13 A' 1093 1026 0.15      
14 A' 980 920 1.77      
15 A' 903 848 1.73      
16 A' 517 486 0.89      
17 A' 349 328 0.03      
18 A' 161 151 5.93      
19 A" 3217 3022 41.66      
20 A" 3198 3004 1.23      
21 A" 3181 2987 0.00      
22 A" 1547 1453 9.15      
23 A" 1356 1274 0.08      
24 A" 1291 1212 0.08      
25 A" 1155 1085 0.00      
26 A" 896 841 0.19      
27 A" 760 713 2.03      
28 A" 357 335 0.15      
29 A" 246 231 0.00      
30 A" 97 91 4.26      

Unscaled Zero Point Vibrational Energy (zpe) 23050.6 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 21649.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.79706 0.07524 0.07147

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.626 0.327 0.000
C2 -1.456 0.479 0.000
C3 0.000 0.635 0.000
C4 0.730 -0.711 0.000
C5 2.241 -0.525 0.000
H6 0.283 1.218 0.878
H7 0.283 1.218 -0.878
H8 0.421 -1.281 0.876
H9 0.421 -1.281 -0.876
H10 2.750 -1.487 0.000
H11 2.566 0.026 0.882
H12 2.566 0.026 -0.882

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.18062.64453.51364.94193.16693.16693.55523.55525.67435.27595.2759
C21.18061.46402.48903.83102.08312.08312.71802.71804.64264.14254.1425
C32.64451.46401.53192.52401.09101.09102.14902.14903.47382.78132.7813
C43.51362.48901.53191.52272.16612.16611.09011.09012.16372.16652.1665
C54.94193.83102.52401.52272.76462.76462.15762.15761.08821.08961.0896
H63.16692.08311.09102.16612.76461.75622.50283.05633.76462.57563.1195
H73.16692.08311.09102.16612.76461.75623.05632.50283.76463.11952.5756
H83.55522.71802.14901.09012.15762.50283.05631.75232.49732.51293.0668
H93.55522.71802.14901.09012.15763.05632.50281.75232.49733.06682.5129
H105.67434.64263.47382.16371.08823.76463.76462.49732.49731.76101.7610
H115.27594.14252.78132.16651.08962.57563.11952.51293.06681.76101.7638
H125.27594.14252.78132.16651.08963.11952.57563.06682.51291.76101.7638

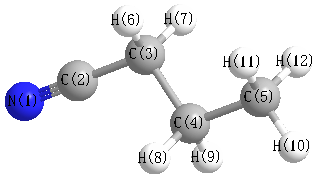
picture of Butanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 178.725 C2 C3 C4 112.343
C2 C3 H6 108.355 C2 C3 H7 108.355
C3 C4 C5 111.434 C3 C4 H8 108.925
C3 C4 H9 108.925 C4 C3 H6 110.214
C4 C3 H7 110.214 C4 C5 H10 110.832
C4 C5 H11 110.978 C4 C5 H12 110.978
C5 C4 H8 110.234 C5 C4 H9 110.234
H6 C3 H7 107.198 H8 C4 H9 106.970
H10 C5 H11 107.924 H10 C5 H12 107.924
H11 C5 H12 108.073
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability