Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -268.802881 |
| Energy at 298.15K | -268.812571 |
| Nuclear repulsion energy | 198.467300 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3254 |
3056 |
3.84 |
|
|
|
| 2 |
A |
3192 |
2997 |
57.50 |
|
|
|
| 3 |
A |
3104 |
2916 |
45.43 |
|
|
|
| 4 |
A |
3098 |
2910 |
27.73 |
|
|
|
| 5 |
A |
1567 |
1472 |
5.35 |
|
|
|
| 6 |
A |
1550 |
1456 |
0.14 |
|
|
|
| 7 |
A |
1537 |
1443 |
4.63 |
|
|
|
| 8 |
A |
1506 |
1414 |
0.40 |
|
|
|
| 9 |
A |
1356 |
1274 |
0.50 |
|
|
|
| 10 |
A |
1236 |
1160 |
37.79 |
|
|
|
| 11 |
A |
1202 |
1129 |
15.08 |
|
|
|
| 12 |
A |
1163 |
1092 |
74.88 |
|
|
|
| 13 |
A |
944 |
886 |
2.24 |
|
|
|
| 14 |
A |
610 |
573 |
6.07 |
|
|
|
| 15 |
A |
321 |
302 |
5.88 |
|
|
|
| 16 |
A |
169 |
158 |
0.43 |
|
|
|
| 17 |
A |
98 |
92 |
2.59 |
|
|
|
| 18 |
B |
3253 |
3056 |
33.85 |
|
|
|
| 19 |
B |
3192 |
2998 |
7.30 |
|
|
|
| 20 |
B |
3180 |
2986 |
41.49 |
|
|
|
| 21 |
B |
3099 |
2911 |
96.95 |
|
|
|
| 22 |
B |
1559 |
1464 |
7.12 |
|
|
|
| 23 |
B |
1538 |
1444 |
3.61 |
|
|
|
| 24 |
B |
1518 |
1425 |
5.29 |
|
|
|
| 25 |
B |
1451 |
1362 |
16.98 |
|
|
|
| 26 |
B |
1272 |
1195 |
15.63 |
|
|
|
| 27 |
B |
1201 |
1128 |
1.27 |
|
|
|
| 28 |
B |
1189 |
1117 |
200.33 |
|
|
|
| 29 |
B |
1094 |
1028 |
136.17 |
|
|
|
| 30 |
B |
961 |
903 |
77.99 |
|
|
|
| 31 |
B |
460 |
432 |
5.60 |
|
|
|
| 32 |
B |
238 |
224 |
21.15 |
|
|
|
| 33 |
B |
134 |
126 |
3.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25621.9 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 24064.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.945 |
| H2 |
-0.710 |
0.556 |
1.561 |
| H3 |
0.710 |
-0.556 |
1.561 |
| O4 |
0.789 |
0.873 |
0.174 |
| O5 |
-0.789 |
-0.873 |
0.174 |
| C6 |
0.000 |
1.796 |
-0.579 |
| C7 |
0.000 |
-1.796 |
-0.579 |
| H8 |
0.700 |
2.447 |
-1.092 |
| H9 |
-0.700 |
-2.447 |
-1.092 |
| H10 |
-0.627 |
1.279 |
-1.304 |
| H11 |
-0.633 |
2.389 |
0.085 |
| H12 |
0.627 |
-1.279 |
-1.304 |
| H13 |
0.633 |
-2.389 |
0.085 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
O4 |
O5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.0922 | 1.0922 | 1.4068 | 1.4068 | 2.3549 | 2.3549 | 3.2594 | 3.2594 | 2.6620 | 2.6163 | 2.6620 | 2.6163 |
H2 | 1.0922 | | 1.8037 | 2.0669 | 1.9932 | 2.5729 | 3.2576 | 3.5497 | 4.0064 | 2.9560 | 2.3542 | 3.6555 | 3.5570 | H3 | 1.0922 | 1.8037 | | 1.9932 | 2.0669 | 3.2576 | 2.5729 | 4.0064 | 3.5497 | 3.6555 | 3.5570 | 2.9560 | 2.3542 | O4 | 1.4068 | 2.0669 | 1.9932 | | 2.3530 | 1.4280 | 2.8826 | 2.0211 | 3.8518 | 2.0863 | 2.0802 | 2.6152 | 3.2664 | O5 | 1.4068 | 1.9932 | 2.0669 | 2.3530 | | 2.8826 | 1.4280 | 3.8518 | 2.0211 | 2.6152 | 3.2664 | 2.0863 | 2.0802 | C6 | 2.3549 | 2.5729 | 3.2576 | 1.4280 | 2.8826 | | 3.5911 | 1.0849 | 4.3299 | 1.0893 | 1.0927 | 3.2202 | 4.2836 | C7 | 2.3549 | 3.2576 | 2.5729 | 2.8826 | 1.4280 | 3.5911 | | 4.3299 | 1.0849 | 3.2202 | 4.2836 | 1.0893 | 1.0927 | H8 | 3.2594 | 3.5497 | 4.0064 | 2.0211 | 3.8518 | 1.0849 | 4.3299 | | 5.0894 | 1.7806 | 1.7797 | 3.7319 | 4.9768 | H9 | 3.2594 | 4.0064 | 3.5497 | 3.8518 | 2.0211 | 4.3299 | 1.0849 | 5.0894 | | 3.7319 | 4.9768 | 1.7806 | 1.7797 | H10 | 2.6620 | 2.9560 | 3.6555 | 2.0863 | 2.6152 | 1.0893 | 3.2202 | 1.7806 | 3.7319 | | 1.7784 | 2.8483 | 4.1192 | H11 | 2.6163 | 2.3542 | 3.5570 | 2.0802 | 3.2664 | 1.0927 | 4.2836 | 1.7797 | 4.9768 | 1.7784 | | 4.1192 | 4.9423 | H12 | 2.6620 | 3.6555 | 2.9560 | 2.6152 | 2.0863 | 3.2202 | 1.0893 | 3.7319 | 1.7806 | 2.8483 | 4.1192 | | 1.7784 | H13 | 2.6163 | 3.5570 | 2.3542 | 3.2664 | 2.0802 | 4.2836 | 1.0927 | 4.9768 | 1.7797 | 4.1192 | 4.9423 | 1.7784 | |
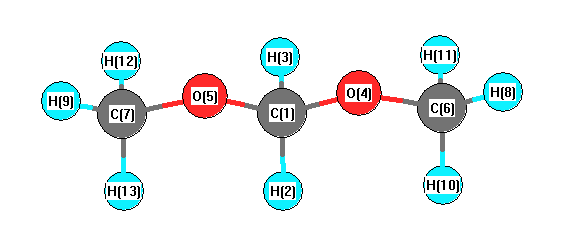 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C6 |
112.342 |
|
C1 |
O5 |
C7 |
112.342 |
| H2 |
C1 |
H3 |
111.327 |
|
H2 |
C1 |
O4 |
110.977 |
| H2 |
C1 |
O5 |
105.102 |
|
H3 |
C1 |
O4 |
105.102 |
| H3 |
C1 |
O5 |
110.977 |
|
O4 |
C1 |
O5 |
113.500 |
| O4 |
C6 |
H8 |
106.283 |
|
O4 |
C6 |
H10 |
111.238 |
| O4 |
C6 |
H11 |
110.518 |
|
O5 |
C7 |
H9 |
106.283 |
| O5 |
C7 |
H12 |
111.238 |
|
O5 |
C7 |
H13 |
110.518 |
| H8 |
C6 |
H10 |
109.963 |
|
H8 |
C6 |
H11 |
109.623 |
| H9 |
C7 |
H12 |
109.963 |
|
H9 |
C7 |
H13 |
109.623 |
| H10 |
C6 |
H11 |
109.178 |
|
H12 |
C7 |
H13 |
109.178 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -268.791266 |
| Energy at 298.15K | -268.800639 |
| Nuclear repulsion energy | 192.891522 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3255 |
3057 |
16.64 |
|
|
|
| 2 |
A1 |
3082 |
2895 |
61.34 |
|
|
|
| 3 |
A1 |
2988 |
2806 |
68.31 |
|
|
|
| 4 |
A1 |
1597 |
1500 |
1.58 |
|
|
|
| 5 |
A1 |
1559 |
1464 |
3.39 |
|
|
|
| 6 |
A1 |
1514 |
1422 |
0.04 |
|
|
|
| 7 |
A1 |
1291 |
1212 |
14.55 |
|
|
|
| 8 |
A1 |
1174 |
1102 |
6.37 |
|
|
|
| 9 |
A1 |
1054 |
990 |
58.78 |
|
|
|
| 10 |
A1 |
480 |
451 |
2.32 |
|
|
|
| 11 |
A1 |
220 |
206 |
2.25 |
|
|
|
| 12 |
A2 |
3157 |
2965 |
0.00 |
|
|
|
| 13 |
A2 |
1539 |
1445 |
0.00 |
|
|
|
| 14 |
A2 |
1270 |
1193 |
0.00 |
|
|
|
| 15 |
A2 |
1197 |
1124 |
0.00 |
|
|
|
| 16 |
A2 |
221 |
208 |
0.00 |
|
|
|
| 17 |
A2 |
59 |
55 |
0.00 |
|
|
|
| 18 |
B1 |
3157 |
2965 |
99.87 |
|
|
|
| 19 |
B1 |
3031 |
2846 |
105.03 |
|
|
|
| 20 |
B1 |
1538 |
1445 |
14.04 |
|
|
|
| 21 |
B1 |
1216 |
1142 |
23.51 |
|
|
|
| 22 |
B1 |
1168 |
1097 |
4.73 |
|
|
|
| 23 |
B1 |
218 |
204 |
7.51 |
|
|
|
| 24 |
B1 |
81 |
76 |
0.34 |
|
|
|
| 25 |
B2 |
3255 |
3057 |
19.20 |
|
|
|
| 26 |
B2 |
3081 |
2893 |
64.83 |
|
|
|
| 27 |
B2 |
1562 |
1467 |
5.40 |
|
|
|
| 28 |
B2 |
1526 |
1433 |
12.77 |
|
|
|
| 29 |
B2 |
1472 |
1382 |
80.83 |
|
|
|
| 30 |
B2 |
1246 |
1170 |
207.36 |
|
|
|
| 31 |
B2 |
1182 |
1110 |
213.23 |
|
|
|
| 32 |
B2 |
1016 |
954 |
4.22 |
|
|
|
| 33 |
B2 |
434 |
408 |
2.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25417.6 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 23872.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.305 |
| H2 |
-0.897 |
0.000 |
0.946 |
| H3 |
0.897 |
0.000 |
0.946 |
| O4 |
0.000 |
1.116 |
-0.538 |
| O5 |
0.000 |
-1.116 |
-0.538 |
| C6 |
0.000 |
2.324 |
0.212 |
| C7 |
0.000 |
-2.324 |
0.212 |
| H8 |
0.000 |
3.133 |
-0.510 |
| H9 |
0.000 |
-3.133 |
-0.510 |
| H10 |
-0.891 |
2.397 |
0.842 |
| H11 |
0.891 |
2.397 |
0.842 |
| H12 |
0.891 |
-2.397 |
0.842 |
| H13 |
-0.891 |
-2.397 |
0.842 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
O4 |
O5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.1028 | 1.1028 | 1.3989 | 1.3989 | 2.3254 | 2.3254 | 3.2371 | 3.2371 | 2.6133 | 2.6133 | 2.6133 | 2.6133 |
H2 | 1.1028 | | 1.7944 | 2.0626 | 2.0626 | 2.5966 | 2.5966 | 3.5692 | 3.5692 | 2.3996 | 2.9925 | 2.9925 | 2.3996 | H3 | 1.1028 | 1.7944 | | 2.0626 | 2.0626 | 2.5966 | 2.5966 | 3.5692 | 3.5692 | 2.9925 | 2.3996 | 2.3996 | 2.9925 | O4 | 1.3989 | 2.0626 | 2.0626 | | 2.2327 | 1.4214 | 3.5208 | 2.0169 | 4.2495 | 2.0830 | 2.0830 | 3.8787 | 3.8787 | O5 | 1.3989 | 2.0626 | 2.0626 | 2.2327 | | 3.5208 | 1.4214 | 4.2495 | 2.0169 | 3.8787 | 3.8787 | 2.0830 | 2.0830 | C6 | 2.3254 | 2.5966 | 2.5966 | 1.4214 | 3.5208 | | 4.6470 | 1.0844 | 5.5040 | 1.0934 | 1.0934 | 4.8453 | 4.8453 | C7 | 2.3254 | 2.5966 | 2.5966 | 3.5208 | 1.4214 | 4.6470 | | 5.5040 | 1.0844 | 4.8453 | 4.8453 | 1.0934 | 1.0934 | H8 | 3.2371 | 3.5692 | 3.5692 | 2.0169 | 4.2495 | 1.0844 | 5.5040 | | 6.2660 | 1.7778 | 1.7778 | 5.7623 | 5.7623 | H9 | 3.2371 | 3.5692 | 3.5692 | 4.2495 | 2.0169 | 5.5040 | 1.0844 | 6.2660 | | 5.7623 | 5.7623 | 1.7778 | 1.7778 | H10 | 2.6133 | 2.3996 | 2.9925 | 2.0830 | 3.8787 | 1.0934 | 4.8453 | 1.7778 | 5.7623 | | 1.7816 | 5.1150 | 4.7947 | H11 | 2.6133 | 2.9925 | 2.3996 | 2.0830 | 3.8787 | 1.0934 | 4.8453 | 1.7778 | 5.7623 | 1.7816 | | 4.7947 | 5.1150 | H12 | 2.6133 | 2.9925 | 2.3996 | 3.8787 | 2.0830 | 4.8453 | 1.0934 | 5.7623 | 1.7778 | 5.1150 | 4.7947 | | 1.7816 | H13 | 2.6133 | 2.3996 | 2.9925 | 3.8787 | 2.0830 | 4.8453 | 1.0934 | 5.7623 | 1.7778 | 4.7947 | 5.1150 | 1.7816 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C6 |
111.075 |
|
C1 |
O5 |
C7 |
111.075 |
| H2 |
C1 |
H3 |
108.892 |
|
H2 |
C1 |
O4 |
110.513 |
| H2 |
C1 |
O5 |
110.513 |
|
H3 |
C1 |
O4 |
110.513 |
| H3 |
C1 |
O5 |
110.513 |
|
O4 |
C1 |
O5 |
105.882 |
| O4 |
C6 |
H8 |
106.420 |
|
O4 |
C6 |
H10 |
111.186 |
| O4 |
C6 |
H11 |
111.186 |
|
O5 |
C7 |
H9 |
106.420 |
| O5 |
C7 |
H12 |
111.186 |
|
O5 |
C7 |
H13 |
111.186 |
| H8 |
C6 |
H10 |
109.438 |
|
H8 |
C6 |
H11 |
109.438 |
| H9 |
C7 |
H12 |
109.438 |
|
H9 |
C7 |
H13 |
109.438 |
| H10 |
C6 |
H11 |
109.120 |
|
H12 |
C7 |
H13 |
109.120 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
