return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-213.130018
Energy at 298.15K-213.142803
HF Energy-212.336545
Nuclear repulsion energy188.878588
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3571 3354 0.09      
2 A' 3231 3034 39.10      
3 A' 3210 3015 55.41      
4 A' 3149 2957 46.07      
5 A' 3119 2929 12.54      
6 A' 3027 2843 173.68      
7 A' 1571 1476 0.84      
8 A' 1548 1454 3.58      
9 A' 1538 1444 15.29      
10 A' 1470 1380 3.64      
11 A' 1432 1345 0.58      
12 A' 1345 1263 5.08      
13 A' 1264 1188 1.82      
14 A' 1196 1123 11.63      
15 A' 1097 1030 8.31      
16 A' 925 869 7.65      
17 A' 848 796 3.65      
18 A' 795 747 80.77      
19 A' 433 407 0.28      
20 A' 267 251 1.62      
21 A' 191 179 0.90      
22 A' 115 108 1.77      
23 A" 3231 3034 6.13      
24 A" 3210 3015 16.55      
25 A" 3148 2957 0.50      
26 A" 3119 2929 35.29      
27 A" 3025 2841 12.92      
28 A" 1561 1466 2.08      
29 A" 1546 1452 0.17      
30 A" 1542 1448 8.29      
31 A" 1507 1416 38.80      
32 A" 1454 1365 12.98      
33 A" 1388 1303 20.00      
34 A" 1312 1233 2.84      
35 A" 1197 1124 44.22      
36 A" 1144 1074 8.95      
37 A" 1095 1029 3.88      
38 A" 972 913 0.31      
39 A" 828 777 0.46      
40 A" 437 410 0.41      
41 A" 268 252 0.63      
42 A" 109 103 1.26      

Unscaled Zero Point Vibrational Energy (zpe) 33715.1 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 31665.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.59385 0.07043 0.06637

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.283 0.000
C2 0.017 0.519 1.218
C3 0.017 0.519 -1.218
C4 0.017 -0.372 2.447
C5 0.017 -0.372 -2.447
H6 -0.798 -0.890 0.000
H7 -0.835 1.215 1.260
H8 0.922 1.130 1.203
H9 -0.835 1.215 -1.260
H10 0.922 1.130 -1.203
H11 0.048 0.225 3.358
H12 -0.884 -0.985 2.480
H13 0.880 -1.034 2.430
H14 0.048 0.225 -3.358
H15 -0.884 -0.985 -2.480
H16 0.880 -1.034 -2.430

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.45871.45872.44882.44881.01682.13462.06412.13462.06413.39612.73062.68613.39612.73062.6861
C21.45872.43631.51823.77212.03381.10071.09152.71102.65552.16012.16052.15094.58554.09314.0582
C31.45872.43633.77211.51822.03382.71102.65551.10071.09154.58554.09314.05822.16012.16052.1509
C42.44881.51823.77214.89432.63102.15732.14994.12134.04901.08901.09021.08775.83565.04664.9973
C52.44883.77211.51824.89432.63104.12134.04902.15732.14995.83565.04664.99731.08901.09021.0877
H61.01682.03382.03382.63102.63102.45372.91312.45372.91313.63792.48362.95703.63792.48362.9570
H72.13461.10072.71102.15734.12132.45371.76002.51953.02622.48232.51623.06084.80434.33934.6492
H82.06411.09152.65552.14994.04902.91311.76003.02622.40542.49553.06012.48784.73084.61484.2286
H92.13462.71101.10074.12132.15732.45372.51953.02621.76004.80434.33934.64922.48232.51623.0608
H102.06412.65551.09154.04902.14992.91313.02622.40541.76004.73084.61484.22862.49553.06012.4878
H113.39612.16014.58551.08905.83563.63792.48232.49554.80434.73081.76091.77106.71566.03455.9815
H122.73062.16054.09311.09025.04662.48362.51623.06014.33934.61481.76091.76526.03454.96075.2181
H132.68612.15094.05821.08774.99732.95703.06082.48784.64924.22861.77101.76525.98155.21814.8607
H143.39614.58552.16015.83561.08903.63794.80434.73082.48232.49556.71566.03455.98151.76091.7710
H152.73064.09312.16055.04661.09022.48364.33934.61482.51623.06016.03454.96075.21811.76091.7652
H162.68614.05822.15094.99731.08772.95704.64924.22863.06082.48785.98155.21814.86071.77101.7652

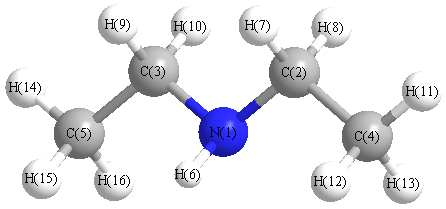
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 110.678 N1 C2 H7 112.281
N1 C2 H8 107.200 N1 C3 C5 110.678
N1 C3 H9 112.281 N1 C3 H10 107.200
C2 N1 C3 113.255 C2 N1 H6 109.184
C2 C4 H11 110.815 C2 C4 H12 110.776
C2 C4 H13 110.168 C3 N1 H6 109.184
C3 C5 H14 110.815 C3 C5 H15 110.776
C3 C5 H16 110.168 C4 C2 H7 109.898
C4 C2 H8 109.856 C5 C3 H8 150.798
C5 C3 H10 109.856 H7 C2 H8 106.802
H9 C3 H10 106.802 H11 C4 H12 107.811
H11 C4 H13 108.897 H12 C4 H13 108.294
H14 C5 H15 107.811 H14 C5 H16 108.897
H15 C5 H16 108.294
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability