return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HCOOC2H5 (Ethyl formate)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS trans 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-267.653052
Energy at 298.15K-267.660277
HF Energy-266.842132
Nuclear repulsion energy177.227851
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3231 3034 16.37      
2 A' 3170 2978 44.51      
3 A' 3152 2961 16.13      
4 A' 3134 2943 14.12      
5 A' 1774 1667 320.71      
6 A' 1563 1468 6.68      
7 A' 1547 1453 3.49      
8 A' 1472 1383 20.58      
9 A' 1427 1340 2.73      
10 A' 1422 1336 5.77      
11 A' 1236 1161 422.01      
12 A' 1165 1094 5.74      
13 A' 1064 999 16.22      
14 A' 874 821 14.97      
15 A' 787 739 2.48      
16 A' 386 362 5.13      
17 A' 229 216 7.65      
18 A" 3246 3049 24.96      
19 A" 3214 3018 6.60      
20 A" 1534 1440 7.14      
21 A" 1322 1242 1.12      
22 A" 1209 1136 4.51      
23 A" 1044 981 0.05      
24 A" 832 782 0.18      
25 A" 350 328 23.48      
26 A" 247 232 3.05      
27 A" 65 61 0.48      

Unscaled Zero Point Vibrational Energy (zpe) 20348.1 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 19110.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.58886 0.09656 0.08561

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.178 -0.248 0.000
C2 -0.703 -0.563 0.000
O3 0.000 0.708 0.000
C4 1.343 0.616 0.000
O5 1.987 -0.418 0.000
H6 -2.748 -1.175 0.000
H7 -2.448 0.325 0.884
H8 -2.448 0.325 -0.884
H9 -0.399 -1.123 -0.882
H10 -0.399 -1.123 0.882
H11 1.774 1.620 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 C4 O5 H6 H7 H8 H9 H10 H11
C11.50782.37843.62564.16791.08881.08721.08722.16962.16964.3708
C21.50781.45262.36182.69372.13472.14792.14791.08851.08853.3015
O32.37841.45261.34662.28393.33172.63092.63092.07192.07191.9944
C43.62562.36181.34661.21804.46653.90403.90402.61552.61551.0923
O54.16792.69372.28391.21804.79494.58254.58252.63972.63972.0494
H61.08882.13473.33174.46654.79491.76671.76672.50952.50955.3162
H71.08722.14792.63093.90404.58251.76671.76793.06802.50874.5038
H81.08722.14792.63093.90404.58251.76671.76792.50873.06804.5038
H92.16961.08852.07192.61552.63972.50953.06802.50871.76433.6093
H102.16961.08852.07192.61552.63972.50952.50873.06801.76433.6093
H114.37083.30151.99441.09232.04945.31624.50384.50383.60933.6093

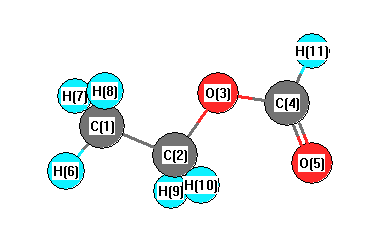
picture of Ethyl formate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 106.895 C1 C2 H9 112.366
C1 C2 H10 112.366 C2 C1 H6 109.533
C2 C1 H7 110.677 C2 C1 H8 110.677
C2 O3 C4 115.023 O3 C2 H9 108.396
O3 C2 H10 108.396 O3 C4 O5 125.803
O3 C4 H11 109.277 O5 C4 H11 124.920
H6 C1 H7 108.555 H6 C1 H8 108.555
H7 C1 H8 108.786 H9 C2 H10 108.278
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability