return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (cyclohexene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-233.913697
Energy at 298.15K-233.925358
HF Energy-233.038447
Nuclear repulsion energy237.511128
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3246 3049 39.54      
2 A 3167 2975 40.21      
3 A 3146 2955 0.73      
4 A 3105 2916 23.97      
5 A 3091 2903 0.00      
6 A 1723 1618 4.23      
7 A 1541 1447 1.78      
8 A 1517 1425 0.16      
9 A 1418 1332 0.03      
10 A 1409 1323 0.27      
11 A 1293 1214 0.47      
12 A 1273 1195 0.09      
13 A 1181 1110 0.05      
14 A 1120 1051 0.01      
15 A 1112 1044 0.05      
16 A 992 931 0.00      
17 A 938 881 1.00      
18 A 858 806 0.06      
19 A 843 792 0.98      
20 A 503 472 0.02      
21 A 406 381 0.02      
22 A 286 269 0.03      
23 B 3223 3027 7.63      
24 B 3172 2979 49.10      
25 B 3146 2955 54.06      
26 B 3107 2918 27.45      
27 B 3091 2903 63.59      
28 B 1532 1438 5.97      
29 B 1525 1432 7.63      
30 B 1451 1363 0.01      
31 B 1410 1324 1.73      
32 B 1378 1295 1.27      
33 B 1317 1237 2.36      
34 B 1180 1108 6.91      
35 B 1081 1016 3.28      
36 B 1039 976 0.85      
37 B 956 898 5.74      
38 B 917 861 4.38      
39 B 743 698 15.04      
40 B 659 619 28.62      
41 B 460 432 0.85      
42 B 173 162 0.20      

Unscaled Zero Point Vibrational Energy (zpe) 32864.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 30866.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.15969 0.15220 0.08613

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.008 -0.672 1.296
C2 0.008 0.672 1.296
C3 0.000 1.497 0.039
C4 0.000 -1.497 0.039
C5 -0.385 0.658 -1.177
C6 0.385 -0.658 -1.177
H7 -0.027 -1.199 2.244
H8 0.027 1.199 2.244
H9 0.989 1.942 -0.114
H10 -0.989 -1.942 -0.114
H11 -0.692 2.334 0.156
H12 0.692 -2.334 0.156
H13 -1.457 0.444 -1.143
H14 1.457 -0.444 -1.143
H15 -0.199 1.218 -2.095
H16 0.199 -1.218 -2.095

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.34432.50721.50362.83352.50411.08442.09743.13292.13663.28742.13363.04882.85413.88703.4411
C21.34431.50362.50722.50412.83352.09741.08442.13663.13292.13363.28742.85413.04883.44113.8870
C32.50721.50362.99431.52672.50473.48272.22521.09523.58161.09253.89522.15152.69952.16173.4591
C41.50362.50722.99432.50471.52672.22523.48273.58161.09523.89521.09252.69952.15153.45912.1617
C52.83352.50411.52672.50471.52543.90893.48782.15982.87302.16333.44861.09362.14691.09132.1688
C62.50412.83352.50471.52671.52543.48783.90892.87302.15983.44862.16332.14691.09362.16881.0913
H71.08442.09743.48272.22523.90893.48782.39784.05672.65324.15722.48334.02653.77404.96964.3451
H82.09741.08442.22523.48273.48783.90892.39782.65324.05672.48334.15723.77404.02654.34514.9696
H93.13292.13661.09523.58162.15982.87304.05672.65324.35831.74774.29483.04702.63972.42043.8119
H102.13663.13293.58161.09522.87302.15982.65324.05674.35834.29481.74772.63973.04703.81192.4204
H113.28742.13361.09253.89522.16333.44864.15722.48331.74774.29484.86942.41773.74492.56104.2986
H122.13363.28743.89521.09253.44862.16332.48334.15724.29481.74774.86943.74492.41774.29862.5610
H133.04882.85412.15152.69951.09362.14694.02653.77403.04702.63972.41773.74493.04621.75762.5316
H142.85413.04882.69952.15152.14691.09363.77404.02652.63973.04703.74492.41773.04622.53161.7576
H153.88703.44112.16173.45911.09132.16884.96964.34512.42043.81192.56104.29861.75762.53162.4676
H163.44113.88703.45912.16172.16881.09134.34514.96963.81192.42044.29862.56102.53161.75762.4676

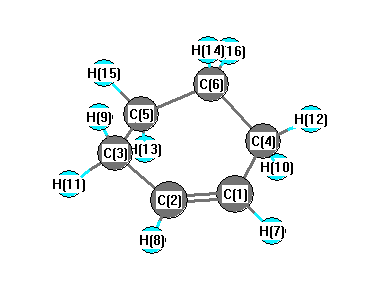
picture of cyclohexene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.272 C1 C2 H8 119.053
C1 C4 C6 111.450 C1 C4 H10 109.592
C1 C4 H12 109.516 C2 C1 C4 123.272
C2 C1 H7 119.053 C2 C3 C5 111.450
C2 C3 H9 109.592 C2 C3 H11 109.516
C3 C2 H8 117.675 C3 C5 C6 110.297
C3 C5 H13 109.278 C3 C5 H15 110.213
C4 C1 H7 117.675 C4 C6 C5 110.297
C4 C6 H14 109.278 C4 C6 H16 110.213
C5 C3 H9 109.829 C5 C3 H11 110.268
C5 C6 H14 109.003 C5 C6 H16 110.864
C6 C4 H10 109.829 C6 C4 H12 110.268
C6 C5 H13 109.003 C6 C5 H15 110.864
H9 C3 H11 106.038 H10 C4 H12 106.038
H13 C5 H15 107.110 H14 C6 H16 107.110
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability