return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Cyclobutane, methylene-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-194.682137
Energy at 298.15K-194.690693
HF Energy-193.959197
Nuclear repulsion energy168.200906
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3224 3028 6.22      
2 A' 3223 3027 39.83      
3 A' 3195 3001 9.17      
4 A' 3149 2958 42.19      
5 A' 3128 2937 11.67      
6 A' 1765 1658 16.85      
7 A' 1546 1452 0.10      
8 A' 1520 1428 3.12      
9 A' 1478 1389 0.63      
10 A' 1288 1210 0.39      
11 A' 1238 1163 0.26      
12 A' 1111 1043 2.16      
13 A' 998 937 0.76      
14 A' 923 867 0.09      
15 A' 870 817 54.61      
16 A' 768 721 2.11      
17 A' 674 633 2.03      
18 A' 376 353 6.38      
19 A' 133 125 0.42      
20 A" 3323 3121 10.37      
21 A" 3199 3005 20.57      
22 A" 3125 2935 39.90      
23 A" 1509 1417 3.21      
24 A" 1312 1232 0.00      
25 A" 1268 1191 0.54      
26 A" 1237 1162 0.72      
27 A" 1205 1132 1.46      
28 A" 1054 990 0.22      
29 A" 951 893 1.81      
30 A" 934 878 0.62      
31 A" 829 778 0.11      
32 A" 666 625 0.11      
33 A" 355 334 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 25786.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 24218.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.34413 0.15542 0.11802

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.244 -0.413 1.084
C2 0.244 -0.413 -1.084
C3 0.000 0.618 0.000
C4 -0.019 -1.491 0.000
C5 -0.471 1.867 0.000
H6 -0.399 -0.391 1.963
H7 -0.399 -0.391 -1.963
H8 1.288 -0.422 1.404
H9 1.288 -0.422 -1.404
H10 -0.665 2.392 0.925
H11 -0.665 2.392 -0.925
H12 0.619 -2.373 0.000
H13 -1.063 -1.800 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C12.16851.51571.55142.62381.08923.11441.09182.69862.95313.56832.27082.1928
C22.16851.51571.55142.62383.11441.08922.69861.09183.56832.95312.27082.1928
C31.51571.51572.10841.33502.24282.24282.17062.17062.10882.10883.05372.6409
C41.55141.55142.10843.38792.28182.28182.19622.19624.04354.04351.08841.0887
C52.62382.62381.33503.38792.99292.99293.20993.20991.08151.08154.37743.7143
H61.08923.11442.24282.28182.99293.92591.77733.76632.98244.01982.96902.5058
H73.11441.08922.24282.28182.99293.92593.76631.77734.01982.98242.96902.5058
H81.09182.69862.17062.19623.20991.77733.76632.80853.45884.14252.49533.0658
H92.69861.09182.17062.19623.20993.76631.77732.80854.14253.45882.49533.0658
H102.95313.56832.10884.04351.08152.98244.01983.45884.14251.85035.02074.3112
H113.56832.95312.10884.04351.08154.01982.98244.14253.45881.85035.02074.3112
H122.27082.27083.05371.08844.37742.96902.96902.49532.49535.02075.02071.7765
H132.19282.19282.64091.08873.71432.50582.50583.06583.06584.31124.31121.7765

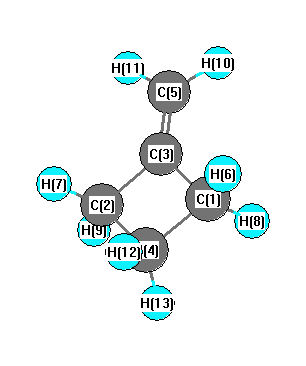
picture of Cyclobutane, methylene- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C2 91.346 C1 C3 C5 133.875
C1 C4 C2 88.677 C1 C4 H12 117.616
C1 C4 H13 111.107 C2 C3 C5 133.875
C2 C4 H12 117.616 C2 C4 H13 111.107
C3 C1 C4 86.849 C3 C1 H6 117.930
C3 C1 H8 111.678 C3 C2 C4 86.849
C3 C2 H7 117.930 C3 C2 H9 111.678
C3 C5 H10 121.189 C3 C5 H11 121.189
C4 C1 H6 118.516 C4 C1 H8 111.195
C4 C2 H7 118.516 C4 C2 H9 111.195
H6 C1 H8 109.162 H7 C2 H9 109.162
H10 C5 H11 117.622 H12 C4 H13 109.369
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability