Jump to
S1C2
S2C1
S2C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -1817.020061 |
| Energy at 298.15K | -1817.021443 |
| Nuclear repulsion energy | 66.852654 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3199 |
3005 |
25.19 |
|
|
|
| 2 |
A1 |
1395 |
1310 |
18.21 |
|
|
|
| 3 |
A1 |
550 |
517 |
28.25 |
|
|
|
| 4 |
B1 |
630 |
592 |
83.28 |
|
|
|
| 5 |
B2 |
3316 |
3115 |
4.28 |
|
|
|
| 6 |
B2 |
575 |
540 |
1.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4832.8 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 4538.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
0.000 |
0.000 |
0.441 |
| C2 |
0.000 |
0.000 |
-1.506 |
| H3 |
0.000 |
0.911 |
-2.095 |
| H4 |
0.000 |
-0.911 |
-2.095 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
H3 |
H4 |
| Zn1 | | 1.9471 | 2.6940 | 2.6940 |
C2 | 1.9471 | | 1.0841 | 1.0841 | H3 | 2.6940 | 1.0841 | | 1.8211 | H4 | 2.6940 | 1.0841 | 1.8211 | |
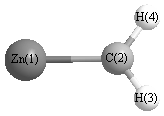 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Zn1 |
C2 |
H3 |
122.866 |
|
Zn1 |
C2 |
H4 |
122.866 |
| H3 |
C2 |
H4 |
114.267 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -1817.020061 |
| Energy at 298.15K | -1817.021443 |
| Nuclear repulsion energy | 66.852654 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2v
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -1816.985839 |
| Energy at 298.15K | -1816.987518 |
| Nuclear repulsion energy | 71.327577 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3244 |
3047 |
10.94 |
|
|
|
| 2 |
A1 |
1246 |
1170 |
56.62 |
|
|
|
| 3 |
A1 |
686 |
645 |
10.66 |
|
|
|
| 4 |
B1 |
16366 |
15371 |
0.00 |
|
|
|
| 5 |
B2 |
3435 |
3226 |
23.02 |
|
|
|
| 6 |
B2 |
671 |
630 |
56.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12823.8 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12044.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
0.000 |
0.000 |
0.407 |
| C2 |
0.000 |
0.000 |
-1.413 |
| H3 |
0.000 |
0.978 |
-1.865 |
| H4 |
0.000 |
-0.978 |
-1.865 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
H3 |
H4 |
| Zn1 | | 1.8205 | 2.4733 | 2.4733 |
C2 | 1.8205 | | 1.0768 | 1.0768 | H3 | 2.4733 | 1.0768 | | 1.9553 | H4 | 2.4733 | 1.0768 | 1.9553 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -1817.007126 |
| Energy at 298.15K | -1817.008657 |
| Nuclear repulsion energy | 69.659059 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2998 |
2816 |
39.32 |
|
|
|
| 2 |
A' |
1377 |
1293 |
25.30 |
|
|
|
| 3 |
A' |
786 |
738 |
209.07 |
|
|
|
| 4 |
A' |
555 |
521 |
3.17 |
|
|
|
| 5 |
A" |
3098 |
2910 |
23.14 |
|
|
|
| 6 |
A" |
727 |
682 |
26.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4769.8 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 4479.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
-0.032 |
-0.411 |
0.000 |
| C2 |
-0.032 |
1.480 |
0.000 |
| H3 |
0.575 |
1.729 |
0.889 |
| H4 |
0.575 |
1.729 |
-0.889 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
H3 |
H4 |
| Zn1 | | 1.8915 | 2.3954 | 2.3954 |
C2 | 1.8915 | | 1.1048 | 1.1048 | H3 | 2.3954 | 1.1048 | | 1.7777 | H4 | 2.3954 | 1.1048 | 1.7777 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
