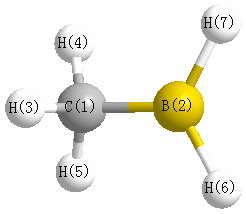
Vibrational Frequencies calculated at wB97X-D/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3112 |
2963 |
16.08 |
|
|
|
| 2 |
A' |
3006 |
2862 |
3.35 |
|
|
|
| 3 |
A' |
2569 |
2446 |
124.32 |
|
|
|
| 4 |
A' |
1494 |
1423 |
4.76 |
|
|
|
| 5 |
A' |
1348 |
1284 |
80.65 |
|
|
|
| 6 |
A' |
1256 |
1196 |
22.35 |
|
|
|
| 7 |
A' |
1084 |
1032 |
72.25 |
|
|
|
| 8 |
A' |
971 |
925 |
12.68 |
|
|
|
| 9 |
A' |
571 |
544 |
0.29 |
|
|
|
| 10 |
A" |
3164 |
3013 |
17.28 |
|
|
|
| 11 |
A" |
2640 |
2514 |
181.61 |
|
|
|
| 12 |
A" |
1445 |
1376 |
6.51 |
|
|
|
| 13 |
A" |
1068 |
1017 |
20.63 |
|
|
|
| 14 |
A" |
692 |
659 |
1.06 |
|
|
|
| 15 |
A" |
127 |
121 |
1.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12272.0 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 11686.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.632 |
|
|
|
| 2 |
B |
0.182 |
|
|
|
| 3 |
H |
0.170 |
|
|
|
| 4 |
H |
0.172 |
|
|
|
| 5 |
H |
0.172 |
|
|
|
| 6 |
H |
-0.032 |
|
|
|
| 7 |
H |
-0.032 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.114 |
-0.631 |
0.000 |
0.642 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.684 |
-0.164 |
0.000 |
| y |
-0.164 |
-16.500 |
0.000 |
| z |
0.000 |
0.000 |
-15.697 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.414 |
-0.164 |
0.000 |
| y |
-0.164 |
-1.809 |
0.000 |
| z |
0.000 |
0.000 |
-0.605 |
|
| Polar |
|---|
| 3z2-r2 | -1.210 |
|---|
| x2-y2 | 2.816 |
|---|
| xy | -0.164 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.029 |
-0.020 |
0.000 |
| y |
-0.020 |
4.608 |
0.000 |
| z |
0.000 |
0.000 |
4.068 |
<r2> (average value of r
2) Å
2
| <r2> |
29.414 |
| (<r2>)1/2 |
5.423 |