return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CClCHCH2 (1,3-Butadiene, 2-chloro-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-614.538169
Energy at 298.15K 
HF Energy-613.832879
Nuclear repulsion energy197.489191
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3360 3156 1.96 51.90 0.75 0.86
2 A' 3356 3152 3.49 66.44 0.68 0.81
3 A' 3255 3057 3.67 183.43 0.13 0.24
4 A' 3250 3052 1.24 79.85 0.14 0.24
5 A' 3239 3042 3.64 24.14 0.73 0.84
6 A' 1720 1616 5.08 150.68 0.20 0.33
7 A' 1658 1557 32.65 5.18 0.28 0.44
8 A' 1496 1405 0.59 52.45 0.40 0.57
9 A' 1442 1354 2.96 3.26 0.46 0.63
10 A' 1342 1261 0.02 25.62 0.24 0.39
11 A' 1281 1203 47.10 11.73 0.57 0.73
12 A' 1059 995 5.60 3.55 0.59 0.75
13 A' 932 876 8.35 0.63 0.75 0.86
14 A' 660 620 19.25 9.60 0.09 0.16
15 A' 536 503 0.72 5.17 0.55 0.71
16 A' 396 372 1.21 3.51 0.74 0.85
17 A' 251 236 0.07 2.37 0.69 0.82
18 A" 1004 943 31.47 0.11 0.75 0.86
19 A" 908 853 42.06 4.13 0.75 0.86
20 A" 861 809 50.80 4.38 0.75 0.86
21 A" 735 690 2.26 3.54 0.75 0.86
22 A" 648 609 0.73 1.15 0.75 0.86
23 A" 374 351 8.17 0.29 0.75 0.86
24 A" 109 102 0.48 0.26 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 16936.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 15906.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.18085 0.12421 0.07364

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.292 1.889 0.000
C2 0.000 0.577 0.000
C3 1.368 0.067 0.000
C4 1.734 -1.225 0.000
Cl5 -1.293 -0.590 0.000
H6 0.513 2.611 0.000
H7 -1.308 2.249 0.000
H8 2.129 0.839 0.000
H9 2.779 -1.496 0.000
H10 1.006 -2.023 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 Cl5 H6 H7 H8 H9 H10
C11.34412.46463.71512.67381.08021.07822.63904.57014.1215
C21.34411.45992.50091.74172.09712.12272.14533.46682.7877
C32.46461.45991.34302.74052.68373.45281.08432.10552.1205
C43.71512.50091.34303.09254.02574.61772.10161.07941.0793
Cl52.67381.74172.74053.09253.67462.83903.70844.17102.7089
H61.08022.09712.68374.02573.67461.85612.39874.69044.6595
H71.07822.12273.45284.61772.83901.85613.71545.54314.8582
H82.63902.14531.08432.10163.70842.39873.71542.42343.0738
H94.57013.46682.10551.07944.17104.69045.54312.42341.8488
H104.12152.78772.12051.07932.70894.65954.85823.07381.8488

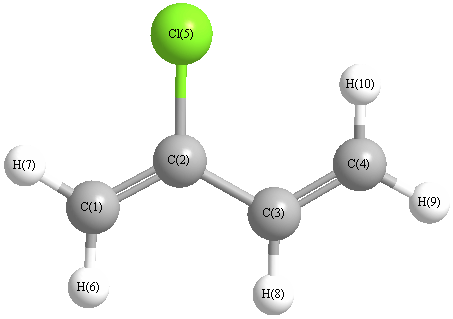
picture of 1,3-Butadiene, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 122.985 C1 C2 Cl5 119.547
C2 C1 H6 119.371 C2 C1 H7 122.019
C2 C3 C4 126.268 C2 C3 H8 114.158
C3 C2 Cl5 117.468 C3 C4 H9 120.339
C3 C4 H10 121.812 C4 C3 H8 119.575
H6 C1 H7 118.610 H9 C4 H10 117.849
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability