return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3C(NH2)HCH2CH3 (2-Butanamine)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-213.142310
Energy at 298.15K-213.155189
HF Energy-212.344561
Nuclear repulsion energy194.748680
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3676 3453 1.60      
2 A 3570 3353 0.33      
3 A 3221 3025 29.48      
4 A 3215 3020 28.36      
5 A 3206 3011 37.28      
6 A 3204 3009 25.19      
7 A 3170 2978 12.66      
8 A 3113 2924 3.96      
9 A 3113 2924 62.16      
10 A 3110 2921 15.85      
11 A 3022 2839 66.41      
12 A 1697 1594 53.31      
13 A 1558 1464 9.65      
14 A 1546 1452 4.26      
15 A 1546 1452 5.67      
16 A 1538 1445 4.66      
17 A 1521 1429 0.93      
18 A 1462 1373 4.20      
19 A 1453 1364 11.16      
20 A 1446 1358 7.13      
21 A 1416 1330 18.61      
22 A 1368 1285 2.41      
23 A 1329 1248 1.79      
24 A 1284 1206 0.60      
25 A 1232 1157 2.18      
26 A 1194 1121 6.86      
27 A 1103 1036 1.49      
28 A 1064 999 4.05      
29 A 1043 979 3.10      
30 A 1008 947 2.14      
31 A 971 912 8.50      
32 A 884 830 116.23      
33 A 838 787 10.80      
34 A 792 744 3.46      
35 A 493 463 7.72      
36 A 469 440 6.17      
37 A 387 363 0.63      
38 A 318 298 36.68      
39 A 271 255 1.50      
40 A 243 228 4.68      
41 A 236 221 11.36      
42 A 121 114 1.69      

Unscaled Zero Point Vibrational Energy (zpe) 33724.8 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 31674.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.26397 0.11511 0.08839

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.551 1.371 -0.230
H2 -0.227 1.940 0.084
H3 1.398 1.838 0.079
C4 1.774 -0.706 -0.013
H5 1.899 -0.741 -1.094
H6 2.637 -0.195 0.415
H7 1.760 -1.724 0.375
C8 0.483 0.018 0.332
H9 0.377 0.030 1.428
C10 -0.727 -0.709 -0.242
H11 -0.600 -0.773 -1.324
H12 -0.727 -1.728 0.150
C13 -2.058 -0.041 0.086
H14 -2.894 -0.648 -0.259
H15 -2.144 0.931 -0.396
H16 -2.169 0.097 1.163

Atom - Atom Distances (Å)
  N1 H2 H3 C4 H5 H6 H7 C8 H9 C10 H11 H12 C13 H14 H15 H16
N11.01371.01462.42062.65062.68673.37791.46702.13952.44162.66893.37472.98393.99332.73643.3114
H21.01371.62863.31903.61863.58734.17842.06372.41182.71463.07913.70262.69723.73132.21872.8864
H31.01461.62862.57322.87692.40383.59242.05242.47593.33213.57464.15203.93374.97103.68724.1148
C42.42063.31902.57321.08941.09061.08921.51992.13802.51172.71352.70703.89064.67454.26404.1924
H52.65063.61862.87691.08941.76721.77322.14833.04532.76102.50973.06904.18784.86534.43044.7269
H62.68673.58732.40381.09061.76721.76332.16582.48693.46603.72043.70684.70895.58984.97854.8727
H73.37794.17843.59241.08921.77321.76332.16072.47022.75683.06042.49774.18284.81864.78464.4022
C81.46702.06372.05241.51992.14832.16582.16071.10081.52402.13192.13272.55373.49222.87552.7805
H92.13952.41182.47592.13803.04532.48692.47021.10082.13363.02922.43812.78083.74173.23962.5606
C102.44162.71463.33212.51172.76103.46602.75681.52402.13361.09221.09261.52452.16732.17272.1680
H112.66893.07913.57462.71352.50973.72043.06042.13193.02921.09221.76192.15652.53192.47973.0667
H123.37473.70264.15202.70703.06903.70682.49772.13272.43811.09261.76192.15002.45523.06282.5370
C132.98392.69723.93373.89064.18784.70894.18282.55372.78081.52452.15652.15001.08891.08891.0910
H143.99333.73134.97104.67454.86535.58984.81863.49223.74172.16732.53192.45521.08891.75341.7607
H152.73642.21873.68724.26404.43044.97854.78462.87553.23962.17272.47973.06281.08891.75341.7686
H163.31142.88644.11484.19244.72694.87274.40222.78052.56062.16803.06672.53701.09101.76071.7686

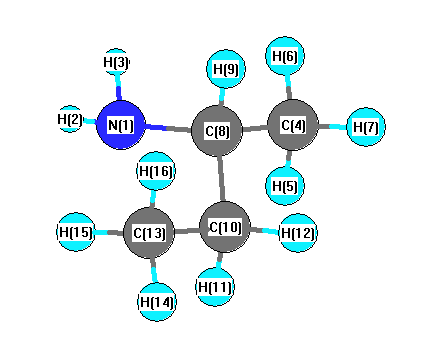
picture of 2-Butanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C8 C4 108.252 N1 C8 H9 112.062
N1 C8 C10 109.417 H2 N1 H3 106.817
H2 N1 C8 111.273 H3 N1 C8 110.251
C4 C8 H9 108.265 C4 C8 C10 111.210
H5 C4 H6 108.315 H5 C4 H7 108.956
H5 C4 C8 109.738 H6 C4 H7 107.980
H6 C4 C8 111.054 H7 C4 C8 110.729
C8 C10 H11 108.014 C8 C10 H12 108.057
C8 C10 C13 113.799 H9 C8 C10 107.657
C10 C13 H14 110.957 C10 C13 H15 111.395
C10 C13 H16 110.893 H11 C10 H12 107.498
H11 C10 C13 109.903 H12 C10 C13 109.365
H14 C13 H15 107.242 H14 C13 H16 107.749
H15 C13 H16 108.448
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability