return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (cis-2-butenal)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-230.567471
Energy at 298.15K-230.573204
Nuclear repulsion energy154.973311
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3248 3050 17.13      
2 A' 3242 3045 1.17      
3 A' 3221 3025 10.59      
4 A' 3113 2924 14.24      
5 A' 3033 2849 136.10      
6 A' 1753 1647 31.24      
7 A' 1715 1611 185.85      
8 A' 1535 1441 22.72      
9 A' 1460 1371 2.27      
10 A' 1452 1363 2.87      
11 A' 1346 1264 14.43      
12 A' 1345 1263 5.96      
13 A' 1171 1100 4.79      
14 A' 1056 991 13.46      
15 A' 918 862 29.00      
16 A' 745 700 37.38      
17 A' 398 374 3.04      
18 A' 207 194 5.19      
19 A" 3195 3000 9.11      
20 A" 1520 1427 8.11      
21 A" 1091 1025 4.28      
22 A" 1022 960 8.47      
23 A" 1001 940 32.39      
24 A" 752 706 0.70      
25 A" 231 217 4.63      
26 A" 190 178 0.99      
27 A" 115 108 6.90      

Unscaled Zero Point Vibrational Energy (zpe) 20036.3 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 18818.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.63464 0.08702 0.07762

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.469 0.560 0.000
C2 0.000 0.694 0.000
C3 0.807 -0.381 0.000
C4 2.298 -0.330 0.000
O5 -2.072 -0.514 0.000
H6 -2.037 1.504 0.000
H7 0.406 1.699 0.000
H8 0.335 -1.358 0.000
H9 2.663 0.695 0.000
H10 2.697 -0.842 0.876
H11 2.697 -0.842 -0.876

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.47552.46373.87091.23121.10212.19492.63354.13434.48254.4825
C21.47551.34522.51592.39872.19211.08412.08012.66273.22543.2254
C32.46371.34521.49132.88273.41282.11921.08532.14482.13332.1333
C43.87092.51591.49134.37394.70702.77432.21591.08801.09041.0904
O51.23122.39872.88274.37392.01863.32302.55114.88674.86034.8603
H61.10212.19213.41284.70702.01862.45113.71784.76885.35595.3559
H72.19491.08412.11922.77433.32302.45113.05882.46983.53203.5320
H82.63352.08011.08532.21592.55113.71783.05883.10392.57182.5718
H94.13432.66272.14481.08804.88674.76882.46983.10391.76941.7694
H104.48253.22542.13331.09044.86035.35593.53202.57181.76941.7520
H114.48253.22542.13331.09044.86035.35593.53202.57181.76941.7520

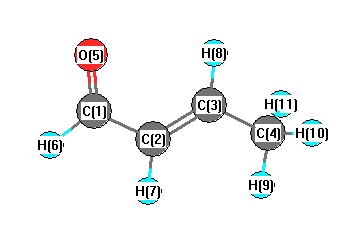
picture of cis-2-butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.650 C1 C2 H7 117.257
C2 C1 O5 124.555 C2 C1 H6 115.758
C2 C3 C4 124.918 C2 C3 H8 117.304
C3 C2 H7 121.093 C3 C4 H9 111.564
C3 C4 H10 110.481 C3 C4 H11 110.481
C4 C3 H8 117.778 O5 C1 H6 119.687
H9 C4 H10 108.634 H9 C4 H11 108.634
H10 C4 H11 106.905
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-31+G** Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.266      
2 C -0.159      
3 C 0.072      
4 C -0.593      
5 O -0.521      
6 H 0.125      
7 H 0.164      
8 H 0.198      
9 H 0.144      
10 H 0.152      
11 H 0.152      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000