return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3 (Tert-butyl radical)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3V 2A"
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-157.276779
Energy at 298.15K-157.285474
HF Energy-156.690943
Nuclear repulsion energy125.831142
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3155 2963 47.13      
2 A1 3051 2866 53.69      
3 A1 1542 1448 21.59      
4 A1 1466 1377 2.60      
5 A1 1146 1077 1.23      
6 A1 793 745 1.80      
7 A1 292 274 3.08      
8 A2 3206 3011 0.00      
9 A2 1517 1425 0.00      
10 A2 992 932 0.00      
11 A2 154 145 0.00      
12 E 3209 3014 38.56      
13 E 3155 2963 13.81      
14 E 3046 2861 34.54      
15 E 1540 1447 3.87      
15 E 1540 1447 3.87      
16 E 1521 1428 3.31      
16 E 1521 1428 3.31      
17 E 1445 1357 7.19      
17 E 1445 1357 7.19      
18 E 1339 1257 4.29      
18 E 1339 1257 4.29      
19 E 1045 982 1.25      
19 E 1045 982 1.25      
20 E 960 902 0.32      
20 E 960 902 0.32      
21 E 383 360 0.17      
21 E 383 360 0.17      
22 E 173 162 0.00      
22 E 173 162 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 26472.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 24862.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.26938 0.26938 0.14676

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C3v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.206
C2 0.000 1.476 -0.022
C3 1.278 -0.738 -0.022
C4 -1.278 -0.738 -0.022
H5 0.000 1.711 -1.095
H6 1.482 -0.855 -1.095
H7 -1.482 -0.855 -1.095
H8 -0.884 1.946 0.408
H9 0.884 1.946 0.408
H10 2.127 -0.207 0.408
H11 1.243 -1.739 0.408
H12 -1.243 -1.739 0.408
H13 -2.127 -0.207 0.408

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.49321.49321.49322.14982.14982.14982.14692.14692.14692.14692.14692.1469
C21.49322.55582.55581.09872.96322.96321.08971.08972.74643.47303.47302.7464
C31.49322.55582.55582.96321.09872.96323.47302.74641.08971.08972.74643.4730
C41.49322.55582.55582.96322.96321.09872.74643.47303.47302.74641.08971.0897
H52.14981.09872.96322.96322.96322.96321.75951.75953.23483.96273.96273.2348
H62.14982.96321.09872.96322.96322.96323.96273.23481.75951.75953.23483.9627
H72.14982.96322.96321.09872.96322.96323.23483.96273.96273.23481.75951.7595
H82.14691.08973.47302.74641.75953.96273.23481.76813.70224.25483.70222.4867
H92.14691.08972.74643.47301.75953.23483.96271.76812.48673.70224.25483.7022
H102.14692.74641.08973.47303.23481.75953.96273.70222.48671.76813.70224.2548
H112.14693.47301.08972.74643.96271.75953.23484.25483.70221.76812.48673.7022
H122.14693.47302.74641.08973.96273.23481.75953.70224.25483.70222.48671.7681
H132.14692.74643.47301.08973.23483.96271.75952.48673.70224.25483.70221.7681

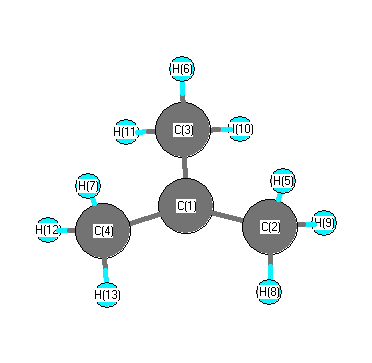
picture of Tert-butyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H5 111.165 C1 C2 H8 111.487
C1 C2 H9 111.487 C1 C3 H6 111.165
C1 C3 H10 111.487 C1 C3 H11 111.487
C1 C4 H7 111.165 C1 C4 H12 111.487
C1 C4 H13 111.487 C2 C1 C3 117.701
C2 C1 C4 117.701 C3 C1 C4 117.700
H5 C2 H8 107.022 H5 C2 H9 107.022
H6 C3 H10 107.022 H6 C3 H11 107.022
H7 C4 H12 107.022 H7 C4 H13 107.022
H8 C2 H9 108.438 H10 C3 H11 108.438
H12 C4 H13 108.438
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability