return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (2H-Thiopyran, tetrahydro-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-593.585313
Energy at 298.15K-593.597774
HF Energy-592.698207
Nuclear repulsion energy312.998258
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3191 2997 21.81      
2 A 3172 2979 43.89      
3 A 3160 2968 32.41      
4 A 3112 2923 21.10      
5 A 3106 2918 27.50      
6 A 3097 2909 24.28      
7 A 1534 1441 2.44      
8 A 1523 1430 12.11      
9 A 1501 1409 7.91      
10 A 1420 1334 0.78      
11 A 1382 1298 6.46      
12 A 1303 1223 0.69      
13 A 1273 1196 6.47      
14 A 1117 1049 2.71      
15 A 1064 999 0.60      
16 A 1006 945 8.95      
17 A 859 806 4.35      
18 A 849 797 0.41      
19 A 695 653 2.84      
20 A 512 481 0.25      
21 A 373 350 0.21      
22 A 351 330 0.09      
23 A 189 177 1.81      
24 A 3192 2998 7.66      
25 A 3167 2974 18.96      
26 A 3115 2926 24.12      
27 A 3107 2918 42.36      
28 A 1516 1424 3.35      
29 A 1496 1406 1.32      
30 A 1423 1336 0.20      
31 A 1403 1318 0.01      
32 A 1346 1264 17.55      
33 A 1317 1237 2.50      
34 A 1198 1125 0.00      
35 A 1138 1069 0.19      
36 A 1123 1055 0.46      
37 A 977 917 0.35      
38 A 938 881 2.98      
39 A 824 774 0.12      
40 A 724 680 0.15      
41 A 400 375 0.30      
42 A 247 232 0.35      

Unscaled Zero Point Vibrational Energy (zpe) 32218.4 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 30259.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.13442 0.10037 0.06415

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 1.507 0.000
C2 0.070 -1.719 0.000
C3 0.386 -0.936 1.273
C4 0.386 -0.936 -1.273
C5 -0.389 0.371 1.358
C6 -0.389 0.371 -1.358
H7 0.630 -2.656 0.000
H8 -0.991 -1.987 0.000
H9 -1.464 0.173 1.366
H10 -1.464 0.173 -1.366
H11 0.144 -1.547 2.146
H12 0.144 -1.547 -2.146
H13 1.456 -0.720 1.314
H14 1.456 -0.720 -1.314
H15 -0.147 0.906 -2.276
H16 -0.147 0.906 2.276

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
S13.22642.78182.78181.81261.81264.21013.63212.40632.40633.73533.73532.96792.96792.35902.3590
C23.22641.52711.52712.53462.53461.09151.09482.79252.79252.15412.15412.15522.15523.48093.4809
C32.78181.52712.54541.52293.03862.15312.15002.15943.40861.09303.48131.09222.80754.03402.1644
C42.78181.52712.54543.03861.52292.15312.15003.40862.15943.48131.09302.80751.09222.16444.0340
C51.81262.53461.52293.03862.71633.47092.78751.09282.93552.14144.03032.14503.42623.68161.0898
C61.81262.53463.03861.52292.71633.47092.78752.93551.09284.03032.14143.42622.14501.08983.6816
H74.21011.09152.15312.15313.47093.47091.75333.77513.77512.46392.46392.48112.48114.29764.2976
H83.63211.09482.15002.15002.78752.78751.75332.59932.59932.46722.46723.05323.05323.77693.7769
H92.40632.79252.15943.40861.09282.93553.77512.59932.73272.48004.22843.05434.06333.94231.7608
H102.40632.79253.40862.15942.93551.09283.77512.59932.73274.22842.48004.06333.05431.76083.9423
H113.73532.15411.09303.48132.14144.03032.46392.46722.48004.22844.29211.76033.79185.06542.4733
H123.73532.15413.48131.09304.03032.14142.46392.46724.22842.48004.29213.79181.76032.47335.0654
H132.96792.15521.09222.80752.14503.42622.48113.05323.05434.06331.76033.79182.62804.25502.4780
H142.96792.15522.80751.09223.42622.14502.48113.05324.06333.05433.79181.76032.62802.47804.2550
H152.35903.48094.03402.16443.68161.08984.29763.77693.94231.76085.06542.47334.25502.47804.5528
H162.35903.48092.16444.03401.08983.68164.29763.77691.76083.94232.47335.06542.47804.25504.5528

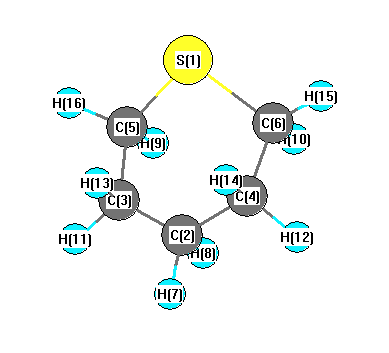
picture of 2H-Thiopyran, tetrahydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C5 C3 112.732 S1 C5 H9 109.318
S1 C5 H16 106.042 S1 C6 C4 112.732
S1 C6 H10 109.318 S1 C6 H15 106.042
C2 C3 C5 112.404 C2 C3 H11 109.488
C2 C3 H13 109.613 C2 C4 C6 112.404
C2 C4 H12 109.488 C2 C4 H14 109.613
C3 C2 C4 112.904 C3 C2 H7 109.493
C3 C2 H8 109.059 C3 C5 H9 110.200
C3 C5 H16 110.782 C4 C2 H7 109.493
C4 C2 H8 109.059 C4 C6 H10 110.200
C4 C6 H15 110.782 C5 S1 C6 97.053
C5 C3 H11 108.774 C5 C3 H13 109.099
C6 C4 H12 108.774 C6 C4 H14 109.099
H7 C2 H8 106.636 H9 C5 H16 107.562
H10 C6 H15 107.562 H11 C3 H13 107.329
H12 C4 H14 107.329
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability