return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H5Cl (1-chloro-1-propene(Z))

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-576.561463
Energy at 298.15K-576.566315
HF Energy-575.984045
Nuclear repulsion energy145.979000
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3309 3108 8.33      
2 A' 3251 3053 9.89      
3 A' 3239 3042 1.91      
4 A' 3122 2932 19.06      
5 A' 1715 1611 24.53      
6 A' 1534 1440 4.30      
7 A' 1462 1373 3.98      
8 A' 1395 1310 35.62      
9 A' 1289 1211 1.86      
10 A' 1127 1058 1.66      
11 A' 968 909 20.22      
12 A' 794 746 38.88      
13 A' 576 541 0.61      
14 A' 231 217 0.88      
15 A" 3200 3005 12.37      
16 A" 1531 1438 7.73      
17 A" 1085 1019 2.96      
18 A" 946 888 0.50      
19 A" 720 676 57.36      
20 A" 393 369 2.12      
21 A" 90 84 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 15987.0 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 15015.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.48145 0.11943 0.09741

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.915 0.000
C2 -1.244 0.423 0.000
C3 -1.643 -1.016 0.000
Cl4 1.436 -0.054 0.000
H5 0.207 1.974 0.000
H6 -2.035 1.163 0.000
H7 -0.774 -1.668 0.000
H8 -2.247 -1.245 0.878
H9 -2.247 -1.245 -0.878

Atom - Atom Distances (Å)
  C1 C2 C3 Cl4 H5 H6 H7 H8 H9
C11.33802.53571.73251.07872.05062.69643.23783.2378
C21.33801.49332.72242.12371.08392.14272.13472.1347
C32.53571.49333.22623.51592.21481.08591.08981.0898
Cl41.73252.72243.22622.37123.67892.73703.96913.9691
H51.07872.12373.51592.37122.38413.77134.14114.1411
H62.05061.08392.21483.67892.38413.09942.57202.5720
H72.69642.14271.08592.73703.77133.09941.76591.7659
H83.23782.13471.08983.96914.14112.57201.76591.7562
H93.23782.13471.08983.96914.14112.57201.76591.7562

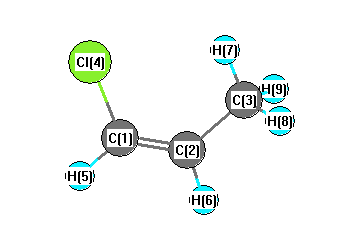
picture of 1-chloro-1-propene(Z) state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 127.080 C1 C2 H6 115.308
C2 C1 Cl4 124.412 C2 C1 H5 122.635
C2 C3 H7 111.373 C2 C3 H8 110.484
C2 C3 H9 110.484 C3 C2 H6 117.612
Cl4 C1 H5 112.953 H7 C3 H8 108.509
H7 C3 H9 108.509 H8 C3 H9 107.361
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability