Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -246.584192 |
| Energy at 298.15K | -246.589681 |
| HF Energy | -245.801353 |
| Nuclear repulsion energy | 156.489272 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3262 |
3063 |
13.07 |
|
|
|
| 2 |
A' |
3089 |
2901 |
55.91 |
|
|
|
| 3 |
A' |
3078 |
2891 |
23.32 |
|
|
|
| 4 |
A' |
2220 |
2085 |
8.03 |
|
|
|
| 5 |
A' |
1562 |
1467 |
4.41 |
|
|
|
| 6 |
A' |
1547 |
1453 |
4.85 |
|
|
|
| 7 |
A' |
1514 |
1422 |
0.50 |
|
|
|
| 8 |
A' |
1437 |
1349 |
40.97 |
|
|
|
| 9 |
A' |
1245 |
1169 |
45.29 |
|
|
|
| 10 |
A' |
1173 |
1102 |
127.83 |
|
|
|
| 11 |
A' |
999 |
938 |
32.58 |
|
|
|
| 12 |
A' |
967 |
908 |
10.30 |
|
|
|
| 13 |
A' |
533 |
501 |
0.83 |
|
|
|
| 14 |
A' |
376 |
353 |
1.57 |
|
|
|
| 15 |
A' |
173 |
163 |
2.02 |
|
|
|
| 16 |
A" |
3169 |
2977 |
38.93 |
|
|
|
| 17 |
A" |
3129 |
2938 |
23.43 |
|
|
|
| 18 |
A" |
1539 |
1445 |
8.04 |
|
|
|
| 19 |
A" |
1277 |
1199 |
3.34 |
|
|
|
| 20 |
A" |
1200 |
1127 |
4.92 |
|
|
|
| 21 |
A" |
1051 |
987 |
2.78 |
|
|
|
| 22 |
A" |
337 |
316 |
1.02 |
|
|
|
| 23 |
A" |
223 |
210 |
4.94 |
|
|
|
| 24 |
A" |
80 |
75 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17589.7 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 16520.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.138 |
-0.347 |
0.000 |
| O2 |
-0.737 |
-0.608 |
0.000 |
| C3 |
0.000 |
0.599 |
0.000 |
| C4 |
1.428 |
0.266 |
0.000 |
| N5 |
2.584 |
0.035 |
0.000 |
| H6 |
-2.626 |
-1.315 |
0.000 |
| H7 |
-2.430 |
0.213 |
0.892 |
| H8 |
-2.430 |
0.213 |
-0.892 |
| H9 |
-0.224 |
1.201 |
-0.887 |
| H10 |
-0.224 |
1.201 |
0.887 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4249 | 2.3378 | 3.6180 | 4.7376 | 1.0841 | 1.0926 | 1.0926 | 2.6164 | 2.6164 |
O2 | 1.4249 | | 1.4145 | 2.3348 | 3.3831 | 2.0169 | 2.0824 | 2.0824 | 2.0792 | 2.0792 | C3 | 2.3378 | 1.4145 | | 1.4662 | 2.6454 | 3.2495 | 2.6175 | 2.6175 | 1.0953 | 1.0953 | C4 | 3.6180 | 2.3348 | 1.4662 | | 1.1795 | 4.3513 | 3.9603 | 3.9603 | 2.0952 | 2.0952 | N5 | 4.7376 | 3.3831 | 2.6454 | 1.1795 | | 5.3824 | 5.0966 | 5.0966 | 3.1678 | 3.1678 | H6 | 1.0841 | 2.0169 | 3.2495 | 4.3513 | 5.3824 | | 1.7795 | 1.7795 | 3.5897 | 3.5897 | H7 | 1.0926 | 2.0824 | 2.6175 | 3.9603 | 5.0966 | 1.7795 | | 1.7834 | 3.0016 | 2.4175 | H8 | 1.0926 | 2.0824 | 2.6175 | 3.9603 | 5.0966 | 1.7795 | 1.7834 | | 2.4175 | 3.0016 | H9 | 2.6164 | 2.0792 | 1.0953 | 2.0952 | 3.1678 | 3.5897 | 3.0016 | 2.4175 | | 1.7747 | H10 | 2.6164 | 2.0792 | 1.0953 | 2.0952 | 3.1678 | 3.5897 | 2.4175 | 3.0016 | 1.7747 | |
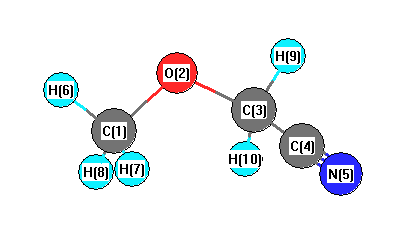 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.836 |
|
O2 |
C1 |
H6 |
106.199 |
| O2 |
C1 |
H7 |
110.924 |
|
O2 |
C1 |
H8 |
110.924 |
| O2 |
C3 |
C4 |
108.276 |
|
O2 |
C3 |
H9 |
111.242 |
| O2 |
C3 |
H10 |
111.242 |
|
C3 |
C4 |
N5 |
178.193 |
| C4 |
C3 |
H9 |
108.902 |
|
C4 |
C3 |
H10 |
108.902 |
| H6 |
C1 |
H7 |
109.672 |
|
H6 |
C1 |
H8 |
109.672 |
| H7 |
C1 |
H8 |
109.398 |
|
H9 |
C3 |
H10 |
108.226 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -246.586905 |
| Energy at 298.15K | -246.592541 |
| HF Energy | -245.802987 |
| Nuclear repulsion energy | 159.551045 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3262 |
3064 |
13.49 |
|
|
|
| 2 |
A |
3216 |
3021 |
3.11 |
|
|
|
| 3 |
A |
3183 |
2990 |
29.85 |
|
|
|
| 4 |
A |
3099 |
2911 |
28.26 |
|
|
|
| 5 |
A |
3096 |
2908 |
38.45 |
|
|
|
| 6 |
A |
2202 |
2069 |
7.82 |
|
|
|
| 7 |
A |
1558 |
1464 |
9.37 |
|
|
|
| 8 |
A |
1543 |
1449 |
7.25 |
|
|
|
| 9 |
A |
1527 |
1434 |
3.10 |
|
|
|
| 10 |
A |
1515 |
1423 |
0.30 |
|
|
|
| 11 |
A |
1414 |
1328 |
18.61 |
|
|
|
| 12 |
A |
1329 |
1248 |
5.62 |
|
|
|
| 13 |
A |
1238 |
1162 |
40.39 |
|
|
|
| 14 |
A |
1205 |
1131 |
8.24 |
|
|
|
| 15 |
A |
1173 |
1102 |
103.62 |
|
|
|
| 16 |
A |
1048 |
984 |
7.13 |
|
|
|
| 17 |
A |
954 |
896 |
38.79 |
|
|
|
| 18 |
A |
915 |
860 |
16.08 |
|
|
|
| 19 |
A |
591 |
555 |
2.23 |
|
|
|
| 20 |
A |
392 |
368 |
3.24 |
|
|
|
| 21 |
A |
337 |
316 |
0.88 |
|
|
|
| 22 |
A |
255 |
240 |
10.65 |
|
|
|
| 23 |
A |
174 |
163 |
4.73 |
|
|
|
| 24 |
A |
118 |
111 |
9.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17672.1 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 16597.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.534 |
-0.796 |
0.139 |
| O2 |
1.094 |
0.435 |
-0.435 |
| C3 |
-0.052 |
0.949 |
0.209 |
| C4 |
-1.229 |
0.071 |
0.038 |
| N5 |
-2.158 |
-0.644 |
-0.099 |
| H6 |
2.450 |
-1.058 |
-0.380 |
| H7 |
0.793 |
-1.584 |
-0.004 |
| H8 |
1.736 |
-0.674 |
1.205 |
| H9 |
-0.259 |
1.915 |
-0.243 |
| H10 |
0.121 |
1.088 |
1.281 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4280 | 2.3593 | 2.8975 | 3.7032 | 1.0842 | 1.0910 | 1.0926 | 3.2732 | 2.6179 |
O2 | 1.4280 | | 1.4112 | 2.3978 | 3.4426 | 2.0177 | 2.0862 | 2.0815 | 2.0142 | 2.0779 | C3 | 2.3593 | 1.4112 | | 1.4780 | 2.6582 | 3.2608 | 2.6784 | 2.6122 | 1.0871 | 1.0950 | C4 | 2.8975 | 2.3978 | 1.4780 | | 1.1803 | 3.8703 | 2.6128 | 3.2726 | 2.1028 | 2.0986 | N5 | 3.7032 | 3.4426 | 2.6582 | 1.1803 | | 4.6352 | 3.0988 | 4.1073 | 3.1900 | 3.1781 | H6 | 1.0842 | 2.0177 | 3.2608 | 3.8703 | 4.6352 | | 1.7784 | 1.7800 | 4.0244 | 3.5758 | H7 | 1.0910 | 2.0862 | 2.6784 | 2.6128 | 3.0988 | 1.7784 | | 1.7838 | 3.6617 | 3.0404 | H8 | 1.0926 | 2.0815 | 2.6122 | 3.2726 | 4.1073 | 1.7800 | 1.7838 | | 3.5753 | 2.3914 | H9 | 3.2732 | 2.0142 | 1.0871 | 2.1028 | 3.1900 | 4.0244 | 3.6617 | 3.5753 | | 1.7758 | H10 | 2.6179 | 2.0779 | 1.0950 | 2.0986 | 3.1781 | 3.5758 | 3.0404 | 2.3914 | 1.7758 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.392 |
|
O2 |
C1 |
H6 |
106.058 |
| O2 |
C1 |
H7 |
111.120 |
|
O2 |
C1 |
H8 |
110.636 |
| O2 |
C3 |
C4 |
112.161 |
|
O2 |
C3 |
H9 |
106.737 |
| O2 |
C3 |
H10 |
111.385 |
|
C3 |
C4 |
N5 |
179.191 |
| C4 |
C3 |
H9 |
109.172 |
|
C4 |
C3 |
H10 |
108.385 |
| H6 |
C1 |
H7 |
109.692 |
|
H6 |
C1 |
H8 |
109.714 |
| H7 |
C1 |
H8 |
109.561 |
|
H9 |
C3 |
H10 |
108.936 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
