Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Geometric Data calculated at MP2=FULL/6-31+G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -235.096462 |
| Energy at 298.15K | -235.110083 |
| Nuclear repulsion energy | 256.121651 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3216 |
3021 |
53.39 |
|
|
|
| 2 |
A' |
3198 |
3004 |
40.42 |
|
|
|
| 3 |
A' |
3195 |
3001 |
25.77 |
|
|
|
| 4 |
A' |
3184 |
2990 |
18.16 |
|
|
|
| 5 |
A' |
3148 |
2957 |
35.70 |
|
|
|
| 6 |
A' |
3122 |
2932 |
7.74 |
|
|
|
| 7 |
A' |
3098 |
2910 |
33.80 |
|
|
|
| 8 |
A' |
3093 |
2905 |
32.53 |
|
|
|
| 9 |
A' |
1556 |
1462 |
2.21 |
|
|
|
| 10 |
A' |
1546 |
1452 |
2.74 |
|
|
|
| 11 |
A' |
1536 |
1442 |
7.80 |
|
|
|
| 12 |
A' |
1522 |
1430 |
0.04 |
|
|
|
| 13 |
A' |
1453 |
1364 |
2.58 |
|
|
|
| 14 |
A' |
1438 |
1351 |
11.45 |
|
|
|
| 15 |
A' |
1353 |
1270 |
6.10 |
|
|
|
| 16 |
A' |
1339 |
1258 |
2.83 |
|
|
|
| 17 |
A' |
1267 |
1190 |
0.44 |
|
|
|
| 18 |
A' |
1233 |
1158 |
0.26 |
|
|
|
| 19 |
A' |
1043 |
980 |
0.29 |
|
|
|
| 20 |
A' |
1019 |
957 |
0.65 |
|
|
|
| 21 |
A' |
981 |
921 |
0.15 |
|
|
|
| 22 |
A' |
952 |
895 |
0.35 |
|
|
|
| 23 |
A' |
900 |
845 |
0.29 |
|
|
|
| 24 |
A' |
726 |
682 |
0.78 |
|
|
|
| 25 |
A' |
612 |
575 |
1.13 |
|
|
|
| 26 |
A' |
405 |
381 |
0.71 |
|
|
|
| 27 |
A' |
359 |
337 |
0.02 |
|
|
|
| 28 |
A' |
187 |
175 |
0.00 |
|
|
|
| 29 |
A" |
3192 |
2998 |
53.01 |
|
|
|
| 30 |
A" |
3188 |
2994 |
0.01 |
|
|
|
| 31 |
A" |
3185 |
2991 |
15.65 |
|
|
|
| 32 |
A" |
3119 |
2929 |
76.12 |
|
|
|
| 33 |
A" |
1546 |
1452 |
8.40 |
|
|
|
| 34 |
A" |
1524 |
1432 |
0.55 |
|
|
|
| 35 |
A" |
1517 |
1425 |
3.93 |
|
|
|
| 36 |
A" |
1305 |
1226 |
0.63 |
|
|
|
| 37 |
A" |
1279 |
1201 |
0.34 |
|
|
|
| 38 |
A" |
1253 |
1176 |
0.03 |
|
|
|
| 39 |
A" |
1211 |
1137 |
1.41 |
|
|
|
| 40 |
A" |
1106 |
1039 |
0.04 |
|
|
|
| 41 |
A" |
1007 |
946 |
0.01 |
|
|
|
| 42 |
A" |
968 |
909 |
2.54 |
|
|
|
| 43 |
A" |
910 |
854 |
0.03 |
|
|
|
| 44 |
A" |
809 |
760 |
0.12 |
|
|
|
| 45 |
A" |
388 |
365 |
0.00 |
|
|
|
| 46 |
A" |
304 |
285 |
0.01 |
|
|
|
| 47 |
A" |
276 |
260 |
0.00 |
|
|
|
| 48 |
A" |
234 |
220 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37500.2 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 35220.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.106 |
0.400 |
0.000 |
| C2 |
0.396 |
-1.708 |
0.000 |
| C3 |
-0.106 |
-0.719 |
1.074 |
| C4 |
-0.106 |
-0.719 |
-1.074 |
| C5 |
1.205 |
1.174 |
0.000 |
| C6 |
-1.291 |
1.348 |
0.000 |
| H7 |
1.482 |
-1.797 |
0.000 |
| H8 |
-0.031 |
-2.710 |
0.000 |
| H9 |
0.509 |
-0.561 |
1.961 |
| H10 |
0.509 |
-0.561 |
-1.961 |
| H11 |
-1.124 |
-0.959 |
1.388 |
| H12 |
-1.124 |
-0.959 |
-1.388 |
| H13 |
2.065 |
0.504 |
0.000 |
| H14 |
-2.230 |
0.794 |
0.000 |
| H15 |
1.274 |
1.810 |
-0.884 |
| H16 |
1.274 |
1.810 |
0.884 |
| H17 |
-1.276 |
1.989 |
0.883 |
| H18 |
-1.276 |
1.989 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1677 | 1.5511 | 1.5511 | 1.5226 | 1.5169 | 2.7105 | 3.1106 | 2.2686 | 2.2686 | 2.1930 | 2.1930 | 2.1742 | 2.1599 | 2.1622 | 2.1622 | 2.1617 | 2.1617 |
C2 | 2.1677 | | 1.5442 | 1.5442 | 2.9937 | 3.4909 | 1.0888 | 1.0887 | 2.2745 | 2.2745 | 2.1905 | 2.1905 | 2.7715 | 3.6274 | 3.7327 | 3.7327 | 4.1533 | 4.1533 | C3 | 1.5511 | 1.5442 | | 2.1476 | 2.5410 | 2.6130 | 2.1990 | 2.2628 | 1.0911 | 3.1002 | 1.0913 | 2.6746 | 2.7140 | 2.8198 | 3.4837 | 2.8877 | 2.9564 | 3.5402 | C4 | 1.5511 | 1.5442 | 2.1476 | | 2.5410 | 2.6130 | 2.1990 | 2.2628 | 3.1002 | 1.0911 | 2.6746 | 1.0913 | 2.7140 | 2.8198 | 2.8877 | 3.4837 | 3.5402 | 2.9564 | C5 | 1.5226 | 2.9937 | 2.5410 | 2.5410 | | 2.5019 | 2.9834 | 4.0756 | 2.7088 | 2.7088 | 3.4495 | 3.4495 | 1.0903 | 3.4560 | 1.0914 | 1.0914 | 2.7564 | 2.7564 | C6 | 1.5169 | 3.4909 | 2.6130 | 2.6130 | 2.5019 | | 4.1920 | 4.2482 | 3.2753 | 3.2753 | 2.6975 | 2.6975 | 3.4606 | 1.0905 | 2.7520 | 2.7520 | 1.0918 | 1.0918 | H7 | 2.7105 | 1.0888 | 2.1990 | 2.1990 | 2.9834 | 4.1920 | | 1.7671 | 2.5131 | 2.5131 | 3.0683 | 3.0683 | 2.3737 | 4.5260 | 3.7194 | 3.7194 | 4.7661 | 4.7661 | H8 | 3.1106 | 1.0887 | 2.2628 | 2.2628 | 4.0756 | 4.2482 | 1.7671 | | 2.9585 | 2.9585 | 2.4867 | 2.4867 | 3.8373 | 4.1359 | 4.7869 | 4.7869 | 4.9405 | 4.9405 | H9 | 2.2686 | 2.2745 | 1.0911 | 3.1002 | 2.7088 | 3.2753 | 2.5131 | 2.9585 | | 3.9212 | 1.7755 | 3.7469 | 2.7202 | 3.6308 | 3.7814 | 2.7140 | 3.2941 | 4.2163 | H10 | 2.2686 | 2.2745 | 3.1002 | 1.0911 | 2.7088 | 3.2753 | 2.5131 | 2.9585 | 3.9212 | | 3.7469 | 1.7755 | 2.7202 | 3.6308 | 2.7140 | 3.7814 | 4.2163 | 3.2941 | H11 | 2.1930 | 2.1905 | 1.0913 | 2.6746 | 3.4495 | 2.6975 | 3.0683 | 2.4867 | 1.7755 | 3.7469 | | 2.7764 | 3.7732 | 2.4947 | 4.3105 | 3.6975 | 2.9955 | 3.7251 | H12 | 2.1930 | 2.1905 | 2.6746 | 1.0913 | 3.4495 | 2.6975 | 3.0683 | 2.4867 | 3.7469 | 1.7755 | 2.7764 | | 3.7732 | 2.4947 | 3.6975 | 4.3105 | 3.7251 | 2.9955 | H13 | 2.1742 | 2.7715 | 2.7140 | 2.7140 | 1.0903 | 3.4606 | 2.3737 | 3.8373 | 2.7202 | 2.7202 | 3.7732 | 3.7732 | | 4.3051 | 1.7646 | 1.7646 | 3.7613 | 3.7613 | H14 | 2.1599 | 3.6274 | 2.8198 | 2.8198 | 3.4560 | 1.0905 | 4.5260 | 4.1359 | 3.6308 | 3.6308 | 2.4947 | 2.4947 | 4.3051 | | 3.7541 | 3.7541 | 1.7667 | 1.7667 | H15 | 2.1622 | 3.7327 | 3.4837 | 2.8877 | 1.0914 | 2.7520 | 3.7194 | 4.7869 | 3.7814 | 2.7140 | 4.3105 | 3.6975 | 1.7646 | 3.7541 | | 1.7682 | 3.1072 | 2.5558 | H16 | 2.1622 | 3.7327 | 2.8877 | 3.4837 | 1.0914 | 2.7520 | 3.7194 | 4.7869 | 2.7140 | 3.7814 | 3.6975 | 4.3105 | 1.7646 | 3.7541 | 1.7682 | | 2.5558 | 3.1072 | H17 | 2.1617 | 4.1533 | 2.9564 | 3.5402 | 2.7564 | 1.0918 | 4.7661 | 4.9405 | 3.2941 | 4.2163 | 2.9955 | 3.7251 | 3.7613 | 1.7667 | 3.1072 | 2.5558 | | 1.7663 | H18 | 2.1617 | 4.1533 | 3.5402 | 2.9564 | 2.7564 | 1.0918 | 4.7661 | 4.9405 | 4.2163 | 3.2941 | 3.7251 | 2.9955 | 3.7613 | 1.7667 | 2.5558 | 3.1072 | 1.7663 | |
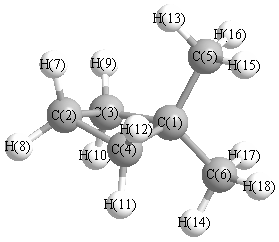 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.906 |
|
C1 |
C3 |
H9 |
117.259 |
| C1 |
C3 |
H11 |
110.987 |
|
C1 |
C4 |
C2 |
88.906 |
| C1 |
C4 |
H10 |
117.259 |
|
C1 |
C4 |
H12 |
110.987 |
| C1 |
C5 |
H13 |
111.553 |
|
C1 |
C5 |
H15 |
110.529 |
| C1 |
C5 |
H16 |
110.529 |
|
C1 |
C6 |
H14 |
110.804 |
| C1 |
C6 |
H17 |
110.870 |
|
C1 |
C6 |
H18 |
110.870 |
| C2 |
C3 |
H9 |
118.318 |
|
C2 |
C3 |
H11 |
111.276 |
| C2 |
C4 |
H10 |
118.318 |
|
C2 |
C4 |
H12 |
111.276 |
| C3 |
C1 |
C4 |
87.623 |
|
C3 |
C1 |
C5 |
111.516 |
| C3 |
C1 |
C6 |
116.792 |
|
C3 |
C2 |
C4 |
88.116 |
| C3 |
C2 |
H7 |
112.108 |
|
C3 |
C2 |
H8 |
117.465 |
| C4 |
C1 |
C5 |
111.516 |
|
C4 |
C1 |
C6 |
116.792 |
| C4 |
C2 |
H7 |
112.108 |
|
C4 |
C2 |
H8 |
117.465 |
| C5 |
C1 |
C6 |
110.797 |
|
H7 |
C2 |
H8 |
108.494 |
| H9 |
C3 |
H11 |
108.885 |
|
H10 |
C4 |
H12 |
108.885 |
| H13 |
C5 |
H15 |
107.955 |
|
H13 |
C5 |
H16 |
107.955 |
| H14 |
C6 |
H17 |
108.106 |
|
H14 |
C6 |
H18 |
108.106 |
| H15 |
C5 |
H16 |
108.198 |
|
H17 |
C6 |
H18 |
107.968 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
