Jump to
S1C2
S1C3
Energy calculated at CCSD/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -1590.512874 |
| Energy at 298.15K | |
| HF Energy | -1589.981141 |
| Nuclear repulsion energy | 347.277830 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
502 |
469 |
0.00 |
|
|
|
| 2 |
A1 |
204 |
191 |
0.00 |
|
|
|
| 3 |
B1 |
450 |
421 |
0.00 |
|
|
|
| 4 |
B2 |
334 |
312 |
0.13 |
|
|
|
| 5 |
E |
429 |
400 |
0.01 |
|
|
|
| 5 |
E |
429 |
400 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1173.5 cm
-1
Scaled (by 0.9338) Zero Point Vibrational Energy (zpe) 1095.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
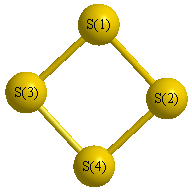
Geometric Data calculated at CCSD/6-31+G**
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.445 |
0.312 |
| S2 |
0.000 |
-1.445 |
0.312 |
| S3 |
-1.445 |
0.000 |
-0.312 |
| S4 |
1.445 |
0.000 |
-0.312 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8906 | 2.1372 | 2.1372 |
S2 | 2.8906 | | 2.1372 | 2.1372 | S3 | 2.1372 | 2.1372 | | 2.8906 | S4 | 2.1372 | 2.1372 | 2.8906 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.448 |
|
S1 |
S3 |
S4 |
47.448 |
| S2 |
S1 |
S3 |
47.448 |
|
S2 |
S4 |
S3 |
47.448 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCSD/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -1590.526064 |
| Energy at 298.15K | |
| HF Energy | -1589.954006 |
| Nuclear repulsion energy | 329.275714 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
671 |
626 |
1.36 |
|
|
|
| 2 |
A1 |
381 |
356 |
8.28 |
|
|
|
| 3 |
A1 |
130 |
121 |
0.56 |
|
|
|
| 4 |
A2 |
194 |
181 |
|
|
|
|
| 5 |
B2 |
656 |
612 |
121.41 |
|
|
|
| 6 |
B2 |
329 |
307 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1179.9 cm
-1
Scaled (by 0.9338) Zero Point Vibrational Energy (zpe) 1101.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.060 |
0.923 |
| S2 |
0.000 |
-1.060 |
0.923 |
| S3 |
0.000 |
1.622 |
-0.923 |
| S4 |
0.000 |
-1.622 |
-0.923 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.1207 | 1.9292 | 3.2562 |
S2 | 2.1207 | | 3.2562 | 1.9292 | S3 | 1.9292 | 3.2562 | | 3.2449 | S4 | 3.2562 | 1.9292 | 3.2449 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
106.940 |
|
S1 |
S3 |
S4 |
73.060 |
| S2 |
S1 |
S3 |
106.940 |
|
S2 |
S4 |
S3 |
73.060 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCSD/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -1590.519284 |
| Energy at 298.15K | |
| HF Energy | -1589.925650 |
| Nuclear repulsion energy | 330.604943 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
739 |
690 |
0.00 |
|
|
|
| 2 |
Ag |
277 |
258 |
0.00 |
|
|
|
| 3 |
Au |
230 |
215 |
0.00 |
|
|
|
| 4 |
B1u |
304i |
284i |
14051.00 |
|
|
|
| 5 |
B2u |
144i |
135i |
20.48 |
|
|
|
| 6 |
B3g |
321 |
300 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 559.4 cm
-1
Scaled (by 0.9338) Zero Point Vibrational Energy (zpe) 522.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-31+G**
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.952 |
1.299 |
| S2 |
0.000 |
0.952 |
-1.299 |
| S3 |
0.000 |
-0.952 |
1.299 |
| S4 |
0.000 |
-0.952 |
-1.299 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5985 | 1.9047 | 3.2219 |
S2 | 2.5985 | | 3.2219 | 1.9047 | S3 | 1.9047 | 3.2219 | | 2.5985 | S4 | 3.2219 | 1.9047 | 2.5985 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
