return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCH3 (Isopropyl radical)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 2A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-118.082376
Energy at 298.15K-118.088702
HF Energy-117.649163
Nuclear repulsion energy75.661149
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3280 3080 27.32      
2 A' 3213 3018 18.85      
3 A' 3154 2962 41.46      
4 A' 3067 2881 34.87      
5 A' 1543 1449 5.58      
6 A' 1529 1436 16.02      
7 A' 1463 1374 4.17      
8 A' 1219 1145 1.61      
9 A' 1074 1009 0.28      
10 A' 918 862 1.59      
11 A' 441 414 25.89      
12 A' 365 342 7.16      
13 A' 168 158 0.22      
14 A" 3214 3019 20.02      
15 A" 3155 2963 5.46      
16 A" 3066 2879 28.38      
17 A" 1532 1439 1.08      
18 A" 1519 1426 1.61      
19 A" 1468 1379 9.76      
20 A" 1413 1327 1.98      
21 A" 1191 1118 0.09      
22 A" 968 909 0.90      
23 A" 963 905 0.26      
24 A" 129 122 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 20024.5 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 18807.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
1.24175 0.28115 0.25089

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.014 0.546 0.000
C2 -0.014 -0.200 1.289
C3 -0.014 -0.200 -1.289
H4 0.317 1.575 0.000
H5 -0.775 -0.983 1.287
H6 -0.775 -0.983 -1.287
H7 0.947 -0.697 1.468
H8 -0.203 0.456 2.137
H9 0.947 -0.697 -1.468
H10 -0.203 0.456 -2.137

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10
C11.48991.48991.08092.13782.13782.15062.14692.15062.1469
C21.48992.57892.21901.09102.79751.09661.08812.96223.4935
C31.48992.57892.21902.79751.09102.96223.49351.09661.0881
H41.08092.21902.21903.06393.06392.77782.46752.77782.4675
H52.13781.09102.79753.06392.57311.75431.76573.26083.7568
H62.13782.79751.09103.06392.57313.26083.75681.75431.7657
H72.15061.09662.96222.77781.75433.26081.76062.93633.9557
H82.14691.08813.49352.46751.76573.75681.76063.95574.2733
H92.15062.96221.09662.77783.26081.75432.93633.95571.7606
H102.14693.49351.08812.46753.75681.76573.95574.27331.7606

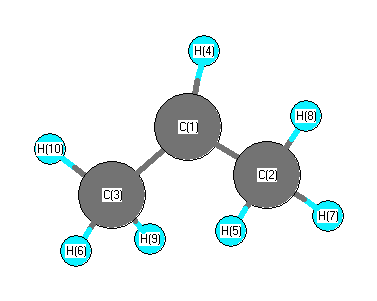
picture of Isopropyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H5 110.907 C1 C2 H7 111.600
C1 C2 H8 111.826 C1 C3 H6 110.907
C1 C3 H9 111.600 C1 C3 H10 111.826
C2 C1 C3 119.872 C2 C1 H4 118.483
C3 C1 H4 118.483 H5 C2 H7 106.634
H5 C2 H8 108.251 H6 C3 H9 106.634
H6 C3 H10 108.251 H7 C2 H8 107.391
H9 C3 H10 107.391
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability