Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Geometric Data calculated at MP2=FULL/6-31+G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -192.867108 |
| Energy at 298.15K | -192.871920 |
| HF Energy | -192.207490 |
| Nuclear repulsion energy | 149.245878 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3346 |
3143 |
0.31 |
|
|
|
| 2 |
A1 |
3332 |
3129 |
3.99 |
|
|
|
| 3 |
A1 |
3306 |
3105 |
0.92 |
|
|
|
| 4 |
A1 |
1595 |
1498 |
2.92 |
|
|
|
| 5 |
A1 |
1501 |
1410 |
5.81 |
|
|
|
| 6 |
A1 |
1190 |
1118 |
3.10 |
|
|
|
| 7 |
A1 |
1119 |
1051 |
11.44 |
|
|
|
| 8 |
A1 |
1080 |
1014 |
15.85 |
|
|
|
| 9 |
A1 |
859 |
807 |
0.27 |
|
|
|
| 10 |
A2 |
852 |
800 |
0.00 |
|
|
|
| 11 |
A2 |
828 |
777 |
0.00 |
|
|
|
| 12 |
A2 |
502 |
471 |
0.00 |
|
|
|
| 13 |
B1 |
885 |
831 |
8.18 |
|
|
|
| 14 |
B1 |
737 |
692 |
52.81 |
|
|
|
| 15 |
B1 |
675 |
634 |
95.71 |
|
|
|
| 16 |
B1 |
486 |
457 |
12.84 |
|
|
|
| 17 |
B2 |
6100 |
5729 |
93141.66 |
|
|
|
| 18 |
B2 |
3313 |
3112 |
76.22 |
|
|
|
| 19 |
B2 |
3304 |
3103 |
11.66 |
|
|
|
| 20 |
B2 |
1413 |
1327 |
101.55 |
|
|
|
| 21 |
B2 |
1333 |
1252 |
0.46 |
|
|
|
| 22 |
B2 |
1190 |
1117 |
15.64 |
|
|
|
| 23 |
B2 |
980 |
921 |
5.15 |
|
|
|
| 24 |
B2 |
869 |
817 |
9.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20397.4 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 19157.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.218 |
| C2 |
0.000 |
1.117 |
0.392 |
| C3 |
0.000 |
0.685 |
-1.001 |
| C4 |
0.000 |
-0.685 |
-1.001 |
| C5 |
0.000 |
-1.117 |
0.392 |
| H6 |
0.000 |
0.000 |
2.294 |
| H7 |
0.000 |
2.146 |
0.717 |
| H8 |
0.000 |
1.338 |
-1.859 |
| H9 |
0.000 |
-1.338 |
-1.859 |
| H10 |
0.000 |
-2.146 |
0.717 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3894 | 2.3223 | 2.3223 | 1.3894 | 1.0764 | 2.2039 | 3.3551 | 3.3551 | 2.2039 |
C2 | 1.3894 | | 1.4581 | 2.2777 | 2.2342 | 2.2063 | 1.0794 | 2.2613 | 3.3305 | 3.2796 | C3 | 2.3223 | 1.4581 | | 1.3705 | 2.2777 | 3.3658 | 2.2554 | 1.0779 | 2.1976 | 3.3121 | C4 | 2.3223 | 2.2777 | 1.3705 | | 1.4581 | 3.3658 | 3.3121 | 2.1976 | 1.0779 | 2.2554 | C5 | 1.3894 | 2.2342 | 2.2777 | 1.4581 | | 2.2063 | 3.2796 | 3.3305 | 2.2613 | 1.0794 | H6 | 1.0764 | 2.2063 | 3.3658 | 3.3658 | 2.2063 | | 2.6634 | 4.3633 | 4.3633 | 2.6634 | H7 | 2.2039 | 1.0794 | 2.2554 | 3.3121 | 3.2796 | 2.6634 | | 2.6998 | 4.3332 | 4.2926 | H8 | 3.3551 | 2.2613 | 1.0779 | 2.1976 | 3.3305 | 4.3633 | 2.6998 | | 2.6761 | 4.3332 | H9 | 3.3551 | 3.3305 | 2.1976 | 1.0779 | 2.2613 | 4.3633 | 4.3332 | 2.6761 | | 2.6998 | H10 | 2.2039 | 3.2796 | 3.3121 | 2.2554 | 1.0794 | 2.6634 | 4.2926 | 4.3332 | 2.6998 | |
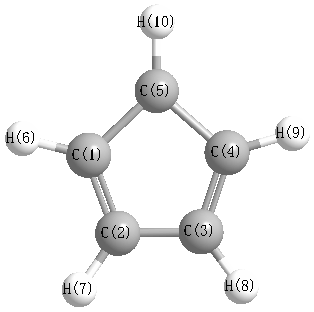 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
109.261 |
|
C1 |
C2 |
H7 |
125.964 |
| C1 |
C5 |
C4 |
109.261 |
|
C1 |
C5 |
H10 |
125.964 |
| C2 |
C1 |
C5 |
107.025 |
|
C2 |
C1 |
H6 |
126.488 |
| C2 |
C3 |
C4 |
107.227 |
|
C2 |
C3 |
H8 |
125.500 |
| C3 |
C2 |
H7 |
124.775 |
|
C3 |
C4 |
C5 |
107.227 |
| C3 |
C4 |
H9 |
127.273 |
|
C4 |
C3 |
H8 |
127.273 |
| C4 |
C5 |
H10 |
124.775 |
|
C5 |
C1 |
H6 |
126.488 |
| C5 |
C4 |
H9 |
125.500 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
