Jump to
S2C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -77.019473 |
| Energy at 298.15K | -77.019147 |
| HF Energy | -76.773374 |
| Nuclear repulsion energy | 23.789184 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3220 |
3025 |
51.16 |
|
|
|
| 2 |
A1 |
1709 |
1605 |
87.58 |
|
|
|
| 3 |
A1 |
1258 |
1182 |
18.13 |
|
|
|
| 4 |
B1 |
753 |
707 |
112.70 |
|
|
|
| 5 |
B2 |
3323 |
3121 |
21.91 |
|
|
|
| 6 |
B2 |
325 |
306 |
7.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5294.7 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 4972.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.482 |
| C2 |
0.000 |
0.000 |
0.824 |
| H3 |
0.000 |
0.938 |
-1.024 |
| H4 |
0.000 |
-0.938 |
-1.024 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3063 | 1.0836 | 1.0836 |
C2 | 1.3063 | | 2.0728 | 2.0728 | H3 | 1.0836 | 2.0728 | | 1.8765 | H4 | 1.0836 | 2.0728 | 1.8765 | |
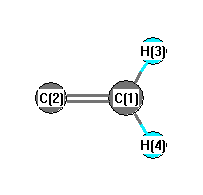 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C2 |
C1 |
H3 |
120.015 |
|
C2 |
C1 |
H4 |
120.015 |
| H3 |
C1 |
H4 |
119.971 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -76.943257 |
| Energy at 298.15K | -76.943382 |
| HF Energy | -76.740255 |
| Nuclear repulsion energy | 23.760310 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3190 |
2996 |
2.64 |
|
|
|
| 2 |
A1 |
1642 |
1542 |
2.10 |
|
|
|
| 3 |
A1 |
1464 |
1375 |
4.49 |
|
|
|
| 4 |
B1 |
953 |
895 |
36.06 |
|
|
|
| 5 |
B2 |
3285 |
3085 |
2.72 |
|
|
|
| 6 |
B2 |
1042 |
979 |
4.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5787.6 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 5435.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.480 |
| C2 |
0.000 |
0.000 |
0.827 |
| H3 |
0.000 |
0.927 |
-1.041 |
| H4 |
0.000 |
-0.927 |
-1.041 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3073 | 1.0839 | 1.0839 |
C2 | 1.3073 | | 2.0858 | 2.0858 | H3 | 1.0839 | 2.0858 | | 1.8550 | H4 | 1.0839 | 2.0858 | 1.8550 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
