return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2 (benzyl radical)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2B1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-270.061023
Energy at 298.15K-270.067782
HF Energy-269.157909
Nuclear repulsion energy262.251481
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3297 3096 11.69      
2 A1 3277 3078 0.91      
3 A1 3261 3063 9.26      
4 A1 3255 3058 5.16      
5 A1 1732 1627 0.69      
6 A1 1565 1470 7.57      
7 A1 1540 1447 1.92      
8 A1 1296 1217 3.84      
9 A1 1216 1142 1.10      
10 A1 1094 1028 2.91      
11 A1 1000 940 0.15      
12 A1 866 814 0.82      
13 A1 549 515 0.58      
14 A2 1041 978 0.00      
15 A2 916 861 0.00      
16 A2 478 449 0.00      
17 A2 403 379 0.00      
18 B1 1009 947 1.23      
19 B1 957 898 1.22      
20 B1 782 734 120.64      
21 B1 638 599 46.83      
22 B1 465 436 11.92      
23 B1 387 364 2.39      
24 B1 198 186 2.43      
25 B2 3371 3166 4.62      
26 B2 3283 3083 25.90      
27 B2 3263 3065 1.66      
28 B2 1713 1609 0.32      
29 B2 1557 1462 6.34      
30 B2 1394 1309 0.40      
31 B2 1306 1227 0.22      
32 B2 1184 1112 2.70      
33 B2 1174 1103 3.77      
34 B2 1013 951 2.89      
35 B2 651 611 0.00      
36 B2 362 340 0.22      

Unscaled Zero Point Vibrational Energy (zpe) 25746.1 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 24180.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.18863 0.09171 0.06171

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.976
C2 0.000 1.204 0.246
C3 0.000 1.198 -1.121
C4 0.000 0.000 -1.820
C5 0.000 -1.198 -1.121
C6 0.000 -1.204 0.246
C7 0.000 0.000 2.387
H8 0.000 2.142 0.786
H9 0.000 2.135 -1.661
H10 0.000 0.000 -2.900
H11 0.000 -2.135 -1.661
H12 0.000 -2.142 0.786
H13 0.000 0.925 2.942
H14 0.000 -0.925 2.942

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14
C11.40762.41442.79552.41441.40761.41112.15033.39233.87603.39232.15032.17252.1725
C21.40761.36692.39122.76332.40772.45581.08242.12203.36903.84493.38912.70963.4345
C32.41441.36691.38692.39552.76333.70632.12771.08162.14523.37613.84574.07134.5832
C42.79552.39121.38691.38692.39124.20663.37332.14081.08052.14083.37334.85034.8503
C52.41442.76332.39551.38691.36693.70633.84573.37612.14521.08162.12774.58324.0713
C61.40762.40772.76332.39121.36692.45583.38913.84493.36902.12201.08243.43452.7096
C71.41112.45583.70634.20663.70632.45582.67394.57605.28714.57602.67391.07841.0784
H82.15031.08242.12773.37333.84573.38912.67392.44684.26364.92734.28392.47523.7484
H93.39232.12201.08162.14083.37613.84494.57602.44682.46874.26984.92734.75855.5264
H103.87603.36902.14521.08052.14523.36905.28714.26362.46872.46874.26365.91465.9146
H113.39233.84493.37612.14081.08162.12204.57604.92734.26982.46872.44685.52644.7585
H122.15033.38913.84573.37332.12771.08242.67394.28394.92734.26362.44683.74842.4752
H132.17252.70964.07134.85034.58323.43451.07842.47524.75855.91465.52643.74841.8497
H142.17253.43454.58324.85034.07132.70961.07843.74845.52645.91464.75852.47521.8497

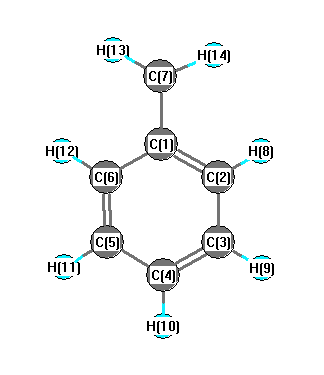
picture of benzyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.958 C1 C2 H8 118.866
C1 C6 C5 120.958 C1 C6 H12 118.866
C1 C7 H13 120.955 C1 C7 H14 120.955
C2 C1 C6 117.573 C2 C1 C7 121.213
C2 C3 C4 120.530 C2 C3 H9 119.696
C3 C2 H8 120.177 C3 C4 C5 119.452
C3 C4 H10 120.274 C4 C3 H9 119.774
C4 C5 C6 120.530 C4 C5 H11 119.774
C5 C1 C7 150.258 C5 C4 H10 120.274
C5 C6 H12 120.177 C6 C5 H11 119.696
H13 C7 H14 118.091
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability