Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Geometric Data calculated at MP2=FULL/6-31+G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -235.093724 |
| Energy at 298.15K | -235.107272 |
| HF Energy | -234.190228 |
| Nuclear repulsion energy | 250.805558 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3201 |
3007 |
65.95 |
|
|
|
| 2 |
A' |
3197 |
3003 |
31.64 |
|
|
|
| 3 |
A' |
3186 |
2992 |
36.94 |
|
|
|
| 4 |
A' |
3139 |
2948 |
24.76 |
|
|
|
| 5 |
A' |
3133 |
2943 |
19.36 |
|
|
|
| 6 |
A' |
3117 |
2927 |
5.62 |
|
|
|
| 7 |
A' |
3104 |
2916 |
30.34 |
|
|
|
| 8 |
A' |
3101 |
2912 |
32.05 |
|
|
|
| 9 |
A' |
1547 |
1453 |
10.35 |
|
|
|
| 10 |
A' |
1541 |
1447 |
1.72 |
|
|
|
| 11 |
A' |
1537 |
1444 |
3.69 |
|
|
|
| 12 |
A' |
1452 |
1363 |
10.73 |
|
|
|
| 13 |
A' |
1449 |
1360 |
0.94 |
|
|
|
| 14 |
A' |
1413 |
1327 |
0.69 |
|
|
|
| 15 |
A' |
1397 |
1312 |
9.20 |
|
|
|
| 16 |
A' |
1306 |
1227 |
4.19 |
|
|
|
| 17 |
A' |
1278 |
1201 |
0.62 |
|
|
|
| 18 |
A' |
1229 |
1154 |
0.08 |
|
|
|
| 19 |
A' |
1171 |
1100 |
0.38 |
|
|
|
| 20 |
A' |
1077 |
1011 |
0.27 |
|
|
|
| 21 |
A' |
985 |
926 |
0.78 |
|
|
|
| 22 |
A' |
979 |
919 |
0.67 |
|
|
|
| 23 |
A' |
885 |
831 |
0.56 |
|
|
|
| 24 |
A' |
827 |
777 |
0.86 |
|
|
|
| 25 |
A' |
656 |
616 |
1.89 |
|
|
|
| 26 |
A' |
445 |
418 |
0.02 |
|
|
|
| 27 |
A' |
335 |
315 |
0.13 |
|
|
|
| 28 |
A' |
129 |
121 |
0.02 |
|
|
|
| 29 |
A" |
3199 |
3005 |
30.08 |
|
|
|
| 30 |
A" |
3197 |
3003 |
25.82 |
|
|
|
| 31 |
A" |
3184 |
2990 |
13.66 |
|
|
|
| 32 |
A" |
3115 |
2925 |
73.75 |
|
|
|
| 33 |
A" |
1538 |
1444 |
4.33 |
|
|
|
| 34 |
A" |
1537 |
1443 |
0.04 |
|
|
|
| 35 |
A" |
1516 |
1424 |
4.75 |
|
|
|
| 36 |
A" |
1319 |
1239 |
0.15 |
|
|
|
| 37 |
A" |
1297 |
1218 |
0.01 |
|
|
|
| 38 |
A" |
1256 |
1180 |
0.01 |
|
|
|
| 39 |
A" |
1163 |
1092 |
0.09 |
|
|
|
| 40 |
A" |
1155 |
1084 |
0.06 |
|
|
|
| 41 |
A" |
1011 |
950 |
2.17 |
|
|
|
| 42 |
A" |
933 |
877 |
0.06 |
|
|
|
| 43 |
A" |
926 |
870 |
0.33 |
|
|
|
| 44 |
A" |
830 |
780 |
0.13 |
|
|
|
| 45 |
A" |
376 |
353 |
0.00 |
|
|
|
| 46 |
A" |
265 |
249 |
0.06 |
|
|
|
| 47 |
A" |
255 |
239 |
0.00 |
|
|
|
| 48 |
A" |
240 |
225 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37562.2 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 35278.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.294 |
0.111 |
1.073 |
| C2 |
0.294 |
0.111 |
-1.073 |
| C3 |
-0.674 |
0.662 |
0.000 |
| C4 |
0.901 |
-0.824 |
0.000 |
| C5 |
-0.996 |
2.142 |
0.000 |
| C6 |
0.294 |
-2.217 |
0.000 |
| H7 |
1.003 |
0.882 |
1.382 |
| H8 |
-0.133 |
-0.354 |
1.963 |
| H9 |
1.003 |
0.882 |
-1.382 |
| H10 |
-0.133 |
-0.354 |
-1.963 |
| H11 |
-1.599 |
0.082 |
0.000 |
| H12 |
1.990 |
-0.898 |
0.000 |
| H13 |
-0.077 |
2.729 |
0.000 |
| H14 |
-0.796 |
-2.170 |
0.000 |
| H15 |
-1.573 |
2.420 |
-0.883 |
| H16 |
-1.573 |
2.420 |
0.883 |
| H17 |
0.604 |
-2.777 |
-0.883 |
| H18 |
0.604 |
-2.777 |
0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1455 | 1.5461 | 1.5474 | 2.6340 | 2.5638 | 1.0918 | 1.0914 | 2.6687 | 3.1006 | 2.1760 | 2.2466 | 2.8527 | 2.7463 | 3.5553 | 2.9752 | 3.5020 | 2.9112 |
C2 | 2.1455 | | 1.5461 | 1.5474 | 2.6340 | 2.5638 | 2.6687 | 3.1006 | 1.0918 | 1.0914 | 2.1760 | 2.2466 | 2.8527 | 2.7463 | 2.9752 | 3.5553 | 2.9112 | 3.5020 | C3 | 1.5461 | 1.5461 | | 2.1653 | 1.5146 | 3.0374 | 2.1838 | 2.2755 | 2.1838 | 2.2755 | 1.0919 | 3.0872 | 2.1513 | 2.8343 | 2.1631 | 2.1631 | 3.7734 | 3.7734 | C4 | 1.5474 | 1.5474 | 2.1653 | | 3.5208 | 1.5199 | 2.1980 | 2.2677 | 2.1980 | 2.2677 | 2.6593 | 1.0915 | 3.6847 | 2.1659 | 4.1744 | 4.1744 | 2.1640 | 2.1640 | C5 | 2.6340 | 2.6340 | 1.5146 | 3.5208 | | 4.5459 | 2.7371 | 3.2907 | 2.7371 | 3.2907 | 2.1462 | 4.2614 | 1.0909 | 4.3162 | 1.0907 | 1.0907 | 5.2473 | 5.2473 | C6 | 2.5638 | 2.5638 | 3.0374 | 1.5199 | 4.5459 | | 3.4668 | 2.7396 | 3.4668 | 2.7396 | 2.9781 | 2.1488 | 4.9597 | 1.0905 | 5.0763 | 5.0763 | 1.0905 | 1.0905 | H7 | 1.0918 | 2.6687 | 2.1838 | 2.1980 | 2.7371 | 3.4668 | | 1.7763 | 2.7640 | 3.7424 | 3.0527 | 2.4607 | 2.5464 | 3.8025 | 3.7588 | 3.0412 | 4.3220 | 3.7146 | H8 | 1.0914 | 3.1006 | 2.2755 | 2.2677 | 3.2907 | 2.7396 | 1.7763 | | 3.7424 | 3.9256 | 2.4886 | 2.9419 | 3.6551 | 2.7547 | 4.2272 | 3.3073 | 3.8093 | 2.7529 | H9 | 2.6687 | 1.0918 | 2.1838 | 2.1980 | 2.7371 | 3.4668 | 2.7640 | 3.7424 | | 1.7763 | 3.0527 | 2.4607 | 2.5464 | 3.8025 | 3.0412 | 3.7588 | 3.7146 | 4.3220 | H10 | 3.1006 | 1.0914 | 2.2755 | 2.2677 | 3.2907 | 2.7396 | 3.7424 | 3.9256 | 1.7763 | | 2.4886 | 2.9419 | 3.6551 | 2.7547 | 3.3073 | 4.2272 | 2.7529 | 3.8093 | H11 | 2.1760 | 2.1760 | 1.0919 | 2.6593 | 2.1462 | 2.9781 | 3.0527 | 2.4886 | 3.0527 | 2.4886 | | 3.7206 | 3.0533 | 2.3908 | 2.4993 | 2.4993 | 3.7156 | 3.7156 | H12 | 2.2466 | 2.2466 | 3.0872 | 1.0915 | 4.2614 | 2.1488 | 2.4607 | 2.9419 | 2.4607 | 2.9419 | 3.7206 | | 4.1744 | 3.0624 | 4.9484 | 4.9484 | 2.4965 | 2.4965 | H13 | 2.8527 | 2.8527 | 2.1513 | 3.6847 | 1.0909 | 4.9597 | 2.5464 | 3.6551 | 2.5464 | 3.6551 | 3.0533 | 4.1744 | | 4.9509 | 1.7647 | 1.7647 | 5.6174 | 5.6174 | H14 | 2.7463 | 2.7463 | 2.8343 | 2.1659 | 4.3162 | 1.0905 | 3.8025 | 2.7547 | 3.8025 | 2.7547 | 2.3908 | 3.0624 | 4.9509 | | 4.7382 | 4.7382 | 1.7626 | 1.7626 | H15 | 3.5553 | 2.9752 | 2.1631 | 4.1744 | 1.0907 | 5.0763 | 3.7588 | 4.2272 | 3.0412 | 3.3073 | 2.4993 | 4.9484 | 1.7647 | 4.7382 | | 1.7656 | 5.6346 | 5.9048 | H16 | 2.9752 | 3.5553 | 2.1631 | 4.1744 | 1.0907 | 5.0763 | 3.0412 | 3.3073 | 3.7588 | 4.2272 | 2.4993 | 4.9484 | 1.7647 | 4.7382 | 1.7656 | | 5.9048 | 5.6346 | H17 | 3.5020 | 2.9112 | 3.7734 | 2.1640 | 5.2473 | 1.0905 | 4.3220 | 3.8093 | 3.7146 | 2.7529 | 3.7156 | 2.4965 | 5.6174 | 1.7626 | 5.6346 | 5.9048 | | 1.7660 | H18 | 2.9112 | 3.5020 | 3.7734 | 2.1640 | 5.2473 | 1.0905 | 3.7146 | 2.7529 | 4.3220 | 3.8093 | 3.7156 | 2.4965 | 5.6174 | 1.7626 | 5.9048 | 5.6346 | 1.7660 | |
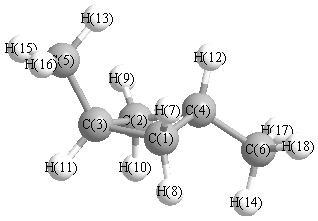 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
87.869 |
|
C1 |
C3 |
C5 |
118.760 |
| C1 |
C3 |
H11 |
109.954 |
|
C1 |
C4 |
C2 |
87.777 |
| C1 |
C4 |
C6 |
113.407 |
|
C1 |
C4 |
H12 |
115.636 |
| C2 |
C3 |
C5 |
118.760 |
|
C2 |
C3 |
H11 |
109.954 |
| C2 |
C4 |
C6 |
113.407 |
|
C2 |
C4 |
H12 |
115.636 |
| C3 |
C1 |
C4 |
88.847 |
|
C3 |
C1 |
H7 |
110.577 |
| C3 |
C1 |
H8 |
118.235 |
|
C3 |
C2 |
C4 |
88.847 |
| C3 |
C2 |
H9 |
110.577 |
|
C3 |
C2 |
H10 |
118.235 |
| C3 |
C5 |
H13 |
110.255 |
|
C3 |
C5 |
H15 |
111.216 |
| C3 |
C5 |
H16 |
111.216 |
|
C4 |
C1 |
H7 |
111.615 |
| C4 |
C1 |
H8 |
117.446 |
|
C4 |
C2 |
H9 |
111.615 |
| C4 |
C2 |
H10 |
117.446 |
|
C4 |
C6 |
H14 |
111.065 |
| C4 |
C6 |
H17 |
110.921 |
|
C4 |
C6 |
H18 |
110.921 |
| C5 |
C3 |
H11 |
109.790 |
|
C6 |
C4 |
H12 |
109.656 |
| H7 |
C1 |
H8 |
108.903 |
|
H9 |
C2 |
H10 |
108.903 |
| H13 |
C5 |
H15 |
107.978 |
|
H13 |
C5 |
H16 |
107.978 |
| H14 |
C6 |
H17 |
107.828 |
|
H14 |
C6 |
H18 |
107.828 |
| H15 |
C5 |
H16 |
108.072 |
|
H17 |
C6 |
H18 |
108.142 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
